return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11Cl (Pentane, 1-chloro-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-656.526594
Energy at 298.15K-656.538748
HF Energy-655.342590
Nuclear repulsion energy285.366730
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3166 3006 26.51      
2 A' 3132 2973 22.58      
3 A' 3099 2942 19.82      
4 A' 3089 2933 37.68      
5 A' 3086 2929 11.68      
6 A' 3068 2913 9.23      
7 A' 1534 1456 8.74      
8 A' 1524 1447 1.62      
9 A' 1513 1436 1.58      
10 A' 1506 1430 0.29      
11 A' 1502 1426 0.05      
12 A' 1425 1353 4.19      
13 A' 1414 1342 0.06      
14 A' 1382 1312 14.08      
15 A' 1327 1260 11.69      
16 A' 1259 1195 4.89      
17 A' 1154 1096 1.08      
18 A' 1106 1050 0.17      
19 A' 1091 1036 6.47      
20 A' 1050 997 1.05      
21 A' 927 880 1.04      
22 A' 787 747 35.60      
23 A' 455 432 2.32      
24 A' 346 329 0.56      
25 A' 263 250 1.39      
26 A' 114 108 1.09      
27 A" 3181 3020 12.41      
28 A" 3159 2999 38.21      
29 A" 3136 2977 18.94      
30 A" 3118 2961 10.14      
31 A" 3092 2936 2.61      
32 A" 1528 1451 7.72      
33 A" 1350 1282 0.68      
34 A" 1341 1273 0.29      
35 A" 1305 1239 0.16      
36 A" 1237 1175 0.13      
37 A" 1124 1067 0.69      
38 A" 982 932 0.02      
39 A" 851 808 0.75      
40 A" 769 730 0.10      
41 A" 749 711 3.67      
42 A" 263 249 0.00      
43 A" 158 150 0.44      
44 A" 103 98 0.20      
45 A" 70 67 1.16      

Unscaled Zero Point Vibrational Energy (zpe) 33916.8 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 32200.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.50877 0.02677 0.02604

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.479 0.097 0.000
Cl2 -2.423 1.615 0.000
C3 0.000 0.375 0.000
C4 0.797 -0.916 0.000
C5 2.293 -0.670 0.000
C6 3.089 -1.960 0.000
H7 4.156 -1.765 0.000
H8 -1.779 -0.453 0.883
H9 -1.779 -0.453 -0.883
H10 0.251 0.968 -0.875
H11 0.251 0.968 0.875
H12 0.532 -1.509 0.874
H13 0.532 -1.509 -0.874
H14 2.555 -0.076 0.873
H15 2.555 -0.076 -0.873
H16 2.856 -2.556 0.878
H17 2.856 -2.556 -0.878

Atom - Atom Distances (Å)
  C1 Cl2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.78731.50502.49143.84885.00965.93471.08261.08262.12562.12562.71792.71794.13154.13155.15815.1581
Cl21.78732.72144.09525.23956.56927.39592.33892.33892.88672.88674.38754.38755.32955.32956.78496.7849
C31.50502.72141.51712.51953.87214.67462.15172.15171.08671.08672.14382.14382.73792.73794.18594.1859
C42.49144.09521.51711.51552.51833.46452.76222.76222.14782.14781.08921.08922.13492.13492.77522.7752
C53.84885.23952.51951.51551.51612.16134.17174.17172.75992.75992.13722.13721.08771.08772.15562.1556
C65.00966.56923.87212.51831.51611.08535.17145.17144.17054.17052.73952.73952.14362.14361.08621.0862
H75.93477.39594.67463.46452.16131.08536.14196.14194.84604.84603.73693.73692.48482.48481.75691.7569
H81.08262.33892.15172.76224.17175.17146.14191.76593.03842.47792.54053.08904.35074.69165.09005.3859
H91.08262.33892.15172.76224.17175.17146.14191.76592.47793.03843.08902.54054.69164.35075.38595.0900
H102.12562.88671.08672.14782.75994.17054.84603.03842.47791.75083.04532.49263.07532.53014.72034.3827
H112.12562.88671.08672.14782.75994.17054.84602.47793.03841.75082.49263.04532.53013.07534.38274.7203
H122.71794.38752.14381.08922.13722.73953.73692.54053.08903.04532.49261.74852.47923.03292.54953.0935
H132.71794.38752.14381.08922.13722.73953.73693.08902.54052.49263.04531.74853.03292.47923.09352.5495
H144.13155.32952.73792.13491.08772.14362.48484.35074.69163.07532.53012.47923.03291.74562.49813.0504
H154.13155.32952.73792.13491.08772.14362.48484.69164.35072.53013.07533.03292.47921.74563.05042.4981
H165.15816.78494.18592.77522.15561.08621.75695.09005.38594.72034.38272.54953.09352.49813.05041.7557
H175.15816.78494.18592.77522.15561.08621.75695.38595.09004.38274.72033.09352.54953.05042.49811.7557

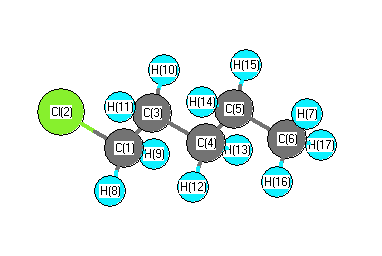
picture of Pentane, 1-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C4 111.051 C1 C3 H10 109.137
C1 C3 H11 109.137 Cl2 C1 C3 111.216
Cl2 C1 H8 106.579 Cl2 C1 H9 106.579
C3 C1 H8 111.471 C3 C1 H9 111.471
C3 C4 C5 112.363 C3 C4 H12 109.588
C3 C4 H13 109.588 C4 C3 H10 110.050
C4 C3 H11 110.050 C4 C5 C6 112.335
C4 C5 H14 109.088 C4 C5 H15 109.088
C5 C4 H12 109.185 C5 C4 H13 109.185
C5 C6 H7 111.288 C5 C6 H16 110.775
C5 C6 H17 110.775 C6 C5 H14 109.724
C6 C5 H15 109.724 H7 C6 H16 108.016
H7 C6 H17 108.016 H8 C1 H9 109.288
H10 C3 H11 107.332 H12 C4 H13 106.773
H14 C5 H15 106.718 H16 C6 H17 107.837
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability