Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -323.616097 |
| Energy at 298.15K | |
| HF Energy | -323.616097 |
| Nuclear repulsion energy | 236.084443 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3140 |
3006 |
25.52 |
|
|
|
| 2 |
A' |
3071 |
2939 |
37.28 |
|
|
|
| 3 |
A' |
3057 |
2926 |
25.14 |
|
|
|
| 4 |
A' |
3054 |
2923 |
10.40 |
|
|
|
| 5 |
A' |
1790 |
1713 |
240.46 |
|
|
|
| 6 |
A' |
1521 |
1456 |
13.65 |
|
|
|
| 7 |
A' |
1507 |
1442 |
1.62 |
|
|
|
| 8 |
A' |
1494 |
1430 |
2.00 |
|
|
|
| 9 |
A' |
1418 |
1357 |
17.76 |
|
|
|
| 10 |
A' |
1415 |
1355 |
2.87 |
|
|
|
| 11 |
A' |
1331 |
1274 |
8.53 |
|
|
|
| 12 |
A' |
1172 |
1121 |
7.29 |
|
|
|
| 13 |
A' |
1098 |
1050 |
34.80 |
|
|
|
| 14 |
A' |
1061 |
1016 |
3.10 |
|
|
|
| 15 |
A' |
939 |
899 |
163.15 |
|
|
|
| 16 |
A' |
875 |
837 |
414.24 |
|
|
|
| 17 |
A' |
687 |
657 |
31.63 |
|
|
|
| 18 |
A' |
392 |
375 |
0.04 |
|
|
|
| 19 |
A' |
350 |
335 |
1.13 |
|
|
|
| 20 |
A' |
153 |
146 |
0.50 |
|
|
|
| 21 |
A" |
3132 |
2998 |
61.37 |
|
|
|
| 22 |
A" |
3112 |
2979 |
3.74 |
|
|
|
| 23 |
A" |
3092 |
2960 |
3.72 |
|
|
|
| 24 |
A" |
1502 |
1437 |
7.78 |
|
|
|
| 25 |
A" |
1320 |
1264 |
0.05 |
|
|
|
| 26 |
A" |
1269 |
1215 |
0.44 |
|
|
|
| 27 |
A" |
1185 |
1134 |
0.68 |
|
|
|
| 28 |
A" |
893 |
855 |
0.97 |
|
|
|
| 29 |
A" |
758 |
725 |
2.57 |
|
|
|
| 30 |
A" |
235 |
225 |
0.01 |
|
|
|
| 31 |
A" |
215 |
206 |
0.59 |
|
|
|
| 32 |
A" |
99 |
95 |
1.50 |
|
|
|
| 33 |
A" |
52i |
50i |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23141.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22149.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.898 |
2.198 |
0.000 |
| C2 |
-1.499 |
0.733 |
0.000 |
| C3 |
0.000 |
0.571 |
0.000 |
| O4 |
0.290 |
-0.831 |
0.000 |
| N5 |
1.663 |
-1.034 |
0.000 |
| O6 |
1.920 |
-2.172 |
0.000 |
| H7 |
-2.980 |
2.306 |
0.000 |
| H8 |
-1.515 |
2.713 |
0.879 |
| H9 |
-1.515 |
2.713 |
-0.879 |
| H10 |
-1.908 |
0.228 |
-0.874 |
| H11 |
-1.908 |
0.228 |
0.874 |
| H12 |
0.444 |
1.029 |
0.884 |
| H13 |
0.444 |
1.029 |
-0.884 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5180 | 2.4999 | 3.7359 | 4.8091 | 5.8026 | 1.0870 | 1.0891 | 1.0891 | 2.1547 | 2.1547 | 2.7627 | 2.7627 |
C2 | 1.5180 | | 1.5074 | 2.3755 | 3.6221 | 4.4861 | 2.1601 | 2.1665 | 2.1665 | 1.0894 | 1.0894 | 2.1543 | 2.1543 | C3 | 2.4999 | 1.5074 | | 1.4311 | 2.3113 | 3.3478 | 3.4480 | 2.7671 | 2.7671 | 2.1265 | 2.1265 | 1.0900 | 1.0900 | O4 | 3.7359 | 2.3755 | 1.4311 | | 1.3884 | 2.1111 | 4.5304 | 4.0727 | 4.0727 | 2.5913 | 2.5913 | 2.0645 | 2.0645 | N5 | 4.8091 | 3.6221 | 2.3113 | 1.3884 | | 1.1659 | 5.7193 | 4.9914 | 4.9914 | 3.8872 | 3.8872 | 2.5543 | 2.5543 | O6 | 5.8026 | 4.4861 | 3.3478 | 2.1111 | 1.1659 | | 6.6373 | 6.0359 | 6.0359 | 4.6020 | 4.6020 | 3.6336 | 3.6336 | H7 | 1.0870 | 2.1601 | 3.4480 | 4.5304 | 5.7193 | 6.6373 | | 1.7567 | 1.7567 | 2.4955 | 2.4955 | 3.7593 | 3.7593 | H8 | 1.0891 | 2.1665 | 2.7671 | 4.0727 | 4.9914 | 6.0359 | 1.7567 | | 1.7587 | 3.0666 | 2.5157 | 2.5832 | 3.1276 | H9 | 1.0891 | 2.1665 | 2.7671 | 4.0727 | 4.9914 | 6.0359 | 1.7567 | 1.7587 | | 2.5157 | 3.0666 | 3.1276 | 2.5832 | H10 | 2.1547 | 1.0894 | 2.1265 | 2.5913 | 3.8872 | 4.6020 | 2.4955 | 3.0666 | 2.5157 | | 1.7485 | 3.0434 | 2.4841 | H11 | 2.1547 | 1.0894 | 2.1265 | 2.5913 | 3.8872 | 4.6020 | 2.4955 | 2.5157 | 3.0666 | 1.7485 | | 2.4841 | 3.0434 | H12 | 2.7627 | 2.1543 | 1.0900 | 2.0645 | 2.5543 | 3.6336 | 3.7593 | 2.5832 | 3.1276 | 3.0434 | 2.4841 | | 1.7680 | H13 | 2.7627 | 2.1543 | 1.0900 | 2.0645 | 2.5543 | 3.6336 | 3.7593 | 3.1276 | 2.5832 | 2.4841 | 3.0434 | 1.7680 | |
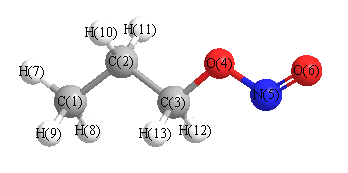 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.446 |
|
C1 |
C2 |
H10 |
110.374 |
| C1 |
C2 |
H11 |
110.374 |
|
C2 |
C1 |
H7 |
110.953 |
| C2 |
C1 |
H8 |
111.342 |
|
C2 |
C1 |
H9 |
111.342 |
| C2 |
C3 |
O4 |
107.859 |
|
C2 |
C3 |
H12 |
111.067 |
| C2 |
C3 |
H13 |
111.067 |
|
C3 |
C2 |
H10 |
108.885 |
| C3 |
C2 |
H11 |
108.885 |
|
C3 |
O4 |
N5 |
110.112 |
| O4 |
C3 |
H12 |
109.209 |
|
O4 |
C3 |
H13 |
109.209 |
| O4 |
N5 |
O6 |
111.179 |
|
H7 |
C1 |
H8 |
107.664 |
| H7 |
C1 |
H9 |
107.664 |
|
H8 |
C1 |
H9 |
107.697 |
| H10 |
C2 |
H11 |
106.743 |
|
H12 |
C3 |
H13 |
108.400 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.367 |
|
|
|
| 2 |
C |
-0.219 |
|
|
|
| 3 |
C |
-0.036 |
|
|
|
| 4 |
O |
-0.170 |
|
|
|
| 5 |
N |
0.106 |
|
|
|
| 6 |
O |
-0.120 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.115 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
H |
0.126 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.099 |
|
|
|
| 13 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.445 |
2.428 |
0.000 |
2.825 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.757 |
1.578 |
0.000 |
| y |
1.578 |
-39.508 |
0.000 |
| z |
0.000 |
0.000 |
-34.617 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.694 |
1.578 |
0.000 |
| y |
1.578 |
-3.321 |
0.000 |
| z |
0.000 |
0.000 |
4.015 |
|
| Polar |
|---|
| 3z2-r2 | 8.030 |
|---|
| x2-y2 | 1.751 |
|---|
| xy | 1.578 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.745 |
-2.191 |
0.000 |
| y |
-2.191 |
9.055 |
0.000 |
| z |
0.000 |
0.000 |
6.103 |
<r2> (average value of r
2) Å
2
| <r2> |
240.843 |
| (<r2>)1/2 |
15.519 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -323.617497 |
| Energy at 298.15K | -323.626526 |
| HF Energy | -323.617497 |
| Nuclear repulsion energy | 243.128345 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3140 |
3005 |
26.56 |
|
|
|
| 2 |
A |
3132 |
2998 |
37.03 |
|
|
|
| 3 |
A |
3118 |
2984 |
20.51 |
|
|
|
| 4 |
A |
3097 |
2964 |
12.69 |
|
|
|
| 5 |
A |
3063 |
2932 |
34.90 |
|
|
|
| 6 |
A |
3060 |
2928 |
19.05 |
|
|
|
| 7 |
A |
3058 |
2927 |
9.38 |
|
|
|
| 8 |
A |
1796 |
1719 |
253.09 |
|
|
|
| 9 |
A |
1507 |
1442 |
8.30 |
|
|
|
| 10 |
A |
1500 |
1436 |
8.33 |
|
|
|
| 11 |
A |
1488 |
1424 |
3.23 |
|
|
|
| 12 |
A |
1478 |
1414 |
2.31 |
|
|
|
| 13 |
A |
1419 |
1358 |
6.52 |
|
|
|
| 14 |
A |
1400 |
1340 |
8.98 |
|
|
|
| 15 |
A |
1381 |
1322 |
0.86 |
|
|
|
| 16 |
A |
1307 |
1251 |
4.58 |
|
|
|
| 17 |
A |
1291 |
1236 |
10.73 |
|
|
|
| 18 |
A |
1189 |
1138 |
10.57 |
|
|
|
| 19 |
A |
1126 |
1078 |
12.08 |
|
|
|
| 20 |
A |
1103 |
1056 |
25.82 |
|
|
|
| 21 |
A |
1018 |
974 |
6.75 |
|
|
|
| 22 |
A |
935 |
895 |
118.09 |
|
|
|
| 23 |
A |
888 |
850 |
1.37 |
|
|
|
| 24 |
A |
856 |
819 |
359.51 |
|
|
|
| 25 |
A |
783 |
750 |
20.71 |
|
|
|
| 26 |
A |
630 |
603 |
30.37 |
|
|
|
| 27 |
A |
454 |
434 |
5.47 |
|
|
|
| 28 |
A |
382 |
366 |
3.26 |
|
|
|
| 29 |
A |
280 |
268 |
0.64 |
|
|
|
| 30 |
A |
244 |
234 |
0.53 |
|
|
|
| 31 |
A |
183 |
175 |
0.45 |
|
|
|
| 32 |
A |
106 |
102 |
0.83 |
|
|
|
| 33 |
A |
47 |
45 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23229.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22233.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.175 |
-0.914 |
0.181 |
| C2 |
-1.748 |
0.457 |
-0.312 |
| C3 |
-0.405 |
0.887 |
0.223 |
| O4 |
0.572 |
-0.030 |
-0.289 |
| N5 |
1.783 |
0.204 |
0.311 |
| O6 |
2.559 |
-0.550 |
-0.141 |
| H7 |
-3.153 |
-1.181 |
-0.214 |
| H8 |
-2.237 |
-0.935 |
1.268 |
| H9 |
-1.466 |
-1.676 |
-0.130 |
| H10 |
-1.712 |
0.474 |
-1.401 |
| H11 |
-2.473 |
1.211 |
-0.008 |
| H12 |
-0.144 |
1.891 |
-0.108 |
| H13 |
-0.378 |
0.853 |
1.311 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5186 | 2.5265 | 2.9240 | 4.1153 | 4.7588 | 1.0874 | 1.0891 | 1.0868 | 2.1552 | 2.1546 | 3.4754 | 2.7621 |
C2 | 1.5186 | | 1.5083 | 2.3703 | 3.5941 | 4.4257 | 2.1598 | 2.1620 | 2.1596 | 1.0894 | 1.0902 | 2.1609 | 2.1600 | C3 | 2.5265 | 1.5083 | | 1.4346 | 2.2936 | 3.3135 | 3.4668 | 2.7880 | 2.7969 | 2.1252 | 2.1067 | 1.0883 | 1.0891 | O4 | 2.9240 | 2.3703 | 1.4346 | | 1.3719 | 2.0588 | 3.8989 | 3.3372 | 2.6242 | 2.5900 | 3.3009 | 2.0588 | 2.0603 | N5 | 4.1153 | 3.5941 | 2.2936 | 1.3719 | | 1.1725 | 5.1530 | 4.2869 | 3.7793 | 3.9014 | 4.3857 | 2.5955 | 2.4685 | O6 | 4.7588 | 4.4257 | 3.3135 | 2.0588 | 1.1725 | | 5.7464 | 5.0138 | 4.1789 | 4.5691 | 5.3331 | 3.6424 | 3.5643 | H7 | 1.0874 | 2.1598 | 3.4668 | 3.8989 | 5.1530 | 5.7464 | | 1.7595 | 1.7605 | 2.4941 | 2.4949 | 4.3008 | 3.7628 | H8 | 1.0891 | 2.1620 | 2.7880 | 3.3372 | 4.2869 | 5.0138 | 1.7595 | | 1.7608 | 3.0635 | 2.5077 | 3.7762 | 2.5797 | H9 | 1.0868 | 2.1596 | 2.7969 | 2.6242 | 3.7793 | 4.1789 | 1.7605 | 1.7608 | | 2.5103 | 3.0609 | 3.8044 | 3.1075 | H10 | 2.1552 | 1.0894 | 2.1252 | 2.5900 | 3.9014 | 4.5691 | 2.4941 | 3.0635 | 2.5103 | | 1.7502 | 2.4776 | 3.0457 | H11 | 2.1546 | 1.0902 | 2.1067 | 3.3009 | 4.3857 | 5.3331 | 2.4949 | 2.5077 | 3.0609 | 1.7502 | | 2.4285 | 2.5012 | H12 | 3.4754 | 2.1609 | 1.0883 | 2.0588 | 2.5955 | 3.6424 | 4.3008 | 3.7762 | 3.8044 | 2.4776 | 2.4285 | | 1.7732 | H13 | 2.7621 | 2.1600 | 1.0891 | 2.0603 | 2.4685 | 3.5643 | 3.7628 | 2.5797 | 3.1075 | 3.0457 | 2.5012 | 1.7732 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.160 |
|
C1 |
C2 |
H10 |
110.366 |
| C1 |
C2 |
H11 |
110.277 |
|
C2 |
C1 |
H7 |
110.859 |
| C2 |
C1 |
H8 |
110.927 |
|
C2 |
C1 |
H9 |
110.881 |
| C2 |
C3 |
O4 |
107.281 |
|
C2 |
C3 |
H12 |
111.629 |
| C2 |
C3 |
H13 |
111.511 |
|
C3 |
C2 |
H10 |
108.715 |
| C3 |
C2 |
H11 |
107.245 |
|
C3 |
O4 |
N5 |
109.601 |
| O4 |
C3 |
H12 |
108.601 |
|
O4 |
C3 |
H13 |
108.676 |
| O4 |
N5 |
O6 |
107.776 |
|
H7 |
C1 |
H8 |
107.879 |
| H7 |
C1 |
H9 |
108.133 |
|
H8 |
C1 |
H9 |
108.038 |
| H10 |
C2 |
H11 |
106.834 |
|
H12 |
C3 |
H13 |
109.047 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.370 |
|
|
|
| 2 |
C |
-0.198 |
|
|
|
| 3 |
C |
-0.071 |
|
|
|
| 4 |
O |
-0.171 |
|
|
|
| 5 |
N |
0.106 |
|
|
|
| 6 |
O |
-0.123 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.117 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.114 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
| 13 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.401 |
-0.874 |
0.703 |
2.650 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.510 |
0.491 |
-0.643 |
| y |
0.491 |
-34.789 |
-0.807 |
| z |
-0.643 |
-0.807 |
-36.836 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.697 |
0.491 |
-0.643 |
| y |
0.491 |
2.884 |
-0.807 |
| z |
-0.643 |
-0.807 |
-0.186 |
|
| Polar |
|---|
| 3z2-r2 | -0.373 |
|---|
| x2-y2 | -3.721 |
|---|
| xy | 0.491 |
|---|
| xz | -0.643 |
|---|
| yz | -0.807 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.705 |
-0.991 |
-0.009 |
| y |
-0.991 |
7.431 |
-0.024 |
| z |
-0.009 |
-0.024 |
6.461 |
<r2> (average value of r
2) Å
2
| <r2> |
192.797 |
| (<r2>)1/2 |
13.885 |
