Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -93.793257 |
| Energy at 298.15K | -93.794510 |
| HF Energy | -93.430231 |
| Nuclear repulsion energy | 28.683319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3490 |
3313 |
22.82 |
|
|
|
| 2 |
A' |
3073 |
2917 |
238.76 |
|
|
|
| 3 |
A' |
2421 |
2298 |
367.20 |
|
|
|
| 4 |
A' |
1011 |
960 |
93.58 |
|
|
|
| 5 |
A' |
732 |
695 |
317.37 |
|
|
|
| 6 |
A" |
917 |
871 |
6.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5821.4 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 5526.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.106 |
0.627 |
0.000 |
| N2 |
0.106 |
-0.571 |
0.000 |
| H3 |
-0.663 |
1.400 |
0.000 |
| H4 |
-0.716 |
-1.166 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.1976 | 1.0909 | 1.9728 |
N2 | 1.1976 | | 2.1158 | 1.0153 | H3 | 1.0909 | 2.1158 | | 2.5672 | H4 | 1.9728 | 1.0153 | 2.5672 | |
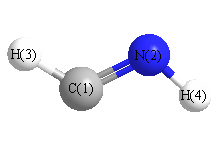 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
125.925 |
|
H3 |
C1 |
N2 |
135.143 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -93.801222 |
| Energy at 298.15K | -93.802540 |
| HF Energy | -93.436257 |
| Nuclear repulsion energy | 28.598225 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3579 |
3398 |
62.20 |
|
|
|
| 2 |
A' |
3103 |
2946 |
146.78 |
|
|
|
| 3 |
A' |
2410 |
2288 |
390.42 |
|
|
|
| 4 |
A' |
1185 |
1125 |
6.63 |
|
|
|
| 5 |
A' |
821 |
779 |
428.22 |
|
|
|
| 6 |
A" |
962 |
914 |
169.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6030.0 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 5724.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.003 |
0.639 |
0.000 |
| N2 |
-0.003 |
-0.569 |
0.000 |
| H3 |
0.892 |
1.260 |
0.000 |
| H4 |
-0.854 |
-1.115 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2083 | 1.0893 | 1.9502 |
N2 | 1.2083 | | 2.0362 | 1.0113 | H3 | 1.0893 | 2.0362 | | 2.9482 | H4 | 1.9502 | 1.0113 | 2.9482 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
122.710 |
|
H3 |
C1 |
N2 |
124.730 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
