Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -93.826919 |
| Energy at 298.15K | -93.828188 |
| HF Energy | -93.431042 |
| Nuclear repulsion energy | 28.541698 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3424 |
3297 |
|
|
|
|
| 2 |
A' |
3025 |
2913 |
|
|
|
|
| 3 |
A' |
2320 |
2234 |
|
|
|
|
| 4 |
A' |
1013 |
976 |
|
|
|
|
| 5 |
A' |
773 |
745 |
|
|
|
|
| 6 |
A" |
918 |
884 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5736.9 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 5524.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.108 |
0.629 |
0.000 |
| N2 |
0.108 |
-0.577 |
0.000 |
| H3 |
-0.660 |
1.407 |
0.000 |
| H4 |
-0.740 |
-1.142 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2057 | 1.0929 | 1.9630 |
N2 | 1.2057 | | 2.1269 | 1.0189 | H3 | 1.0929 | 2.1269 | | 2.5494 | H4 | 1.9630 | 1.0189 | 2.5494 | |
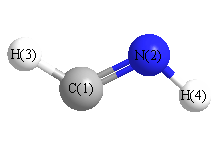 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
123.657 |
|
H3 |
C1 |
N2 |
135.371 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -93.834963 |
| Energy at 298.15K | -93.836288 |
| HF Energy | -93.436748 |
| Nuclear repulsion energy | 28.466931 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3534 |
3403 |
|
|
|
|
| 2 |
A' |
3052 |
2939 |
|
|
|
|
| 3 |
A' |
2323 |
2237 |
|
|
|
|
| 4 |
A' |
1189 |
1145 |
|
|
|
|
| 5 |
A' |
838 |
807 |
|
|
|
|
| 6 |
A" |
967 |
931 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5951.3 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 5730.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.002 |
0.642 |
0.000 |
| N2 |
-0.002 |
-0.574 |
0.000 |
| H3 |
0.895 |
1.265 |
0.000 |
| H4 |
-0.870 |
-1.098 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2157 | 1.0920 | 1.9440 |
N2 | 1.2157 | | 2.0461 | 1.0136 | H3 | 1.0920 | 2.0461 | | 2.9491 | H4 | 1.9440 | 1.0136 | 2.9491 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
121.122 |
|
H3 |
C1 |
N2 |
124.822 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
