return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (Methyl propyl ether)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-233.523105
Energy at 298.15K 
HF Energy-232.227353
Nuclear repulsion energy190.134842
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3183 3022 24.02      
2 A' 3172 3012 23.64      
3 A' 3109 2952 22.08      
4 A' 3091 2935 20.10      
5 A' 3040 2886 64.30      
6 A' 3014 2861 40.13      
7 A' 1553 1474 2.85      
8 A' 1537 1459 10.59      
9 A' 1532 1454 2.55      
10 A' 1517 1440 1.27      
11 A' 1496 1421 0.33      
12 A' 1443 1370 18.52      
13 A' 1426 1354 2.71      
14 A' 1347 1279 1.90      
15 A' 1248 1185 43.06      
16 A' 1197 1137 124.91      
17 A' 1148 1090 15.57      
18 A' 1091 1036 3.15      
19 A' 1012 961 21.24      
20 A' 927 880 6.75      
21 A' 445 423 0.71      
22 A' 412 391 2.97      
23 A' 191 181 1.31      
24 A" 3164 3004 43.28      
25 A" 3140 2981 1.75      
26 A" 3091 2935 47.62      
27 A" 3039 2885 51.34      
28 A" 1530 1452 7.36      
29 A" 1520 1443 6.91      
30 A" 1336 1269 0.29      
31 A" 1288 1222 1.83      
32 A" 1215 1153 5.85      
33 A" 1187 1127 0.22      
34 A" 916 869 1.50      
35 A" 780 740 1.37      
36 A" 252 239 1.11      
37 A" 239 227 2.09      
38 A" 118 112 1.75      
39 A" 108 103 1.26      

Unscaled Zero Point Vibrational Energy (zpe) 30524.7 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 28980.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.66692 0.07220 0.06845

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.292 0.664 0.000
O2 -1.266 -0.296 0.000
C3 0.000 0.316 0.000
C4 1.062 -0.751 0.000
C5 2.455 -0.151 0.000
H6 -3.237 0.136 0.000
H7 -2.240 1.301 0.885
H8 -2.240 1.301 -0.885
H9 0.109 0.956 0.881
H10 0.109 0.956 -0.881
H11 0.919 -1.380 -0.873
H12 0.919 -1.380 0.873
H13 2.612 0.470 0.877
H14 2.612 0.470 -0.877
H15 3.216 -0.924 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.40462.31793.63984.81601.08271.09141.09142.57402.57403.90493.90494.98514.98515.7316
O21.40461.40662.37243.72422.01732.06922.06922.05832.05832.59102.59104.04954.04954.5259
C32.31791.40661.50522.49893.24182.60222.60221.09451.09452.11742.11742.75962.75963.4464
C43.63982.37241.50521.51654.38943.98703.98702.14392.14391.08601.08602.15912.15912.1606
C54.81603.72422.49891.51655.69904.99344.99342.73932.73932.15242.15241.08631.08631.0843
H61.08272.01733.24184.38945.69901.77001.77003.55593.55594.50914.50915.92365.92366.5390
H71.09142.06922.60223.98704.99341.77001.76952.37462.95924.50094.14334.92275.22865.9580
H81.09142.06922.60223.98704.99341.77001.76952.95922.37464.14334.50095.22864.92275.9580
H92.57402.05831.09452.14392.73933.55592.37462.95921.76233.03162.47232.54943.09713.7362
H102.57402.05831.09452.14392.73933.55592.95922.37461.76232.47233.03163.09712.54943.7362
H113.90492.59102.11741.08602.15244.50914.50094.14333.03162.47231.74683.05852.50772.4993
H123.90492.59102.11741.08602.15244.50914.14334.50092.47233.03161.74682.50773.05852.4993
H134.98514.04952.75962.15911.08635.92364.92275.22862.54943.09713.05852.50771.75491.7540
H144.98514.04952.75962.15911.08635.92365.22864.92273.09712.54942.50773.05851.75491.7540
H155.73164.52593.44642.16061.08436.53905.95805.95803.73623.73622.49932.49931.75401.7540

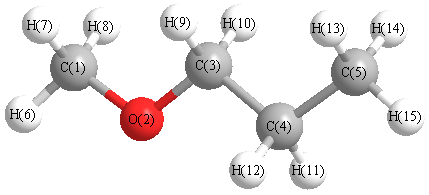
picture of Methyl propyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 111.080 O2 C1 H6 107.693
O2 C1 H7 111.378 O2 C1 H8 111.378
O2 C3 C4 109.081 O2 C3 H9 110.139
O2 C3 H10 110.139 C3 C4 C5 111.584
C3 C4 H11 108.522 C3 C4 H12 108.522
C4 C3 H9 110.114 C4 C3 H10 110.114
C4 C5 H13 111.024 C4 C5 H14 111.024
C4 C5 H15 111.266 C5 C4 H11 110.503
C5 C4 H12 110.503 H6 C1 H7 109.007
H6 C1 H8 109.007 H7 C1 H8 108.328
H9 C3 H10 107.240 H11 C4 H12 107.069
H13 C5 H14 107.743 H13 C5 H15 107.813
H14 C5 H15 107.813
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability