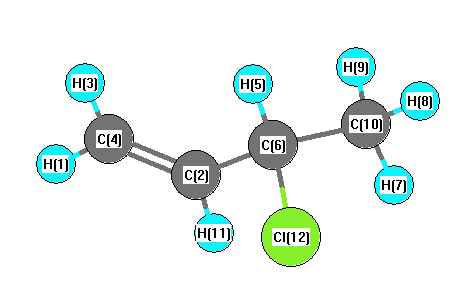
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3250 |
3110 |
9.91 |
|
|
|
| 2 |
A |
3176 |
3039 |
4.53 |
|
|
|
| 3 |
A |
3159 |
3023 |
11.17 |
|
|
|
| 4 |
A |
3156 |
3021 |
10.71 |
|
|
|
| 5 |
A |
3141 |
3006 |
15.90 |
|
|
|
| 6 |
A |
3120 |
2986 |
1.10 |
|
|
|
| 7 |
A |
3062 |
2931 |
13.37 |
|
|
|
| 8 |
A |
1728 |
1654 |
2.85 |
|
|
|
| 9 |
A |
1491 |
1427 |
1.63 |
|
|
|
| 10 |
A |
1485 |
1421 |
9.47 |
|
|
|
| 11 |
A |
1457 |
1394 |
9.17 |
|
|
|
| 12 |
A |
1405 |
1345 |
7.19 |
|
|
|
| 13 |
A |
1334 |
1277 |
1.75 |
|
|
|
| 14 |
A |
1314 |
1258 |
1.70 |
|
|
|
| 15 |
A |
1253 |
1199 |
19.28 |
|
|
|
| 16 |
A |
1201 |
1149 |
10.73 |
|
|
|
| 17 |
A |
1116 |
1068 |
2.41 |
|
|
|
| 18 |
A |
1044 |
999 |
36.66 |
|
|
|
| 19 |
A |
1031 |
987 |
5.06 |
|
|
|
| 20 |
A |
979 |
937 |
10.78 |
|
|
|
| 21 |
A |
968 |
926 |
35.11 |
|
|
|
| 22 |
A |
881 |
843 |
8.08 |
|
|
|
| 23 |
A |
728 |
696 |
35.73 |
|
|
|
| 24 |
A |
642 |
615 |
12.78 |
|
|
|
| 25 |
A |
458 |
438 |
5.10 |
|
|
|
| 26 |
A |
312 |
298 |
1.35 |
|
|
|
| 27 |
A |
297 |
285 |
1.27 |
|
|
|
| 28 |
A |
278 |
266 |
4.47 |
|
|
|
| 29 |
A |
246 |
235 |
0.28 |
|
|
|
| 30 |
A |
105 |
100 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21907.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20967.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.138 |
|
|
|
| 2 |
C |
-0.082 |
|
|
|
| 3 |
H |
0.130 |
|
|
|
| 4 |
C |
-0.311 |
|
|
|
| 5 |
H |
0.147 |
|
|
|
| 6 |
C |
-0.058 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
C |
-0.329 |
|
|
|
| 11 |
H |
0.154 |
|
|
|
| 12 |
Cl |
-0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.515 |
1.446 |
-0.297 |
2.116 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.484 |
-2.055 |
-0.092 |
| y |
-2.055 |
-39.196 |
-0.257 |
| z |
-0.092 |
-0.257 |
-36.400 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.685 |
-2.055 |
-0.092 |
| y |
-2.055 |
-1.754 |
-0.257 |
| z |
-0.092 |
-0.257 |
2.439 |
|
| Polar |
|---|
| 3z2-r2 | 4.879 |
|---|
| x2-y2 | 0.712 |
|---|
| xy | -2.055 |
|---|
| xz | -0.092 |
|---|
| yz | -0.257 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.245 |
-0.249 |
-0.569 |
| y |
-0.249 |
8.187 |
0.026 |
| z |
-0.569 |
0.026 |
7.414 |
<r2> (average value of r
2) Å
2
| <r2> |
172.249 |
| (<r2>)1/2 |
13.124 |