return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-235.395818
Energy at 298.15K-235.408188
HF Energy-234.260200
Nuclear repulsion energy245.880379
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3268 3103 13.07      
2 A' 3192 3031 7.70      
3 A' 3170 3010 18.03      
4 A' 3163 3003 20.70      
5 A' 3157 2997 53.92      
6 A' 3087 2931 19.53      
7 A' 3084 2928 13.17      
8 A' 3081 2925 13.39      
9 A' 1718 1631 11.73      
10 A' 1535 1458 5.63      
11 A' 1531 1453 9.73      
12 A' 1516 1440 12.77      
13 A' 1469 1395 0.96      
14 A' 1421 1349 4.83      
15 A' 1417 1345 8.51      
16 A' 1353 1285 1.87      
17 A' 1330 1263 0.10      
18 A' 1201 1140 1.42      
19 A' 1119 1062 8.56      
20 A' 1013 962 1.48      
21 A' 988 938 0.79      
22 A' 925 879 1.55      
23 A' 752 714 1.22      
24 A' 523 497 0.96      
25 A' 443 420 0.90      
26 A' 331 314 0.16      
27 A' 286 272 0.30      
28 A' 263 250 0.19      
29 A" 3162 3002 10.07      
30 A" 3155 2995 3.25      
31 A" 3146 2987 15.44      
32 A" 3082 2926 21.25      
33 A" 1523 1446 0.25      
34 A" 1511 1434 1.67      
35 A" 1500 1424 8.23      
36 A" 1400 1329 7.65      
37 A" 1356 1287 1.36      
38 A" 1150 1092 2.35      
39 A" 1072 1017 0.00      
40 A" 981 931 0.17      
41 A" 944 896 37.62      
42 A" 934 886 1.62      
43 A" 755 717 0.81      
44 A" 558 530 8.17      
45 A" 272 258 0.06      
46 A" 209 199 0.41      
47 A" 178 169 0.13      
48 A" 26 25 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 36624.4 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 34771.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.15030 0.08504 0.08401

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.744 -1.058 0.000
C2 -0.473 -0.656 0.000
C3 -0.126 0.804 0.000
C4 0.651 -1.643 0.000
C5 0.651 1.195 1.250
C6 0.651 1.195 -1.250
H7 -2.001 -2.105 0.000
H8 -2.553 -0.346 0.000
H9 -1.065 1.353 0.000
H10 0.270 -2.659 0.000
H11 1.285 -1.520 -0.874
H12 1.285 -1.520 0.874
H13 0.813 2.269 1.275
H14 0.116 0.907 2.150
H15 1.626 0.716 1.266
H16 0.813 2.269 -1.275
H17 0.116 0.907 -2.150
H18 1.626 0.716 -1.266

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33292.46672.46503.51763.51761.07751.07792.50532.57213.18583.18584.38533.45604.01324.38533.45604.0132
C21.33291.50081.49532.50052.50052.10522.10302.09542.13542.14452.14453.44032.72302.80913.44032.72302.8091
C32.46671.50082.56691.52281.52283.46042.68551.08863.48472.85522.85522.15712.16602.16332.15712.16602.1633
C42.46501.49532.56693.10083.10082.69153.45633.45311.08471.08701.08704.11743.37842.84904.11743.37842.8490
C53.51762.50051.52283.10082.50014.41373.76842.12934.06883.50462.81251.08651.08591.08662.74853.45392.7405
C63.51762.50051.52283.10082.50014.41373.76842.12934.06882.81253.50462.74853.45392.74051.08651.08591.0866
H71.07752.10523.46042.69154.41374.41371.84343.58242.33723.44993.44995.35444.26324.76575.35444.26324.7657
H81.07792.10302.68553.45633.76843.76841.84342.25843.64904.10724.10724.44863.64894.49374.44863.64894.4937
H92.50532.09541.08863.45312.12932.12933.58242.25844.22833.81343.81342.44772.49333.04212.44772.49333.0421
H102.57212.13543.48471.08474.06884.06882.33723.64904.22831.75831.75835.11844.16683.85065.11844.16683.8506
H113.18582.14452.85521.08703.50462.81253.44994.10723.81341.75831.74884.38104.05033.11353.83862.98072.2948
H123.18582.14452.85521.08702.81253.50463.44994.10723.81341.75831.74883.83862.98072.29484.38104.05033.1135
H134.38533.44032.15714.11741.08652.74855.35444.44862.44775.11844.38103.83861.76221.75322.54963.75083.0868
H143.45602.72302.16603.37841.08593.45394.26323.64892.49334.16684.05032.98071.76221.76083.75084.30003.7399
H154.01322.80912.16332.84901.08662.74054.76574.49373.04213.85063.11352.29481.75321.76083.08683.73992.5317
H164.38533.44032.15714.11742.74851.08655.35444.44862.44775.11843.83864.38102.54963.75083.08681.76221.7532
H173.45602.72302.16603.37843.45391.08594.26323.64892.49334.16682.98074.05033.75084.30003.73991.76221.7608
H184.01322.80912.16332.84902.74051.08664.76574.49373.04213.85062.29483.11353.08683.73992.53171.75321.7608

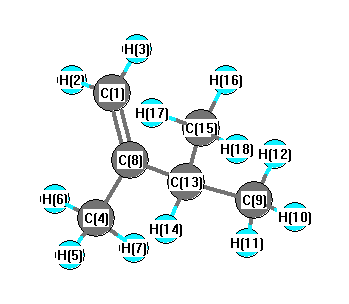
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.915 C1 C2 C4 121.176
C2 C1 H7 121.341 C2 C1 H8 121.093
C2 C3 C5 111.579 C2 C3 C6 111.579
C2 C3 H9 106.962 C2 C4 H10 110.716
C2 C4 H11 111.308 C2 C4 H12 111.308
C3 C2 C4 117.909 C3 C5 H13 110.402
C3 C5 H14 111.156 C3 C5 H15 110.890
C3 C6 H16 110.402 C3 C6 H17 111.156
C3 C6 H18 110.890 C5 C3 C6 110.344
C5 C3 H9 108.097 C6 C3 H9 108.097
H7 C1 H8 117.566 H10 C4 H11 108.122
H10 C4 H12 108.122 H11 C4 H12 107.107
H13 C5 H14 108.428 H13 C5 H15 107.559
H14 C5 H15 108.287 H16 C6 H17 108.428
H16 C6 H18 107.559 H17 C6 H18 108.287
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability