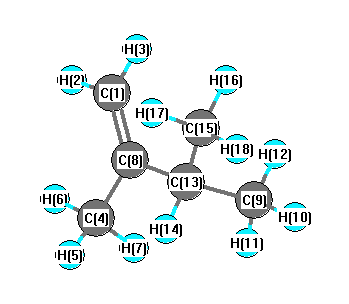
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3234 |
3095 |
19.60 |
|
|
|
| 2 |
A' |
3151 |
3015 |
11.97 |
|
|
|
| 3 |
A' |
3142 |
3007 |
19.82 |
|
|
|
| 4 |
A' |
3132 |
2998 |
27.55 |
|
|
|
| 5 |
A' |
3125 |
2991 |
70.46 |
|
|
|
| 6 |
A' |
3055 |
2924 |
37.34 |
|
|
|
| 7 |
A' |
3053 |
2922 |
0.85 |
|
|
|
| 8 |
A' |
3048 |
2918 |
17.80 |
|
|
|
| 9 |
A' |
1732 |
1658 |
22.23 |
|
|
|
| 10 |
A' |
1511 |
1446 |
9.61 |
|
|
|
| 11 |
A' |
1501 |
1437 |
6.36 |
|
|
|
| 12 |
A' |
1487 |
1423 |
12.75 |
|
|
|
| 13 |
A' |
1446 |
1384 |
1.06 |
|
|
|
| 14 |
A' |
1410 |
1350 |
4.20 |
|
|
|
| 15 |
A' |
1393 |
1333 |
10.57 |
|
|
|
| 16 |
A' |
1340 |
1282 |
2.81 |
|
|
|
| 17 |
A' |
1313 |
1257 |
0.05 |
|
|
|
| 18 |
A' |
1189 |
1138 |
1.74 |
|
|
|
| 19 |
A' |
1107 |
1059 |
10.11 |
|
|
|
| 20 |
A' |
995 |
953 |
1.80 |
|
|
|
| 21 |
A' |
962 |
920 |
0.70 |
|
|
|
| 22 |
A' |
912 |
873 |
1.82 |
|
|
|
| 23 |
A' |
740 |
708 |
1.31 |
|
|
|
| 24 |
A' |
512 |
490 |
1.14 |
|
|
|
| 25 |
A' |
424 |
406 |
0.94 |
|
|
|
| 26 |
A' |
321 |
307 |
0.18 |
|
|
|
| 27 |
A' |
254 |
243 |
0.51 |
|
|
|
| 28 |
A' |
232 |
222 |
0.16 |
|
|
|
| 29 |
A" |
3131 |
2997 |
17.47 |
|
|
|
| 30 |
A" |
3121 |
2987 |
0.22 |
|
|
|
| 31 |
A" |
3104 |
2971 |
20.33 |
|
|
|
| 32 |
A" |
3051 |
2920 |
25.43 |
|
|
|
| 33 |
A" |
1491 |
1427 |
0.34 |
|
|
|
| 34 |
A" |
1487 |
1423 |
0.87 |
|
|
|
| 35 |
A" |
1464 |
1401 |
8.54 |
|
|
|
| 36 |
A" |
1389 |
1330 |
6.75 |
|
|
|
| 37 |
A" |
1342 |
1284 |
1.90 |
|
|
|
| 38 |
A" |
1133 |
1085 |
2.29 |
|
|
|
| 39 |
A" |
1055 |
1009 |
0.22 |
|
|
|
| 40 |
A" |
967 |
925 |
0.16 |
|
|
|
| 41 |
A" |
935 |
895 |
39.87 |
|
|
|
| 42 |
A" |
922 |
882 |
0.91 |
|
|
|
| 43 |
A" |
749 |
717 |
0.82 |
|
|
|
| 44 |
A" |
551 |
527 |
8.38 |
|
|
|
| 45 |
A" |
235 |
225 |
0.15 |
|
|
|
| 46 |
A" |
208 |
199 |
0.43 |
|
|
|
| 47 |
A" |
117 |
112 |
0.00 |
|
|
|
| 48 |
A" |
24 |
23 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36095.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34547.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.354 |
|
|
|
| 2 |
C |
0.108 |
|
|
|
| 3 |
C |
-0.069 |
|
|
|
| 4 |
C |
-0.376 |
|
|
|
| 5 |
C |
-0.373 |
|
|
|
| 6 |
C |
-0.373 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
H |
0.111 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.124 |
|
|
|
| 13 |
H |
0.119 |
|
|
|
| 14 |
H |
0.127 |
|
|
|
| 15 |
H |
0.116 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
| 18 |
H |
0.116 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.272 |
0.405 |
0.000 |
0.488 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.994 |
-0.780 |
0.000 |
| y |
-0.780 |
-39.735 |
0.000 |
| z |
0.000 |
0.000 |
-41.546 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.646 |
-0.780 |
0.000 |
| y |
-0.780 |
0.535 |
0.000 |
| z |
0.000 |
0.000 |
-2.182 |
|
| Polar |
|---|
| 3z2-r2 | -4.363 |
|---|
| x2-y2 | 0.741 |
|---|
| xy | -0.780 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.775 |
0.776 |
0.000 |
| y |
0.776 |
12.176 |
0.000 |
| z |
0.000 |
0.000 |
9.232 |
<r2> (average value of r
2) Å
2
| <r2> |
188.615 |
| (<r2>)1/2 |
13.734 |