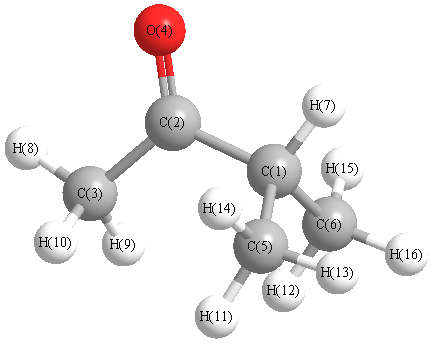
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3179 |
3043 |
8.88 |
|
|
|
| 2 |
A |
3140 |
3006 |
18.08 |
|
|
|
| 3 |
A |
3130 |
2996 |
33.84 |
|
|
|
| 4 |
A |
3129 |
2995 |
46.77 |
|
|
|
| 5 |
A |
3128 |
2993 |
1.76 |
|
|
|
| 6 |
A |
3122 |
2988 |
7.69 |
|
|
|
| 7 |
A |
3085 |
2952 |
3.10 |
|
|
|
| 8 |
A |
3062 |
2931 |
4.54 |
|
|
|
| 9 |
A |
3054 |
2923 |
15.32 |
|
|
|
| 10 |
A |
3052 |
2922 |
21.92 |
|
|
|
| 11 |
A |
1822 |
1744 |
185.84 |
|
|
|
| 12 |
A |
1514 |
1449 |
6.37 |
|
|
|
| 13 |
A |
1508 |
1443 |
21.39 |
|
|
|
| 14 |
A |
1498 |
1433 |
3.24 |
|
|
|
| 15 |
A |
1486 |
1423 |
1.34 |
|
|
|
| 16 |
A |
1471 |
1408 |
9.25 |
|
|
|
| 17 |
A |
1458 |
1395 |
12.79 |
|
|
|
| 18 |
A |
1418 |
1357 |
4.00 |
|
|
|
| 19 |
A |
1395 |
1336 |
4.89 |
|
|
|
| 20 |
A |
1381 |
1321 |
51.58 |
|
|
|
| 21 |
A |
1347 |
1289 |
1.33 |
|
|
|
| 22 |
A |
1327 |
1271 |
1.49 |
|
|
|
| 23 |
A |
1265 |
1211 |
42.56 |
|
|
|
| 24 |
A |
1187 |
1136 |
1.72 |
|
|
|
| 25 |
A |
1138 |
1090 |
0.82 |
|
|
|
| 26 |
A |
1092 |
1045 |
18.65 |
|
|
|
| 27 |
A |
1014 |
971 |
1.16 |
|
|
|
| 28 |
A |
969 |
928 |
1.16 |
|
|
|
| 29 |
A |
942 |
902 |
16.02 |
|
|
|
| 30 |
A |
931 |
891 |
1.91 |
|
|
|
| 31 |
A |
910 |
871 |
4.34 |
|
|
|
| 32 |
A |
732 |
701 |
5.13 |
|
|
|
| 33 |
A |
597 |
571 |
0.40 |
|
|
|
| 34 |
A |
526 |
503 |
7.71 |
|
|
|
| 35 |
A |
486 |
465 |
4.89 |
|
|
|
| 36 |
A |
324 |
310 |
1.19 |
|
|
|
| 37 |
A |
300 |
287 |
1.74 |
|
|
|
| 38 |
A |
260 |
249 |
0.62 |
|
|
|
| 39 |
A |
241 |
230 |
0.03 |
|
|
|
| 40 |
A |
202 |
193 |
0.63 |
|
|
|
| 41 |
A |
147 |
141 |
0.12 |
|
|
|
| 42 |
A |
51 |
49 |
3.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31009.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29679.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.278 |
|
|
|
| 2 |
C |
0.241 |
|
|
|
| 3 |
H |
0.130 |
|
|
|
| 4 |
H |
0.140 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
C |
-0.392 |
|
|
|
| 7 |
H |
0.129 |
|
|
|
| 8 |
C |
-0.102 |
|
|
|
| 9 |
H |
0.135 |
|
|
|
| 10 |
H |
0.135 |
|
|
|
| 11 |
H |
0.119 |
|
|
|
| 12 |
H |
0.117 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
H |
0.127 |
|
|
|
| 15 |
C |
-0.365 |
|
|
|
| 16 |
C |
-0.387 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.990 |
1.689 |
1.243 |
2.891 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.909 |
3.561 |
2.755 |
| y |
3.561 |
-38.866 |
-1.339 |
| z |
2.755 |
-1.339 |
-38.367 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.293 |
3.561 |
2.755 |
| y |
3.561 |
0.772 |
-1.339 |
| z |
2.755 |
-1.339 |
1.521 |
|
| Polar |
|---|
| 3z2-r2 | 3.041 |
|---|
| x2-y2 | -2.043 |
|---|
| xy | 3.561 |
|---|
| xz | 2.755 |
|---|
| yz | -1.339 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.918 |
-0.036 |
-0.004 |
| y |
-0.036 |
9.138 |
0.307 |
| z |
-0.004 |
0.307 |
8.400 |
<r2> (average value of r
2) Å
2
| <r2> |
177.402 |
| (<r2>)1/2 |
13.319 |