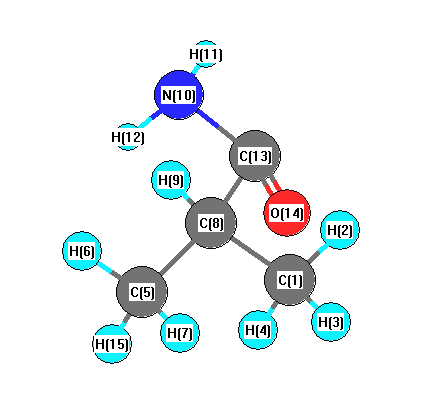
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3773 |
3611 |
36.84 |
|
|
|
| 2 |
A |
3630 |
3474 |
36.37 |
|
|
|
| 3 |
A |
3146 |
3011 |
28.88 |
|
|
|
| 4 |
A |
3143 |
3009 |
2.85 |
|
|
|
| 5 |
A |
3132 |
2997 |
37.84 |
|
|
|
| 6 |
A |
3130 |
2996 |
16.16 |
|
|
|
| 7 |
A |
3060 |
2929 |
14.96 |
|
|
|
| 8 |
A |
3058 |
2926 |
32.78 |
|
|
|
| 9 |
A |
3050 |
2919 |
32.94 |
|
|
|
| 10 |
A |
1805 |
1727 |
281.93 |
|
|
|
| 11 |
A |
1618 |
1549 |
127.25 |
|
|
|
| 12 |
A |
1515 |
1450 |
17.89 |
|
|
|
| 13 |
A |
1499 |
1435 |
10.62 |
|
|
|
| 14 |
A |
1491 |
1427 |
1.68 |
|
|
|
| 15 |
A |
1484 |
1420 |
1.06 |
|
|
|
| 16 |
A |
1425 |
1364 |
74.73 |
|
|
|
| 17 |
A |
1405 |
1345 |
5.63 |
|
|
|
| 18 |
A |
1391 |
1331 |
4.95 |
|
|
|
| 19 |
A |
1333 |
1276 |
0.87 |
|
|
|
| 20 |
A |
1286 |
1231 |
86.51 |
|
|
|
| 21 |
A |
1199 |
1147 |
2.81 |
|
|
|
| 22 |
A |
1135 |
1086 |
5.04 |
|
|
|
| 23 |
A |
1134 |
1085 |
0.17 |
|
|
|
| 24 |
A |
1061 |
1015 |
6.16 |
|
|
|
| 25 |
A |
982 |
940 |
0.22 |
|
|
|
| 26 |
A |
930 |
890 |
2.79 |
|
|
|
| 27 |
A |
929 |
889 |
1.90 |
|
|
|
| 28 |
A |
783 |
749 |
4.16 |
|
|
|
| 29 |
A |
766 |
733 |
3.50 |
|
|
|
| 30 |
A |
614 |
587 |
6.51 |
|
|
|
| 31 |
A |
600 |
574 |
10.10 |
|
|
|
| 32 |
A |
469 |
449 |
2.99 |
|
|
|
| 33 |
A |
318 |
304 |
0.83 |
|
|
|
| 34 |
A |
282 |
270 |
4.94 |
|
|
|
| 35 |
A |
244 |
233 |
5.41 |
|
|
|
| 36 |
A |
243 |
233 |
1.29 |
|
|
|
| 37 |
A |
216 |
207 |
16.58 |
|
|
|
| 38 |
A |
199 |
191 |
154.01 |
|
|
|
| 39 |
A |
28 |
26 |
7.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28750.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 27516.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.349 |
|
|
|
| 2 |
H |
0.115 |
|
|
|
| 3 |
H |
0.138 |
|
|
|
| 4 |
H |
0.113 |
|
|
|
| 5 |
C |
-0.349 |
|
|
|
| 6 |
H |
0.115 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
C |
-0.138 |
|
|
|
| 9 |
H |
0.115 |
|
|
|
| 10 |
N |
-0.254 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
H |
0.176 |
|
|
|
| 13 |
C |
0.246 |
|
|
|
| 14 |
O |
-0.352 |
|
|
|
| 15 |
H |
0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.280 |
-0.002 |
-3.616 |
3.627 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.655 |
0.002 |
4.665 |
| y |
0.002 |
-38.151 |
0.000 |
| z |
4.665 |
0.000 |
-39.053 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.947 |
0.002 |
4.665 |
| y |
0.002 |
-2.297 |
0.000 |
| z |
4.665 |
0.000 |
-3.650 |
|
| Polar |
|---|
| 3z2-r2 | -7.300 |
|---|
| x2-y2 | 5.496 |
|---|
| xy | 0.002 |
|---|
| xz | 4.665 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.551 |
0.000 |
0.354 |
| y |
0.000 |
7.774 |
0.000 |
| z |
0.354 |
0.000 |
8.475 |
<r2> (average value of r
2) Å
2
| <r2> |
170.569 |
| (<r2>)1/2 |
13.060 |