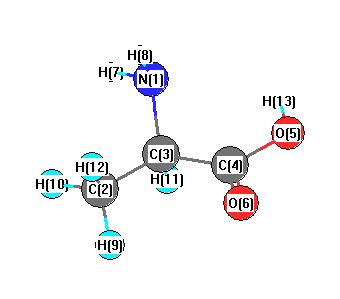
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3638 |
3490 |
10.84 |
|
|
|
| 2 |
A |
3549 |
3404 |
0.32 |
|
|
|
| 3 |
A |
3479 |
3337 |
293.18 |
|
|
|
| 4 |
A |
3160 |
3031 |
5.22 |
|
|
|
| 5 |
A |
3126 |
2998 |
23.62 |
|
|
|
| 6 |
A |
3065 |
2940 |
13.76 |
|
|
|
| 7 |
A |
3054 |
2929 |
10.71 |
|
|
|
| 8 |
A |
1878 |
1801 |
317.79 |
|
|
|
| 9 |
A |
1662 |
1594 |
34.01 |
|
|
|
| 10 |
A |
1500 |
1439 |
16.01 |
|
|
|
| 11 |
A |
1490 |
1429 |
3.23 |
|
|
|
| 12 |
A |
1444 |
1385 |
409.46 |
|
|
|
| 13 |
A |
1408 |
1351 |
10.02 |
|
|
|
| 14 |
A |
1394 |
1337 |
3.86 |
|
|
|
| 15 |
A |
1318 |
1264 |
8.12 |
|
|
|
| 16 |
A |
1251 |
1200 |
16.24 |
|
|
|
| 17 |
A |
1230 |
1179 |
8.45 |
|
|
|
| 18 |
A |
1163 |
1116 |
9.66 |
|
|
|
| 19 |
A |
1100 |
1055 |
39.77 |
|
|
|
| 20 |
A |
1022 |
980 |
0.63 |
|
|
|
| 21 |
A |
944 |
906 |
29.52 |
|
|
|
| 22 |
A |
906 |
869 |
63.92 |
|
|
|
| 23 |
A |
865 |
830 |
110.20 |
|
|
|
| 24 |
A |
816 |
783 |
6.16 |
|
|
|
| 25 |
A |
748 |
718 |
7.33 |
|
|
|
| 26 |
A |
576 |
553 |
3.36 |
|
|
|
| 27 |
A |
536 |
515 |
1.24 |
|
|
|
| 28 |
A |
406 |
389 |
4.57 |
|
|
|
| 29 |
A |
360 |
345 |
13.29 |
|
|
|
| 30 |
A |
287 |
276 |
13.74 |
|
|
|
| 31 |
A |
276 |
265 |
14.73 |
|
|
|
| 32 |
A |
229 |
219 |
0.27 |
|
|
|
| 33 |
A |
67 |
65 |
0.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23972.4 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 22994.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.309 |
|
|
|
| 2 |
C |
-0.249 |
|
|
|
| 3 |
C |
-0.059 |
|
|
|
| 4 |
C |
0.294 |
|
|
|
| 5 |
O |
-0.232 |
|
|
|
| 6 |
O |
-0.307 |
|
|
|
| 7 |
H |
0.135 |
|
|
|
| 8 |
H |
0.138 |
|
|
|
| 9 |
H |
0.111 |
|
|
|
| 10 |
H |
0.083 |
|
|
|
| 11 |
H |
0.102 |
|
|
|
| 12 |
H |
0.085 |
|
|
|
| 13 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.932 |
2.090 |
0.316 |
5.366 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.035 |
-0.235 |
-0.168 |
| y |
-0.235 |
-37.228 |
0.184 |
| z |
-0.168 |
0.184 |
-33.460 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.691 |
-0.235 |
-0.168 |
| y |
-0.235 |
-0.980 |
0.184 |
| z |
-0.168 |
0.184 |
4.671 |
|
| Polar |
|---|
| 3z2-r2 | 9.343 |
|---|
| x2-y2 | -1.807 |
|---|
| xy | -0.235 |
|---|
| xz | -0.168 |
|---|
| yz | 0.184 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.846 |
0.178 |
0.063 |
| y |
0.178 |
8.117 |
-0.074 |
| z |
0.063 |
-0.074 |
5.926 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |