return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-398.400769
Energy at 298.15K-398.410852
HF Energy-396.878996
Nuclear repulsion energy328.083879
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3835 3641 64.85      
2 A 3777 3586 90.66      
3 A 3641 3456 8.59      
4 A 3544 3364 3.30      
5 A 3172 3012 14.07      
6 A 3124 2966 9.14      
7 A 3059 2905 38.07      
8 A 1809 1718 219.27      
9 A 1662 1578 32.77      
10 A 1524 1447 4.97      
11 A 1441 1368 23.28      
12 A 1428 1356 33.99      
13 A 1408 1336 50.29      
14 A 1391 1320 4.64      
15 A 1329 1262 12.86      
16 A 1260 1196 12.33      
17 A 1203 1142 1.20      
18 A 1171 1112 193.97      
19 A 1157 1098 43.77      
20 A 1119 1062 94.62      
21 A 1051 998 48.66      
22 A 1011 959 2.05      
23 A 886 841 155.07      
24 A 829 787 32.60      
25 A 761 722 19.22      
26 A 633 601 86.93      
27 A 601 570 16.85      
28 A 534 507 166.91      
29 A 512 486 23.86      
30 A 460 437 8.84      
31 A 294 279 6.18      
32 A 288 274 27.51      
33 A 272 258 11.24      
34 A 211 201 2.03      
35 A 165 157 4.65      
36 A 40 38 1.23      

Unscaled Zero Point Vibrational Energy (zpe) 25300.0 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 24019.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.11876 0.07912 0.05371

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.769 -0.535 0.014
O2 2.038 -0.349 -0.384
O3 0.388 -1.535 0.574
C4 -0.085 0.670 -0.310
C5 -1.493 0.437 0.189
O6 -2.084 -0.692 -0.397
N7 0.429 1.912 0.231
H8 2.518 -1.151 -0.143
H9 -0.119 0.753 -1.393
H10 -1.469 0.369 1.278
H11 -2.086 1.302 -0.081
H12 -1.609 -1.450 -0.048
H13 1.332 2.126 -0.161
H14 0.542 1.838 1.232

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.34181.20831.51172.46832.88682.47951.86092.10412.72543.39612.54922.72512.6767
O21.34182.24572.35543.66194.13552.84230.96552.62443.94664.45203.82382.58373.1032
O31.20832.24572.42232.75232.78643.46392.27983.05982.75183.82052.09353.85153.4399
C41.51172.35542.42231.51222.42051.44873.18111.08672.12862.11112.62452.03702.0333
C52.46833.66192.75231.51221.40272.42254.32672.11891.09131.08271.90593.30972.6813
O62.88684.13552.78642.42051.40273.67234.63162.63392.07612.01840.96054.43443.9934
N72.47952.84233.46391.44872.42253.67233.72682.06962.66022.60633.94121.00781.0092
H81.86090.96552.27983.18114.32674.63163.72683.48524.49805.21714.13883.48573.8382
H92.10412.62443.05981.08672.11892.63392.06963.48523.01752.42692.98022.34772.9164
H102.72543.94662.75182.12861.09132.07612.66024.49803.01751.75962.25613.60602.4908
H113.39614.45203.82052.11111.08272.01842.60635.21712.42691.75962.79323.51682.9860
H122.54923.82382.09352.62451.90590.96053.94124.13882.98022.25612.79324.63174.1325
H132.72512.58373.85152.03703.30974.43441.00783.48572.34773.60603.51684.63171.6266
H142.67673.10323.43992.03332.68133.99341.00923.83822.91642.49082.98604.13251.6266

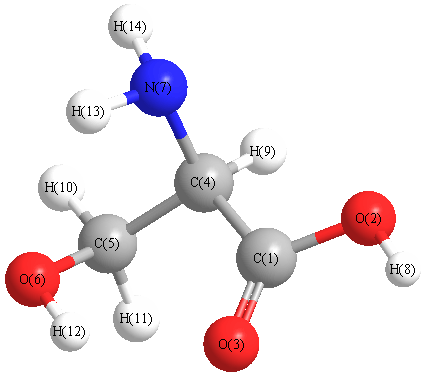
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 106.371 C1 C4 C5 109.423
C1 C4 N7 113.747 C1 C4 H9 107.008
O2 C1 O3 123.354 O2 C1 C4 111.125
O3 C1 C4 125.511 C4 C5 O6 112.225
C4 C5 H10 108.610 C4 C5 H11 107.744
C4 N7 H13 110.754 C4 N7 H14 110.351
C5 C4 N7 109.780 C5 C4 H9 108.115
C5 O6 H12 105.994 O6 C5 H10 112.096
O6 C5 H11 107.918 N7 C4 H9 108.586
H10 C5 H11 108.074 H13 N7 H14 107.496
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability