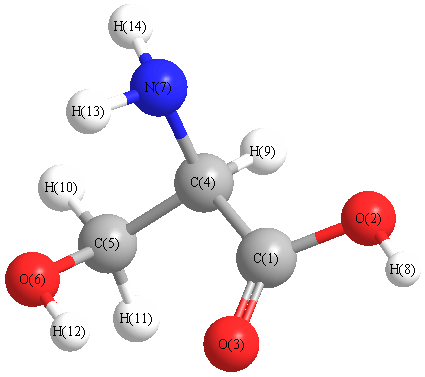
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3820 |
3664 |
67.14 |
|
|
|
| 2 |
A |
3797 |
3642 |
83.27 |
|
|
|
| 3 |
A |
3628 |
3480 |
5.90 |
|
|
|
| 4 |
A |
3540 |
3395 |
1.95 |
|
|
|
| 5 |
A |
3125 |
2998 |
18.61 |
|
|
|
| 6 |
A |
3064 |
2939 |
13.74 |
|
|
|
| 7 |
A |
3001 |
2879 |
48.33 |
|
|
|
| 8 |
A |
1822 |
1748 |
274.56 |
|
|
|
| 9 |
A |
1665 |
1597 |
32.92 |
|
|
|
| 10 |
A |
1503 |
1442 |
4.23 |
|
|
|
| 11 |
A |
1438 |
1379 |
33.74 |
|
|
|
| 12 |
A |
1405 |
1348 |
18.67 |
|
|
|
| 13 |
A |
1396 |
1339 |
59.80 |
|
|
|
| 14 |
A |
1384 |
1327 |
7.36 |
|
|
|
| 15 |
A |
1323 |
1269 |
9.58 |
|
|
|
| 16 |
A |
1248 |
1197 |
14.68 |
|
|
|
| 17 |
A |
1200 |
1152 |
11.05 |
|
|
|
| 18 |
A |
1175 |
1128 |
202.13 |
|
|
|
| 19 |
A |
1145 |
1098 |
27.34 |
|
|
|
| 20 |
A |
1117 |
1072 |
118.36 |
|
|
|
| 21 |
A |
1036 |
994 |
32.31 |
|
|
|
| 22 |
A |
1002 |
961 |
1.56 |
|
|
|
| 23 |
A |
870 |
834 |
154.92 |
|
|
|
| 24 |
A |
823 |
789 |
41.01 |
|
|
|
| 25 |
A |
753 |
722 |
22.78 |
|
|
|
| 26 |
A |
629 |
603 |
88.41 |
|
|
|
| 27 |
A |
586 |
562 |
10.97 |
|
|
|
| 28 |
A |
543 |
521 |
187.68 |
|
|
|
| 29 |
A |
515 |
494 |
7.59 |
|
|
|
| 30 |
A |
447 |
429 |
8.88 |
|
|
|
| 31 |
A |
296 |
284 |
4.82 |
|
|
|
| 32 |
A |
286 |
274 |
9.46 |
|
|
|
| 33 |
A |
256 |
246 |
27.38 |
|
|
|
| 34 |
A |
208 |
199 |
5.05 |
|
|
|
| 35 |
A |
153 |
146 |
4.24 |
|
|
|
| 36 |
A |
29 |
28 |
1.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25114.2 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 24089.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.293 |
|
|
|
| 2 |
O |
-0.248 |
|
|
|
| 3 |
O |
-0.331 |
|
|
|
| 4 |
C |
-0.038 |
|
|
|
| 5 |
C |
0.050 |
|
|
|
| 6 |
O |
-0.351 |
|
|
|
| 7 |
N |
-0.301 |
|
|
|
| 8 |
H |
0.212 |
|
|
|
| 9 |
H |
0.107 |
|
|
|
| 10 |
H |
0.048 |
|
|
|
| 11 |
H |
0.091 |
|
|
|
| 12 |
H |
0.211 |
|
|
|
| 13 |
H |
0.134 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.085 |
0.601 |
0.570 |
3.194 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.884 |
-2.014 |
-1.238 |
| y |
-2.014 |
-43.578 |
1.753 |
| z |
-1.238 |
1.753 |
-40.187 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.998 |
-2.014 |
-1.238 |
| y |
-2.014 |
-3.043 |
1.753 |
| z |
-1.238 |
1.753 |
2.044 |
|
| Polar |
|---|
| 3z2-r2 | 4.089 |
|---|
| x2-y2 | 2.694 |
|---|
| xy | -2.014 |
|---|
| xz | -1.238 |
|---|
| yz | 1.753 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.638 |
0.054 |
-0.224 |
| y |
0.054 |
8.435 |
-0.159 |
| z |
-0.224 |
-0.159 |
6.531 |
<r2> (average value of r
2) Å
2
| <r2> |
213.611 |
| (<r2>)1/2 |
14.615 |