return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6O (2,5-Cyclohexadienone)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-306.925139
Energy at 298.15K-306.931834
HF Energy-305.646162
Nuclear repulsion energy270.422450
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3251 3086 0.02      
2 A1 3205 3043 3.56      
3 A1 3059 2904 4.90      
4 A1 1740 1652 156.87      
5 A1 1689 1603 21.80      
6 A1 1447 1374 17.89      
7 A1 1429 1357 5.02      
8 A1 1204 1143 5.77      
9 A1 974 924 4.69      
10 A1 908 862 10.61      
11 A1 790 750 2.08      
12 A1 504 479 1.76      
13 A2 1221 1159 0.00      
14 A2 1031 979 0.00      
15 A2 774 735 0.00      
16 A2 365 347 0.00      
17 B1 3076 2921 2.62      
18 B1 1044 991 0.56      
19 B1 966 917 29.73      
20 B1 873 829 27.69      
21 B1 585 555 22.61      
22 B1 324 307 2.73      
23 B1 125 118 0.89      
24 B2 3249 3085 9.93      
25 B2 3204 3042 8.70      
26 B2 1673 1588 4.39      
27 B2 1448 1375 40.60      
28 B2 1398 1327 0.38      
29 B2 1292 1227 20.97      
30 B2 1156 1098 6.07      
31 B2 1017 965 5.64      
32 B2 579 549 0.78      
33 B2 453 430 13.42      

Unscaled Zero Point Vibrational Energy (zpe) 23025.0 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 21860.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.17780 0.08985 0.06033

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.103
C2 0.000 0.000 -1.812
C3 0.000 1.244 0.330
C4 0.000 -1.244 0.330
C5 0.000 1.245 -1.007
C6 0.000 -1.245 -1.007
O7 0.000 0.000 2.329
H8 0.000 2.159 0.896
H9 0.000 -2.159 0.896
H10 0.000 2.180 -1.545
H11 0.000 -2.180 -1.545
H12 0.863 0.000 -2.480
H13 -0.863 0.000 -2.480

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13
C12.91491.46451.46452.45042.45041.22542.16922.16923.43003.43003.68623.6862
C22.91492.47682.47681.48231.48234.14033.46303.46302.19632.19631.09211.0921
C31.46452.47682.48771.33742.82542.35401.07613.44982.09573.90363.19243.1924
C41.46452.47682.48772.82541.33742.35403.44981.07613.90362.09573.19243.1924
C52.45041.48231.33742.82542.49013.56062.11123.90011.07853.46702.11322.1132
C62.45041.48232.82541.33742.49013.56063.90012.11123.46701.07852.11322.1132
O71.22544.14032.35402.35403.56063.56062.59142.59144.44484.44484.88594.8859
H82.16923.46301.07613.44982.11123.90012.59144.31852.44064.97854.09954.0995
H92.16923.46303.44981.07613.90012.11122.59144.31854.97852.44064.09954.0995
H103.43002.19632.09573.90361.07853.46704.44482.44064.97854.36012.52462.5246
H113.43002.19633.90362.09573.46701.07854.44484.97852.44064.36012.52462.5246
H123.68621.09213.19243.19242.11322.11324.88594.09954.09952.52462.52461.7268
H133.68621.09213.19243.19242.11322.11324.88594.09954.09952.52462.52461.7268

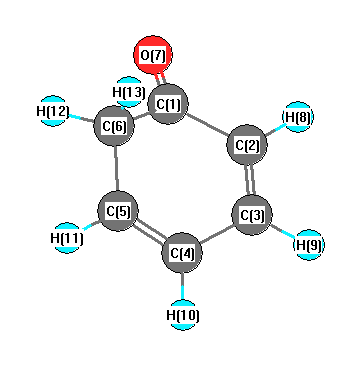
picture of 2,5-Cyclohexadienone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C5 121.915 C1 C3 H8 116.423
C1 C4 C6 121.915 C1 C4 H9 116.423
C2 C5 C3 122.816 C2 C5 H10 117.239
C2 C6 C4 122.816 C2 C6 H11 117.239
C3 C1 C4 116.271 C3 C1 O7 121.864
C3 C5 H10 119.945 C4 C1 O7 121.864
C4 C6 H11 119.945 C5 C2 C6 114.266
C5 C2 H12 109.408 C5 C2 H13 109.408
C5 C3 H8 121.662 C6 C2 H12 109.408
C6 C2 H13 109.408 C6 C4 H9 121.662
H12 C2 H13 104.487
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability