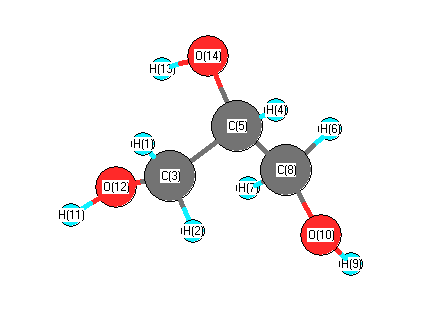
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3906 |
3738 |
50.55 |
|
|
|
| 2 |
A |
3899 |
3732 |
39.69 |
|
|
|
| 3 |
A |
3814 |
3651 |
53.90 |
|
|
|
| 4 |
A |
3092 |
2960 |
36.17 |
|
|
|
| 5 |
A |
3090 |
2958 |
43.11 |
|
|
|
| 6 |
A |
3068 |
2937 |
25.96 |
|
|
|
| 7 |
A |
3039 |
2908 |
32.27 |
|
|
|
| 8 |
A |
3028 |
2898 |
48.72 |
|
|
|
| 9 |
A |
1520 |
1455 |
7.41 |
|
|
|
| 10 |
A |
1506 |
1441 |
10.44 |
|
|
|
| 11 |
A |
1455 |
1393 |
2.21 |
|
|
|
| 12 |
A |
1446 |
1384 |
17.74 |
|
|
|
| 13 |
A |
1418 |
1357 |
38.09 |
|
|
|
| 14 |
A |
1368 |
1309 |
6.27 |
|
|
|
| 15 |
A |
1324 |
1267 |
43.52 |
|
|
|
| 16 |
A |
1284 |
1229 |
17.90 |
|
|
|
| 17 |
A |
1246 |
1193 |
10.01 |
|
|
|
| 18 |
A |
1218 |
1166 |
0.65 |
|
|
|
| 19 |
A |
1188 |
1137 |
30.31 |
|
|
|
| 20 |
A |
1149 |
1100 |
40.51 |
|
|
|
| 21 |
A |
1115 |
1067 |
89.26 |
|
|
|
| 22 |
A |
1099 |
1052 |
90.87 |
|
|
|
| 23 |
A |
1071 |
1025 |
77.33 |
|
|
|
| 24 |
A |
1008 |
964 |
16.18 |
|
|
|
| 25 |
A |
949 |
908 |
9.62 |
|
|
|
| 26 |
A |
840 |
804 |
12.80 |
|
|
|
| 27 |
A |
649 |
621 |
26.60 |
|
|
|
| 28 |
A |
484 |
463 |
6.15 |
|
|
|
| 29 |
A |
447 |
427 |
111.09 |
|
|
|
| 30 |
A |
395 |
378 |
1.79 |
|
|
|
| 31 |
A |
269 |
258 |
3.88 |
|
|
|
| 32 |
A |
249 |
238 |
5.79 |
|
|
|
| 33 |
A |
213 |
204 |
126.59 |
|
|
|
| 34 |
A |
207 |
198 |
55.66 |
|
|
|
| 35 |
A |
156 |
149 |
37.29 |
|
|
|
| 36 |
A |
96 |
92 |
3.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26151.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 25029.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.094 |
|
|
|
| 2 |
H |
0.094 |
|
|
|
| 3 |
C |
-0.056 |
|
|
|
| 4 |
H |
0.125 |
|
|
|
| 5 |
C |
0.039 |
|
|
|
| 6 |
H |
0.091 |
|
|
|
| 7 |
H |
0.085 |
|
|
|
| 8 |
C |
-0.047 |
|
|
|
| 9 |
H |
0.212 |
|
|
|
| 10 |
O |
-0.347 |
|
|
|
| 11 |
H |
0.218 |
|
|
|
| 12 |
O |
-0.361 |
|
|
|
| 13 |
H |
0.217 |
|
|
|
| 14 |
O |
-0.364 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.217 |
-2.058 |
-0.642 |
2.167 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.956 |
3.927 |
-3.061 |
| y |
3.927 |
-37.979 |
-0.220 |
| z |
-3.061 |
-0.220 |
-35.687 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.877 |
3.927 |
-3.061 |
| y |
3.927 |
-5.157 |
-0.220 |
| z |
-3.061 |
-0.220 |
-1.720 |
|
| Polar |
|---|
| 3z2-r2 | -3.439 |
|---|
| x2-y2 | 8.023 |
|---|
| xy | 3.927 |
|---|
| xz | -3.061 |
|---|
| yz | -0.220 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.766 |
0.131 |
-0.086 |
| y |
0.131 |
7.082 |
-0.178 |
| z |
-0.086 |
-0.178 |
6.812 |
<r2> (average value of r
2) Å
2
| <r2> |
171.670 |
| (<r2>)1/2 |
13.102 |