Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -550.799825 |
| Energy at 298.15K | |
| HF Energy | -549.143230 |
| Nuclear repulsion energy | 353.204429 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.019 |
0.000 |
| C2 |
1.019 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.019 |
0.000 |
| C4 |
-1.019 |
0.000 |
0.000 |
| F5 |
0.000 |
2.330 |
0.000 |
| F6 |
2.330 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.330 |
0.000 |
| F8 |
-2.330 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4405 | 2.0371 | 1.4405 | 1.3114 | 2.5429 | 3.3486 | 2.5429 |
C2 | 1.4405 | | 1.4405 | 2.0371 | 2.5429 | 1.3114 | 2.5429 | 3.3486 | C3 | 2.0371 | 1.4405 | | 1.4405 | 3.3486 | 2.5429 | 1.3114 | 2.5429 | C4 | 1.4405 | 2.0371 | 1.4405 | | 2.5429 | 3.3486 | 2.5429 | 1.3114 | F5 | 1.3114 | 2.5429 | 3.3486 | 2.5429 | | 3.2951 | 4.6600 | 3.2951 | F6 | 2.5429 | 1.3114 | 2.5429 | 3.3486 | 3.2951 | | 3.2951 | 4.6600 | F7 | 3.3486 | 2.5429 | 1.3114 | 2.5429 | 4.6600 | 3.2951 | | 3.2951 | F8 | 2.5429 | 3.3486 | 2.5429 | 1.3114 | 3.2951 | 4.6600 | 3.2951 | |
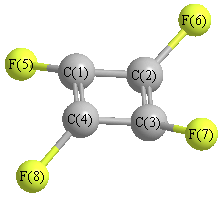 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -550.890902 |
| Energy at 298.15K | -550.890965 |
| HF Energy | -549.213436 |
| Nuclear repulsion energy | 354.709766 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1798 |
1753 |
|
|
|
|
| 2 |
Ag |
1202 |
1171 |
|
|
|
|
| 3 |
Ag |
729 |
710 |
|
|
|
|
| 4 |
Ag |
347 |
338 |
|
|
|
|
| 5 |
Ag |
199 |
194 |
|
|
|
|
| 6 |
Au |
1339 |
1305 |
|
|
|
|
| 7 |
Au |
931 |
908 |
|
|
|
|
| 8 |
Au |
593 |
578 |
|
|
|
|
| 9 |
Au |
247 |
241 |
|
|
|
|
| 10 |
Au |
150 |
146 |
|
|
|
|
| 11 |
Bg |
1358 |
1324 |
|
|
|
|
| 12 |
Bg |
764 |
745 |
|
|
|
|
| 13 |
Bg |
494 |
481 |
|
|
|
|
| 14 |
Bg |
409 |
399 |
|
|
|
|
| 15 |
Bu |
1758 |
1714 |
|
|
|
|
| 16 |
Bu |
992 |
967 |
|
|
|
|
| 17 |
Bu |
309 |
301 |
|
|
|
|
| 18 |
Bu |
170 |
166 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6893.7 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 6720.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.135 |
0.760 |
0.672 |
| C2 |
-0.135 |
-0.760 |
0.672 |
| C3 |
-0.135 |
-0.760 |
-0.672 |
| C4 |
0.135 |
0.760 |
-0.672 |
| F5 |
-0.135 |
1.669 |
1.586 |
| F6 |
0.135 |
-1.669 |
1.586 |
| F7 |
0.135 |
-1.669 |
-1.586 |
| F8 |
-0.135 |
1.669 |
-1.586 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5444 | 2.0480 | 1.3449 | 1.3172 | 2.5959 | 3.3175 | 2.4500 |
C2 | 1.5444 | | 1.3449 | 2.0480 | 2.5959 | 1.3172 | 2.4500 | 3.3175 | C3 | 2.0480 | 1.3449 | | 1.5444 | 3.3175 | 2.4500 | 1.3172 | 2.5959 | C4 | 1.3449 | 2.0480 | 1.5444 | | 2.4500 | 3.3175 | 2.5959 | 1.3172 | F5 | 1.3172 | 2.5959 | 3.3175 | 2.4500 | | 3.3497 | 4.6139 | 3.1730 | F6 | 2.5959 | 1.3172 | 2.4500 | 3.3175 | 3.3497 | | 3.1730 | 4.6139 | F7 | 3.3175 | 2.4500 | 1.3172 | 2.5959 | 4.6139 | 3.1730 | | 3.3497 | F8 | 2.4500 | 3.3175 | 2.5959 | 1.3172 | 3.1730 | 4.6139 | 3.3497 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
130.058 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
133.942 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
130.058 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
133.942 |
| C3 |
C2 |
F6 |
133.942 |
|
C3 |
C4 |
F8 |
130.058 |
| C4 |
C1 |
F5 |
133.942 |
|
C4 |
C3 |
F7 |
130.058 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
