Jump to
S1C2
S1C3
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -224.938246 |
| Energy at 298.15K | -224.944471 |
| HF Energy | -224.079400 |
| Nuclear repulsion energy | 123.602388 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3694 |
3504 |
|
|
|
|
| 2 |
A |
3581 |
3397 |
|
|
|
|
| 3 |
A |
1822 |
1728 |
|
|
|
|
| 4 |
A |
1633 |
1549 |
|
|
|
|
| 5 |
A |
1192 |
1131 |
|
|
|
|
| 6 |
A |
947 |
899 |
|
|
|
|
| 7 |
A |
631 |
599 |
|
|
|
|
| 8 |
A |
468 |
444 |
|
|
|
|
| 9 |
A |
388 |
368 |
|
|
|
|
| 10 |
B |
3693 |
3503 |
|
|
|
|
| 11 |
B |
3579 |
3395 |
|
|
|
|
| 12 |
B |
1640 |
1556 |
|
|
|
|
| 13 |
B |
1420 |
1347 |
|
|
|
|
| 14 |
B |
1068 |
1013 |
|
|
|
|
| 15 |
B |
791 |
750 |
|
|
|
|
| 16 |
B |
584 |
554 |
|
|
|
|
| 17 |
B |
553 |
524 |
|
|
|
|
| 18 |
B |
446 |
423 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14064.4 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 13341.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.148 |
| O2 |
0.000 |
0.000 |
1.363 |
| N3 |
0.000 |
1.161 |
-0.618 |
| N4 |
0.000 |
-1.161 |
-0.618 |
| H5 |
0.248 |
1.971 |
-0.072 |
| H6 |
0.491 |
1.103 |
-1.496 |
| H7 |
-0.248 |
-1.971 |
-0.072 |
| H8 |
-0.491 |
-1.103 |
-1.496 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2157 | 1.3909 | 1.3909 | 1.9981 | 2.0399 | 1.9981 | 2.0399 |
O2 | 1.2157 | | 2.2966 | 2.2966 | 2.4503 | 3.1043 | 2.4503 | 3.1043 | N3 | 1.3909 | 2.2966 | | 2.3220 | 1.0076 | 1.0078 | 3.1885 | 2.4779 | N4 | 1.3909 | 2.2966 | 2.3220 | | 3.1885 | 2.4779 | 1.0076 | 1.0078 | H5 | 1.9981 | 2.4503 | 1.0076 | 3.1885 | | 1.6853 | 3.9721 | 3.4676 | H6 | 2.0399 | 3.1043 | 1.0078 | 2.4779 | 1.6853 | | 3.4676 | 2.4158 | H7 | 1.9981 | 2.4503 | 3.1885 | 1.0076 | 3.9721 | 3.4676 | | 1.6853 | H8 | 2.0399 | 3.1043 | 2.4779 | 1.0078 | 3.4676 | 2.4158 | 1.6853 | |
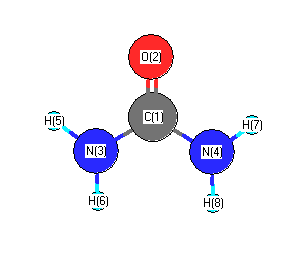 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
111.844 |
|
C1 |
N3 |
H6 |
115.598 |
| C1 |
N4 |
H7 |
111.844 |
|
C1 |
N4 |
H8 |
115.598 |
| O2 |
C1 |
N3 |
123.410 |
|
O2 |
C1 |
N4 |
123.410 |
| N3 |
C1 |
N4 |
113.179 |
|
H5 |
N3 |
H6 |
113.484 |
| H7 |
N4 |
H8 |
113.484 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -224.936234 |
| Energy at 298.15K | -224.942153 |
| HF Energy | -224.077885 |
| Nuclear repulsion energy | 123.637621 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3719 |
3528 |
|
|
|
|
| 2 |
A' |
3602 |
3417 |
|
|
|
|
| 3 |
A' |
1813 |
1720 |
|
|
|
|
| 4 |
A' |
1648 |
1563 |
|
|
|
|
| 5 |
A' |
1191 |
1130 |
|
|
|
|
| 6 |
A' |
955 |
906 |
|
|
|
|
| 7 |
A' |
776 |
736 |
|
|
|
|
| 8 |
A' |
544 |
516 |
|
|
|
|
| 9 |
A' |
500 |
474 |
|
|
|
|
| 10 |
A' |
453 |
430 |
|
|
|
|
| 11 |
A" |
3716 |
3525 |
|
|
|
|
| 12 |
A" |
3594 |
3410 |
|
|
|
|
| 13 |
A" |
1631 |
1547 |
|
|
|
|
| 14 |
A" |
1424 |
1351 |
|
|
|
|
| 15 |
A" |
1032 |
979 |
|
|
|
|
| 16 |
A" |
571 |
542 |
|
|
|
|
| 17 |
A" |
462 |
438 |
|
|
|
|
| 18 |
A" |
223 |
211 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13927.2 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 13211.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
0.143 |
0.000 |
| O2 |
0.045 |
1.360 |
0.000 |
| N3 |
0.045 |
-0.608 |
1.165 |
| N4 |
0.045 |
-0.608 |
-1.165 |
| H5 |
-0.196 |
-0.072 |
1.981 |
| H6 |
-0.334 |
-1.540 |
1.145 |
| H7 |
-0.196 |
-0.072 |
-1.981 |
| H8 |
-0.334 |
-1.540 |
-1.145 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2171 | 1.3868 | 1.3868 | 2.0038 | 2.0642 | 2.0038 | 2.0642 |
O2 | 1.2171 | | 2.2870 | 2.2870 | 2.4566 | 3.1402 | 2.4566 | 3.1402 | N3 | 1.3868 | 2.2870 | | 2.3299 | 1.0061 | 1.0058 | 3.2008 | 2.5191 | N4 | 1.3868 | 2.2870 | 2.3299 | | 3.2008 | 2.5191 | 1.0061 | 1.0058 | H5 | 2.0038 | 2.4566 | 1.0061 | 3.2008 | | 1.6949 | 3.9629 | 3.4562 | H6 | 2.0642 | 3.1402 | 1.0058 | 2.5191 | 1.6949 | | 3.4562 | 2.2894 | H7 | 2.0038 | 2.4566 | 3.2008 | 1.0061 | 3.9629 | 3.4562 | | 1.6949 | H8 | 2.0642 | 3.1402 | 2.5191 | 1.0058 | 3.4562 | 2.2894 | 1.6949 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
112.767 |
|
C1 |
N3 |
H6 |
118.390 |
| C1 |
N4 |
H7 |
112.767 |
|
C1 |
N4 |
H8 |
118.390 |
| O2 |
C1 |
N3 |
122.750 |
|
O2 |
C1 |
N4 |
122.750 |
| N3 |
C1 |
N4 |
114.289 |
|
H5 |
N3 |
H6 |
114.801 |
| H7 |
N4 |
H8 |
114.801 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -224.934524 |
| Energy at 298.15K | |
| HF Energy | -224.078164 |
| Nuclear repulsion energy | 123.925852 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3790 |
3595 |
|
|
|
|
| 2 |
A1 |
3655 |
3467 |
|
|
|
|
| 3 |
A1 |
1801 |
1709 |
|
|
|
|
| 4 |
A1 |
1627 |
1543 |
|
|
|
|
| 5 |
A1 |
1144 |
1085 |
|
|
|
|
| 6 |
A1 |
971 |
921 |
|
|
|
|
| 7 |
A1 |
473 |
448 |
|
|
|
|
| 8 |
A2 |
357 |
338 |
|
|
|
|
| 9 |
A2 |
469i |
445i |
|
|
|
|
| 10 |
B1 |
771 |
731 |
|
|
|
|
| 11 |
B1 |
565 |
536 |
|
|
|
|
| 12 |
B1 |
385i |
365i |
|
|
|
|
| 13 |
B2 |
3788 |
3593 |
|
|
|
|
| 14 |
B2 |
3647 |
3459 |
|
|
|
|
| 15 |
B2 |
1626 |
1542 |
|
|
|
|
| 16 |
B2 |
1433 |
1359 |
|
|
|
|
| 17 |
B2 |
990 |
939 |
|
|
|
|
| 18 |
B2 |
560 |
531 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13171.3 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 12494.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.144 |
| O2 |
0.000 |
0.000 |
1.364 |
| N3 |
0.000 |
1.156 |
-0.599 |
| N4 |
0.000 |
-1.156 |
-0.599 |
| H5 |
0.000 |
2.020 |
-0.094 |
| H6 |
0.000 |
1.169 |
-1.600 |
| H7 |
0.000 |
-2.020 |
-0.094 |
| H8 |
0.000 |
-1.169 |
-1.600 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2199 | 1.3737 | 1.3737 | 2.0341 | 2.0992 | 2.0341 | 2.0992 |
O2 | 1.2199 | | 2.2775 | 2.2775 | 2.4909 | 3.1856 | 2.4909 | 3.1856 | N3 | 1.3737 | 2.2775 | | 2.3115 | 1.0013 | 1.0009 | 3.2159 | 2.5313 | N4 | 1.3737 | 2.2775 | 2.3115 | | 3.2159 | 2.5313 | 1.0013 | 1.0009 | H5 | 2.0341 | 2.4909 | 1.0013 | 3.2159 | | 1.7299 | 4.0404 | 3.5272 | H6 | 2.0992 | 3.1856 | 1.0009 | 2.5313 | 1.7299 | | 3.5272 | 2.3386 | H7 | 2.0341 | 2.4909 | 3.2159 | 1.0013 | 4.0404 | 3.5272 | | 1.7299 | H8 | 2.0992 | 3.1856 | 2.5313 | 1.0009 | 3.5272 | 2.3386 | 1.7299 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.970 |
|
C1 |
N3 |
H6 |
123.498 |
| C1 |
N4 |
H7 |
116.970 |
|
C1 |
N4 |
H8 |
123.498 |
| O2 |
C1 |
N3 |
122.721 |
|
O2 |
C1 |
N4 |
122.721 |
| N3 |
C1 |
N4 |
114.557 |
|
H5 |
N3 |
H6 |
119.532 |
| H7 |
N4 |
H8 |
119.532 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
