Jump to
S1C2
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -225.301601 |
| Energy at 298.15K | -225.307787 |
| HF Energy | -225.301601 |
| Nuclear repulsion energy | 124.278294 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3732 |
3580 |
37.24 |
|
|
|
| 2 |
A |
3618 |
3471 |
3.21 |
|
|
|
| 3 |
A |
1832 |
1758 |
417.23 |
|
|
|
| 4 |
A |
1634 |
1568 |
0.28 |
|
|
|
| 5 |
A |
1186 |
1137 |
3.32 |
|
|
|
| 6 |
A |
969 |
929 |
8.60 |
|
|
|
| 7 |
A |
565 |
542 |
68.54 |
|
|
|
| 8 |
A |
474 |
455 |
2.45 |
|
|
|
| 9 |
A |
386 |
370 |
66.65 |
|
|
|
| 10 |
B |
3732 |
3580 |
30.26 |
|
|
|
| 11 |
B |
3614 |
3466 |
47.72 |
|
|
|
| 12 |
B |
1638 |
1571 |
199.86 |
|
|
|
| 13 |
B |
1433 |
1375 |
196.40 |
|
|
|
| 14 |
B |
1055 |
1012 |
21.59 |
|
|
|
| 15 |
B |
794 |
762 |
57.07 |
|
|
|
| 16 |
B |
583 |
560 |
74.85 |
|
|
|
| 17 |
B |
542 |
520 |
157.31 |
|
|
|
| 18 |
B |
447 |
429 |
156.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14117.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 13541.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.143 |
| O2 |
0.000 |
0.000 |
1.353 |
| N3 |
-0.123 |
1.149 |
-0.608 |
| N4 |
0.123 |
-1.149 |
-0.608 |
| H5 |
0.000 |
1.988 |
-0.069 |
| H6 |
0.306 |
1.166 |
-1.516 |
| H7 |
0.000 |
-1.988 |
-0.069 |
| H8 |
-0.306 |
-1.166 |
-1.516 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2107 | 1.3780 | 1.3780 | 1.9990 | 2.0504 | 1.9990 | 2.0504 |
O2 | 1.2107 | | 2.2765 | 2.2765 | 2.4443 | 3.1124 | 2.4443 | 3.1124 | N3 | 1.3780 | 2.2765 | | 2.3112 | 1.0044 | 1.0046 | 3.1851 | 2.4932 | N4 | 1.3780 | 2.2765 | 2.3112 | | 3.1851 | 2.4932 | 1.0044 | 1.0046 | H5 | 1.9990 | 2.4443 | 1.0044 | 3.1851 | | 1.6923 | 3.9754 | 3.4829 | H6 | 2.0504 | 3.1124 | 1.0046 | 2.4932 | 1.6923 | | 3.4829 | 2.4103 | H7 | 1.9990 | 2.4443 | 3.1851 | 1.0044 | 3.9754 | 3.4829 | | 1.6923 | H8 | 2.0504 | 3.1124 | 2.4932 | 1.0046 | 3.4829 | 2.4103 | 1.6923 | |
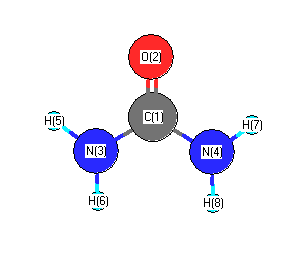 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.144 |
|
C1 |
N3 |
H6 |
117.910 |
| C1 |
N4 |
H7 |
113.144 |
|
C1 |
N4 |
H8 |
117.910 |
| O2 |
C1 |
N3 |
123.008 |
|
O2 |
C1 |
N4 |
123.008 |
| N3 |
C1 |
N4 |
113.985 |
|
H5 |
N3 |
H6 |
114.786 |
| H7 |
N4 |
H8 |
114.786 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.256 |
|
|
|
| 2 |
O |
-0.354 |
|
|
|
| 3 |
N |
-0.249 |
|
|
|
| 4 |
N |
-0.249 |
|
|
|
| 5 |
H |
0.158 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.744 |
3.744 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.241 |
2.285 |
0.000 |
| y |
2.285 |
-17.026 |
0.000 |
| z |
0.000 |
0.000 |
-24.830 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.313 |
2.285 |
0.000 |
| y |
2.285 |
8.010 |
0.000 |
| z |
0.000 |
0.000 |
-3.697 |
|
| Polar |
|---|
| 3z2-r2 | -7.394 |
|---|
| x2-y2 | -8.215 |
|---|
| xy | 2.285 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.008 |
-0.073 |
0.000 |
| y |
-0.073 |
5.221 |
0.000 |
| z |
0.000 |
0.000 |
5.279 |
<r2> (average value of r
2) Å
2
| <r2> |
68.030 |
| (<r2>)1/2 |
8.248 |
Jump to
S1C1
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -225.301614 |
| Energy at 298.15K | -225.307300 |
| HF Energy | -225.301614 |
| Nuclear repulsion energy | 124.326222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3762 |
3608 |
40.41 |
|
|
|
| 2 |
A' |
3641 |
3492 |
7.30 |
|
|
|
| 3 |
A' |
1823 |
1749 |
440.54 |
|
|
|
| 4 |
A' |
1641 |
1574 |
11.17 |
|
|
|
| 5 |
A' |
1179 |
1130 |
2.76 |
|
|
|
| 6 |
A' |
976 |
936 |
10.13 |
|
|
|
| 7 |
A' |
786 |
754 |
21.07 |
|
|
|
| 8 |
A' |
551 |
528 |
25.48 |
|
|
|
| 9 |
A' |
479 |
459 |
12.28 |
|
|
|
| 10 |
A' |
385 |
369 |
353.22 |
|
|
|
| 11 |
A" |
3759 |
3606 |
34.73 |
|
|
|
| 12 |
A" |
3632 |
3484 |
49.95 |
|
|
|
| 13 |
A" |
1630 |
1564 |
226.86 |
|
|
|
| 14 |
A" |
1436 |
1378 |
194.23 |
|
|
|
| 15 |
A" |
1020 |
978 |
18.45 |
|
|
|
| 16 |
A" |
575 |
552 |
22.19 |
|
|
|
| 17 |
A" |
414 |
397 |
51.94 |
|
|
|
| 18 |
A" |
143 |
137 |
41.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13914.8 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 13347.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.009 |
0.140 |
0.000 |
| O2 |
0.034 |
1.352 |
0.000 |
| N3 |
0.034 |
-0.599 |
1.158 |
| N4 |
0.034 |
-0.599 |
-1.158 |
| H5 |
-0.152 |
-0.073 |
1.991 |
| H6 |
-0.252 |
-1.560 |
1.161 |
| H7 |
-0.152 |
-0.073 |
-1.991 |
| H8 |
-0.252 |
-1.560 |
-1.161 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2121 | 1.3734 | 1.3734 | 2.0086 | 2.0748 | 2.0086 | 2.0748 |
O2 | 1.2121 | | 2.2682 | 2.2682 | 2.4552 | 3.1474 | 2.4552 | 3.1474 | N3 | 1.3734 | 2.2682 | | 2.3150 | 1.0027 | 1.0024 | 3.1973 | 2.5263 | N4 | 1.3734 | 2.2682 | 2.3150 | | 3.1973 | 2.5263 | 1.0027 | 1.0024 | H5 | 2.0086 | 2.4552 | 1.0027 | 3.1973 | | 1.7050 | 3.9815 | 3.4864 | H6 | 2.0748 | 3.1474 | 1.0024 | 2.5263 | 1.7050 | | 3.4864 | 2.3228 | H7 | 2.0086 | 2.4552 | 3.1973 | 1.0027 | 3.9815 | 3.4864 | | 1.7050 | H8 | 2.0748 | 3.1474 | 2.5263 | 1.0024 | 3.4864 | 2.3228 | 1.7050 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
114.512 |
|
C1 |
N3 |
H6 |
120.890 |
| C1 |
N4 |
H7 |
114.512 |
|
C1 |
N4 |
H8 |
120.890 |
| O2 |
C1 |
N3 |
122.506 |
|
O2 |
C1 |
N4 |
122.506 |
| N3 |
C1 |
N4 |
114.873 |
|
H5 |
N3 |
H6 |
116.493 |
| H7 |
N4 |
H8 |
116.493 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.254 |
|
|
|
| 2 |
O |
-0.362 |
|
|
|
| 3 |
N |
-0.238 |
|
|
|
| 4 |
N |
-0.238 |
|
|
|
| 5 |
H |
0.161 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.161 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.440 |
-4.056 |
0.000 |
4.305 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.217 |
1.510 |
0.000 |
| y |
1.510 |
-24.264 |
0.000 |
| z |
0.000 |
0.000 |
-17.049 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.561 |
1.510 |
0.000 |
| y |
1.510 |
-3.131 |
0.000 |
| z |
0.000 |
0.000 |
7.692 |
|
| Polar |
|---|
| 3z2-r2 | 15.384 |
|---|
| x2-y2 | -0.953 |
|---|
| xy | 1.510 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.958 |
0.101 |
0.000 |
| y |
0.101 |
5.276 |
0.000 |
| z |
0.000 |
0.000 |
5.222 |
<r2> (average value of r
2) Å
2
| <r2> |
68.063 |
| (<r2>)1/2 |
8.250 |
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -225.301076 |
| Energy at 298.15K | |
| HF Energy | -225.301076 |
| Nuclear repulsion energy | 124.458340 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3800 |
3645 |
53.36 |
|
|
|
| 2 |
A1 |
3668 |
3518 |
6.11 |
|
|
|
| 3 |
A1 |
1815 |
1741 |
472.16 |
|
|
|
| 4 |
A1 |
1628 |
1562 |
2.33 |
|
|
|
| 5 |
A1 |
1151 |
1104 |
0.45 |
|
|
|
| 6 |
A1 |
983 |
943 |
9.17 |
|
|
|
| 7 |
A1 |
477 |
458 |
3.38 |
|
|
|
| 8 |
A2 |
379 |
363 |
0.00 |
|
|
|
| 9 |
A2 |
388i |
373i |
0.00 |
|
|
|
| 10 |
B1 |
786 |
754 |
5.79 |
|
|
|
| 11 |
B1 |
571 |
548 |
8.08 |
|
|
|
| 12 |
B1 |
284i |
273i |
416.58 |
|
|
|
| 13 |
B2 |
3798 |
3643 |
41.92 |
|
|
|
| 14 |
B2 |
3659 |
3510 |
67.05 |
|
|
|
| 15 |
B2 |
1626 |
1560 |
288.54 |
|
|
|
| 16 |
B2 |
1441 |
1382 |
196.70 |
|
|
|
| 17 |
B2 |
993 |
952 |
12.78 |
|
|
|
| 18 |
B2 |
568 |
544 |
14.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13334.5 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 12790.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.140 |
| O2 |
0.000 |
0.000 |
1.354 |
| N3 |
0.000 |
1.153 |
-0.594 |
| N4 |
0.000 |
-1.153 |
-0.594 |
| H5 |
0.000 |
2.014 |
-0.085 |
| H6 |
0.000 |
1.177 |
-1.593 |
| H7 |
0.000 |
-2.014 |
-0.085 |
| H8 |
0.000 |
-1.177 |
-1.593 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2138 | 1.3666 | 1.3666 | 2.0262 | 2.0952 | 2.0262 | 2.0952 |
O2 | 1.2138 | | 2.2634 | 2.2634 | 2.4749 | 3.1736 | 2.4749 | 3.1736 | N3 | 1.3666 | 2.2634 | | 2.3054 | 1.0002 | 0.9998 | 3.2070 | 2.5347 | N4 | 1.3666 | 2.2634 | 2.3054 | | 3.2070 | 2.5347 | 1.0002 | 0.9998 | H5 | 2.0262 | 2.4749 | 1.0002 | 3.2070 | | 1.7252 | 4.0273 | 3.5290 | H6 | 2.0952 | 3.1736 | 0.9998 | 2.5347 | 1.7252 | | 3.5290 | 2.3533 | H7 | 2.0262 | 2.4749 | 3.2070 | 1.0002 | 4.0273 | 3.5290 | | 1.7252 | H8 | 2.0952 | 3.1736 | 2.5347 | 0.9998 | 3.5290 | 2.3533 | 1.7252 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.916 |
|
C1 |
N3 |
H6 |
123.862 |
| C1 |
N4 |
H7 |
116.916 |
|
C1 |
N4 |
H8 |
123.862 |
| O2 |
C1 |
N3 |
122.491 |
|
O2 |
C1 |
N4 |
122.491 |
| N3 |
C1 |
N4 |
115.019 |
|
H5 |
N3 |
H6 |
119.223 |
| H7 |
N4 |
H8 |
119.223 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.251 |
|
|
|
| 2 |
O |
-0.368 |
|
|
|
| 3 |
N |
-0.242 |
|
|
|
| 4 |
N |
-0.242 |
|
|
|
| 5 |
H |
0.164 |
|
|
|
| 6 |
H |
0.137 |
|
|
|
| 7 |
H |
0.164 |
|
|
|
| 8 |
H |
0.137 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.360 |
4.360 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.571 |
0.000 |
0.000 |
| y |
0.000 |
-16.365 |
0.000 |
| z |
0.000 |
0.000 |
-23.825 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.477 |
0.000 |
0.000 |
| y |
0.000 |
8.334 |
0.000 |
| z |
0.000 |
0.000 |
-2.857 |
|
| Polar |
|---|
| 3z2-r2 | -5.714 |
|---|
| x2-y2 | -9.207 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.898 |
0.000 |
0.000 |
| y |
0.000 |
5.195 |
0.000 |
| z |
0.000 |
0.000 |
5.243 |
<r2> (average value of r
2) Å
2
| <r2> |
67.987 |
| (<r2>)1/2 |
8.245 |
