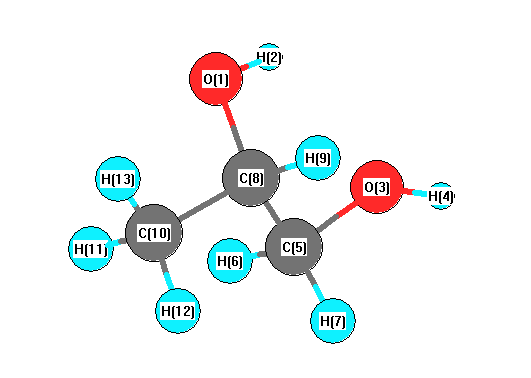
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3869 |
3703 |
31.42 |
|
|
|
| 2 |
A |
3809 |
3646 |
54.17 |
|
|
|
| 3 |
A |
3148 |
3013 |
18.00 |
|
|
|
| 4 |
A |
3138 |
3004 |
30.32 |
|
|
|
| 5 |
A |
3112 |
2978 |
22.98 |
|
|
|
| 6 |
A |
3063 |
2931 |
19.80 |
|
|
|
| 7 |
A |
3038 |
2908 |
42.44 |
|
|
|
| 8 |
A |
2950 |
2823 |
69.77 |
|
|
|
| 9 |
A |
1500 |
1436 |
1.82 |
|
|
|
| 10 |
A |
1493 |
1429 |
4.56 |
|
|
|
| 11 |
A |
1484 |
1421 |
5.26 |
|
|
|
| 12 |
A |
1445 |
1383 |
26.22 |
|
|
|
| 13 |
A |
1409 |
1348 |
39.69 |
|
|
|
| 14 |
A |
1391 |
1331 |
9.64 |
|
|
|
| 15 |
A |
1380 |
1321 |
4.12 |
|
|
|
| 16 |
A |
1350 |
1292 |
36.09 |
|
|
|
| 17 |
A |
1292 |
1236 |
47.39 |
|
|
|
| 18 |
A |
1226 |
1173 |
17.47 |
|
|
|
| 19 |
A |
1192 |
1140 |
29.40 |
|
|
|
| 20 |
A |
1104 |
1056 |
37.09 |
|
|
|
| 21 |
A |
1092 |
1045 |
11.19 |
|
|
|
| 22 |
A |
1064 |
1019 |
114.94 |
|
|
|
| 23 |
A |
944 |
904 |
3.88 |
|
|
|
| 24 |
A |
923 |
884 |
16.75 |
|
|
|
| 25 |
A |
859 |
822 |
12.27 |
|
|
|
| 26 |
A |
537 |
514 |
61.77 |
|
|
|
| 27 |
A |
497 |
476 |
87.51 |
|
|
|
| 28 |
A |
471 |
451 |
4.01 |
|
|
|
| 29 |
A |
355 |
340 |
16.92 |
|
|
|
| 30 |
A |
309 |
296 |
68.93 |
|
|
|
| 31 |
A |
243 |
233 |
2.56 |
|
|
|
| 32 |
A |
216 |
207 |
0.48 |
|
|
|
| 33 |
A |
152 |
146 |
4.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25026.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23952.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.351 |
|
|
|
| 2 |
H |
0.212 |
|
|
|
| 3 |
O |
-0.355 |
|
|
|
| 4 |
H |
0.218 |
|
|
|
| 5 |
C |
-0.053 |
|
|
|
| 6 |
H |
0.116 |
|
|
|
| 7 |
H |
0.085 |
|
|
|
| 8 |
C |
0.063 |
|
|
|
| 9 |
H |
0.071 |
|
|
|
| 10 |
C |
-0.354 |
|
|
|
| 11 |
H |
0.116 |
|
|
|
| 12 |
H |
0.104 |
|
|
|
| 13 |
H |
0.129 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.495 |
-1.641 |
1.474 |
2.260 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.753 |
1.956 |
2.741 |
| y |
1.956 |
-32.427 |
0.085 |
| z |
2.741 |
0.085 |
-29.743 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.668 |
1.956 |
2.741 |
| y |
1.956 |
-1.179 |
0.085 |
| z |
2.741 |
0.085 |
2.846 |
|
| Polar |
|---|
| 3z2-r2 | 5.693 |
|---|
| x2-y2 | -0.326 |
|---|
| xy | 1.956 |
|---|
| xz | 2.741 |
|---|
| yz | 0.085 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.461 |
0.219 |
0.140 |
| y |
0.219 |
6.630 |
-0.036 |
| z |
0.140 |
-0.036 |
6.289 |
<r2> (average value of r
2) Å
2
| <r2> |
130.926 |
| (<r2>)1/2 |
11.442 |