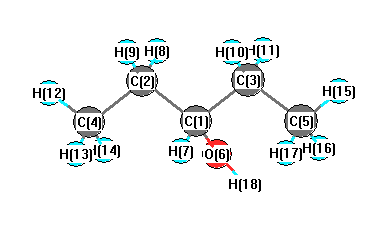
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3884 |
3718 |
23.87 |
|
|
|
| 2 |
A |
3147 |
3012 |
23.62 |
|
|
|
| 3 |
A |
3128 |
2994 |
29.18 |
|
|
|
| 4 |
A |
3124 |
2990 |
49.70 |
|
|
|
| 5 |
A |
3117 |
2983 |
50.42 |
|
|
|
| 6 |
A |
3094 |
2961 |
15.55 |
|
|
|
| 7 |
A |
3086 |
2954 |
0.72 |
|
|
|
| 8 |
A |
3056 |
2925 |
43.79 |
|
|
|
| 9 |
A |
3055 |
2924 |
38.61 |
|
|
|
| 10 |
A |
3049 |
2918 |
10.37 |
|
|
|
| 11 |
A |
3046 |
2916 |
17.42 |
|
|
|
| 12 |
A |
2970 |
2843 |
46.18 |
|
|
|
| 13 |
A |
1509 |
1444 |
7.40 |
|
|
|
| 14 |
A |
1507 |
1442 |
5.91 |
|
|
|
| 15 |
A |
1497 |
1432 |
7.77 |
|
|
|
| 16 |
A |
1495 |
1431 |
6.02 |
|
|
|
| 17 |
A |
1480 |
1416 |
3.54 |
|
|
|
| 18 |
A |
1469 |
1406 |
0.52 |
|
|
|
| 19 |
A |
1436 |
1374 |
9.92 |
|
|
|
| 20 |
A |
1410 |
1349 |
1.39 |
|
|
|
| 21 |
A |
1402 |
1342 |
5.18 |
|
|
|
| 22 |
A |
1394 |
1334 |
11.90 |
|
|
|
| 23 |
A |
1366 |
1308 |
0.20 |
|
|
|
| 24 |
A |
1326 |
1269 |
13.96 |
|
|
|
| 25 |
A |
1308 |
1252 |
1.71 |
|
|
|
| 26 |
A |
1282 |
1227 |
6.84 |
|
|
|
| 27 |
A |
1252 |
1198 |
28.31 |
|
|
|
| 28 |
A |
1181 |
1130 |
28.01 |
|
|
|
| 29 |
A |
1160 |
1110 |
3.81 |
|
|
|
| 30 |
A |
1099 |
1051 |
5.45 |
|
|
|
| 31 |
A |
1079 |
1032 |
26.36 |
|
|
|
| 32 |
A |
1042 |
997 |
2.92 |
|
|
|
| 33 |
A |
1028 |
984 |
1.59 |
|
|
|
| 34 |
A |
983 |
941 |
49.24 |
|
|
|
| 35 |
A |
942 |
902 |
9.49 |
|
|
|
| 36 |
A |
863 |
826 |
3.65 |
|
|
|
| 37 |
A |
780 |
747 |
1.50 |
|
|
|
| 38 |
A |
765 |
732 |
0.31 |
|
|
|
| 39 |
A |
491 |
470 |
3.50 |
|
|
|
| 40 |
A |
470 |
450 |
6.93 |
|
|
|
| 41 |
A |
393 |
377 |
0.95 |
|
|
|
| 42 |
A |
302 |
289 |
7.88 |
|
|
|
| 43 |
A |
248 |
237 |
47.00 |
|
|
|
| 44 |
A |
228 |
218 |
54.56 |
|
|
|
| 45 |
A |
212 |
203 |
6.52 |
|
|
|
| 46 |
A |
183 |
175 |
1.07 |
|
|
|
| 47 |
A |
108 |
103 |
1.55 |
|
|
|
| 48 |
A |
91 |
88 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36265.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34710.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.080 |
|
|
|
| 2 |
C |
-0.212 |
|
|
|
| 3 |
C |
-0.201 |
|
|
|
| 4 |
C |
-0.369 |
|
|
|
| 5 |
C |
-0.395 |
|
|
|
| 6 |
O |
-0.350 |
|
|
|
| 7 |
H |
0.086 |
|
|
|
| 8 |
H |
0.111 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
| 10 |
H |
0.117 |
|
|
|
| 11 |
H |
0.108 |
|
|
|
| 12 |
H |
0.114 |
|
|
|
| 13 |
H |
0.104 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
H |
0.106 |
|
|
|
| 17 |
H |
0.120 |
|
|
|
| 18 |
H |
0.208 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.124 |
-0.542 |
0.852 |
1.511 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.449 |
2.061 |
0.212 |
| y |
2.061 |
-39.267 |
1.266 |
| z |
0.212 |
1.266 |
-40.125 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.247 |
2.061 |
0.212 |
| y |
2.061 |
0.520 |
1.266 |
| z |
0.212 |
1.266 |
-0.767 |
|
| Polar |
|---|
| 3z2-r2 | -1.534 |
|---|
| x2-y2 | -0.182 |
|---|
| xy | 2.061 |
|---|
| xz | 0.212 |
|---|
| yz | 1.266 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.462 |
0.090 |
0.043 |
| y |
0.090 |
9.156 |
0.147 |
| z |
0.043 |
0.147 |
8.691 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |