return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (Z)-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-156.913392
Energy at 298.15K-156.921178
HF Energy-156.165329
Nuclear repulsion energy118.329046
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3217 3055 37.97      
2 A1 3189 3028 7.87      
3 A1 3090 2933 11.27      
4 A1 1745 1657 4.29      
5 A1 1523 1445 5.32      
6 A1 1430 1358 5.32      
7 A1 1307 1240 0.00      
8 A1 1038 985 0.68      
9 A1 891 846 0.54      
10 A1 299 283 0.00      
11 A2 3142 2983 0.00      
12 A2 1508 1431 0.00      
13 A2 1073 1018 0.00      
14 A2 1022 970 0.00      
15 A2 397 377 0.00      
16 A2 142 134 0.00      
17 B1 3143 2984 27.32      
18 B1 1514 1437 11.59      
19 B1 1081 1026 4.23      
20 B1 721 685 37.02      
21 B1 136 129 0.90      
22 B2 3187 3025 13.12      
23 B2 3171 3010 1.01      
24 B2 3087 2931 29.89      
25 B2 1514 1438 4.86      
26 B2 1456 1382 8.65      
27 B2 1398 1327 0.75      
28 B2 1170 1111 0.34      
29 B2 1005 954 13.53      
30 B2 565 537 5.77      

Unscaled Zero Point Vibrational Energy (zpe) 24079.2 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 22860.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.54757 0.17302 0.13810

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.669 0.647
C2 0.000 -0.669 0.647
C3 0.000 1.612 -0.511
C4 0.000 -1.612 -0.511
H5 0.000 1.151 1.612
H6 0.000 -1.151 1.612
H7 0.000 1.119 -1.471
H8 0.000 -1.119 -1.471
H9 -0.872 2.261 -0.478
H10 0.872 2.261 -0.478
H11 0.872 -2.261 -0.478
H12 -0.872 -2.261 -0.478

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.33811.49332.55831.07862.06022.16542.77182.13592.13593.25793.2579
C21.33812.55831.49332.06021.07862.77182.16543.25793.25792.13592.1359
C31.49332.55833.22452.17213.48461.07982.89491.08771.08773.97073.9707
C42.55831.49333.22453.48462.17212.89491.07983.97073.97071.08771.0877
H51.07862.06022.17213.48462.30263.08323.82862.52222.52224.09584.0958
H62.06021.07863.48462.17212.30263.82863.08324.09584.09582.52222.5222
H72.16542.77181.07982.89493.08323.82862.23741.74701.74703.62923.6292
H82.77182.16542.89491.07983.82863.08322.23743.62923.62921.74701.7470
H92.13593.25791.08773.97072.52224.09581.74703.62921.74434.84744.5226
H102.13593.25791.08773.97072.52224.09581.74703.62921.74434.52264.8474
H113.25792.13593.97071.08774.09582.52223.62921.74704.84744.52261.7443
H123.25792.13593.97071.08774.09582.52223.62921.74704.52264.84741.7443

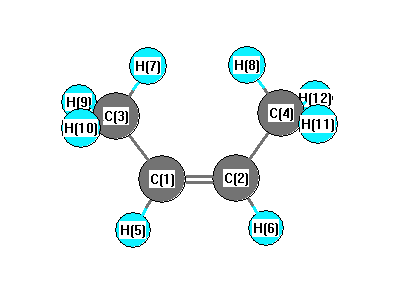
picture of 2-Butene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 129.167 C1 C2 H6 116.555
C1 C3 H7 113.634 C1 C3 H9 110.717
C1 C3 H10 110.717 C2 C1 C3 129.167
C2 C1 H5 116.555 C2 C4 H8 113.634
C2 C4 H11 110.717 C2 C4 H12 110.717
C3 C1 H5 114.277 C4 C2 H6 114.277
H7 C3 H9 107.417 H7 C3 H10 107.417
H8 C4 H11 107.417 H8 C4 H12 107.417
H9 C3 H10 106.617 H11 C4 H12 106.617
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability