return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,4-Pentadiene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-194.921861
Energy at 298.15K-194.929470
HF Energy-194.020030
Nuclear repulsion energy156.508503
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3279 3113 8.83      
2 A 3206 3044 1.12      
3 A 3192 3030 3.82      
4 A 3075 2920 16.51      
5 A 1715 1628 0.76      
6 A 1493 1418 6.70      
7 A 1466 1392 0.17      
8 A 1328 1261 0.15      
9 A 1280 1215 0.06      
10 A 1095 1040 3.67      
11 A 1043 990 13.46      
12 A 964 915 38.25      
13 A 933 886 2.60      
14 A 699 663 10.83      
15 A 373 354 0.46      
16 A 308 292 0.13      
17 A 88 84 0.02      
18 A 3279 3113 11.76      
19 A 3205 3043 5.06      
20 A 3192 3030 15.98      
21 A 3118 2960 10.14      
22 A 1704 1617 18.49      
23 A 1471 1397 4.59      
24 A 1336 1269 0.34      
25 A 1310 1244 1.79      
26 A 1189 1129 3.33      
27 A 1048 995 18.03      
28 A 992 942 2.38      
29 A 962 914 35.51      
30 A 905 859 6.94      
31 A 630 598 11.46      
32 A 456 433 2.69      
33 A 103 98 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 25218.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 23942.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.64532 0.08062 0.08004

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.686
C2 0.000 1.232 -0.160
C3 0.000 -1.232 -0.160
C4 0.992 2.116 -0.201
C5 -0.992 -2.116 -0.201
H6 -0.879 -0.014 1.328
H7 0.879 0.014 1.328
H8 -0.864 1.377 -0.791
H9 0.864 -1.377 -0.791
H10 0.949 2.979 -0.842
H11 -0.949 -2.979 -0.842
H12 1.869 1.992 0.414
H13 -1.869 -1.992 0.414

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.49411.49412.49942.49941.08891.08892.19592.19593.47963.47962.74532.7453
C21.49412.46321.32973.49172.13042.11441.07952.81942.10194.36952.09813.7704
C31.49412.46323.49171.32972.11442.13042.81941.07954.36952.10193.77042.0981
C42.49941.32973.49174.67393.22092.60172.08333.54461.07625.48941.07805.0439
C52.49943.49171.32974.67392.60173.22093.54462.08335.48941.07625.04391.0780
H61.08892.13042.11443.22092.60171.75822.53443.06364.12393.67523.52322.3938
H71.08892.11442.13042.60173.22091.75823.06362.53443.67524.12392.39383.5232
H82.19591.07952.81942.08333.54462.53443.06363.25142.41974.35693.04973.7163
H92.19592.81941.07953.54462.08333.06362.53443.25144.35692.41973.71633.0497
H103.47962.10194.36951.07625.48944.12393.67522.41974.35696.25241.84365.8505
H113.47964.36952.10195.48941.07623.67524.12394.35692.41976.25245.85051.8436
H122.74532.09813.77041.07805.04393.52322.39383.04973.71631.84365.85055.4635
H132.74533.77042.09815.04391.07802.39383.52323.71633.04975.85051.84365.4635

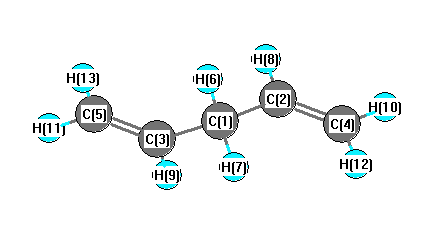
picture of 1,4-Pentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.435 C1 C2 H8 116.202
C1 C3 C5 124.435 C1 C3 H9 116.202
C2 C1 C3 111.042 C2 C1 H6 110.151
C2 C1 H7 108.880 C2 C4 H10 121.412
C2 C4 H12 120.888 C3 C1 H6 108.880
C3 C1 H7 110.151 C3 C5 H11 121.412
C3 C5 H13 120.888 C4 C2 H8 119.344
C5 C3 H9 119.344 H6 C1 H7 107.676
H10 C4 H12 117.700 H11 C5 H13 117.700
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/cc-pVTZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.135      
2 C -0.126      
3 C -0.126      
4 C -0.349      
5 C -0.349      
6 H 0.107      
7 H 0.107      
8 H 0.156      
9 H 0.156      
10 H 0.143      
11 H 0.143      
12 H 0.136      
13 H 0.136      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.673 2.589 0.000
y 2.589 10.409 0.000
z 0.000 0.000 6.574


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000