Jump to
S1C2
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -616.156932 |
| Energy at 298.15K | |
| HF Energy | -615.102876 |
| Nuclear repulsion energy | 206.416165 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
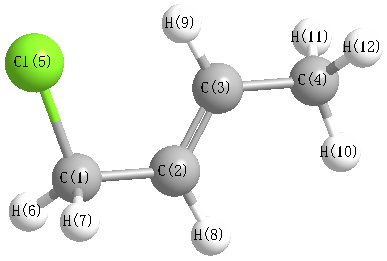
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.314 |
0.308 |
0.000 |
| C2 |
0.000 |
1.015 |
0.000 |
| C3 |
1.212 |
0.464 |
0.000 |
| C4 |
2.487 |
1.242 |
0.000 |
| Cl5 |
-1.188 |
-1.478 |
0.000 |
| H6 |
-1.895 |
0.573 |
0.879 |
| H7 |
-1.895 |
0.573 |
-0.879 |
| H8 |
-0.090 |
2.094 |
0.000 |
| H9 |
1.289 |
-0.612 |
0.000 |
| H10 |
2.297 |
2.314 |
0.000 |
| H11 |
3.089 |
1.001 |
0.876 |
| H12 |
3.089 |
1.001 |
-0.876 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4925 | 2.5310 | 3.9147 | 1.7896 | 1.0866 | 1.0866 | 2.1659 | 2.7611 | 4.1309 | 4.5429 | 4.5429 |
C2 | 1.4925 | | 1.3315 | 2.4976 | 2.7615 | 2.1348 | 2.1348 | 1.0828 | 2.0763 | 2.6385 | 3.2110 | 3.2110 | C3 | 2.5310 | 1.3315 | | 1.4940 | 3.0872 | 3.2305 | 3.2305 | 2.0863 | 1.0791 | 2.1442 | 2.1400 | 2.1400 | C4 | 3.9147 | 2.4976 | 1.4940 | | 4.5724 | 4.5189 | 4.5189 | 2.7143 | 2.2079 | 1.0879 | 1.0898 | 1.0898 | Cl5 | 1.7896 | 2.7615 | 3.0872 | 4.5724 | | 2.3408 | 2.3408 | 3.7369 | 2.6242 | 5.1497 | 5.0207 | 5.0207 | H6 | 1.0866 | 2.1348 | 3.2305 | 4.5189 | 2.3408 | | 1.7586 | 2.5184 | 3.5094 | 4.6227 | 5.0021 | 5.3011 | H7 | 1.0866 | 2.1348 | 3.2305 | 4.5189 | 2.3408 | 1.7586 | | 2.5184 | 3.5094 | 4.6227 | 5.3011 | 5.0021 | H8 | 2.1659 | 1.0828 | 2.0863 | 2.7143 | 3.7369 | 2.5184 | 2.5184 | | 3.0376 | 2.3969 | 3.4740 | 3.4740 | H9 | 2.7611 | 2.0763 | 1.0791 | 2.2079 | 2.6242 | 3.5094 | 3.5094 | 3.0376 | | 3.0944 | 2.5711 | 2.5711 | H10 | 4.1309 | 2.6385 | 2.1442 | 1.0879 | 5.1497 | 4.6227 | 4.6227 | 2.3969 | 3.0944 | | 1.7655 | 1.7655 | H11 | 4.5429 | 3.2110 | 2.1400 | 1.0898 | 5.0207 | 5.0021 | 5.3011 | 3.4740 | 2.5711 | 1.7655 | | 1.7519 | H12 | 4.5429 | 3.2110 | 2.1400 | 1.0898 | 5.0207 | 5.3011 | 5.0021 | 3.4740 | 2.5711 | 1.7655 | 1.7519 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.246 |
|
C1 |
C2 |
H8 |
113.541 |
| C2 |
C1 |
Cl5 |
114.264 |
|
C2 |
C1 |
H6 |
110.746 |
| C2 |
C1 |
H7 |
110.746 |
|
C2 |
C3 |
C4 |
124.149 |
| C2 |
C3 |
H9 |
118.559 |
|
C3 |
C2 |
H8 |
119.214 |
| C3 |
C4 |
H10 |
111.326 |
|
C3 |
C4 |
H11 |
110.865 |
| C3 |
C4 |
H12 |
110.865 |
|
C4 |
C3 |
H9 |
117.292 |
| Cl5 |
C1 |
H6 |
106.366 |
|
Cl5 |
C1 |
H7 |
106.366 |
| H6 |
C1 |
H7 |
108.037 |
|
H10 |
C4 |
H11 |
108.328 |
| H10 |
C4 |
H12 |
108.328 |
|
H11 |
C4 |
H12 |
106.976 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -616.149039 |
| Energy at 298.15K | |
| HF Energy | -615.093773 |
| Nuclear repulsion energy | 205.436923 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C1
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
