Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -616.858432 |
| Energy at 298.15K | -616.865567 |
| HF Energy | -616.858432 |
| Nuclear repulsion energy | 206.890058 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3186 |
3049 |
6.39 |
|
|
|
| 2 |
A' |
3143 |
3008 |
14.82 |
|
|
|
| 3 |
A' |
3132 |
2998 |
16.25 |
|
|
|
| 4 |
A' |
3084 |
2952 |
17.26 |
|
|
|
| 5 |
A' |
3047 |
2916 |
26.98 |
|
|
|
| 6 |
A' |
1774 |
1698 |
0.79 |
|
|
|
| 7 |
A' |
1491 |
1427 |
12.88 |
|
|
|
| 8 |
A' |
1473 |
1409 |
2.88 |
|
|
|
| 9 |
A' |
1412 |
1351 |
3.08 |
|
|
|
| 10 |
A' |
1327 |
1270 |
0.56 |
|
|
|
| 11 |
A' |
1324 |
1267 |
29.31 |
|
|
|
| 12 |
A' |
1294 |
1238 |
4.88 |
|
|
|
| 13 |
A' |
1132 |
1083 |
0.44 |
|
|
|
| 14 |
A' |
1055 |
1009 |
13.32 |
|
|
|
| 15 |
A' |
916 |
877 |
10.70 |
|
|
|
| 16 |
A' |
729 |
697 |
19.47 |
|
|
|
| 17 |
A' |
589 |
564 |
3.95 |
|
|
|
| 18 |
A' |
334 |
320 |
1.17 |
|
|
|
| 19 |
A' |
168 |
161 |
0.80 |
|
|
|
| 20 |
A" |
3127 |
2993 |
4.12 |
|
|
|
| 21 |
A" |
3101 |
2968 |
16.29 |
|
|
|
| 22 |
A" |
1478 |
1415 |
6.85 |
|
|
|
| 23 |
A" |
1200 |
1149 |
2.15 |
|
|
|
| 24 |
A" |
1072 |
1026 |
0.95 |
|
|
|
| 25 |
A" |
1000 |
957 |
34.64 |
|
|
|
| 26 |
A" |
953 |
912 |
0.01 |
|
|
|
| 27 |
A" |
740 |
708 |
0.40 |
|
|
|
| 28 |
A" |
246 |
236 |
4.23 |
|
|
|
| 29 |
A" |
212 |
202 |
2.23 |
|
|
|
| 30 |
A" |
127 |
122 |
1.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21931.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20990.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.307 |
0.301 |
0.000 |
| C2 |
0.000 |
1.011 |
0.000 |
| C3 |
1.207 |
0.473 |
0.000 |
| C4 |
2.477 |
1.249 |
0.000 |
| Cl5 |
-1.184 |
-1.479 |
0.000 |
| H6 |
-1.892 |
0.564 |
0.878 |
| H7 |
-1.892 |
0.564 |
-0.878 |
| H8 |
-0.102 |
2.092 |
0.000 |
| H9 |
1.291 |
-0.607 |
0.000 |
| H10 |
2.293 |
2.321 |
0.000 |
| H11 |
3.082 |
1.006 |
0.874 |
| H12 |
3.082 |
1.006 |
-0.874 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4874 | 2.5199 | 3.9007 | 1.7844 | 1.0875 | 1.0875 | 2.1586 | 2.7520 | 4.1285 | 4.5300 | 4.5300 |
C2 | 1.4874 | | 1.3216 | 2.4882 | 2.7573 | 2.1330 | 2.1330 | 1.0860 | 2.0696 | 2.6414 | 3.2032 | 3.2032 | C3 | 2.5199 | 1.3216 | | 1.4882 | 3.0867 | 3.2221 | 3.2221 | 2.0824 | 1.0827 | 2.1440 | 2.1360 | 2.1360 | C4 | 3.9007 | 2.4882 | 1.4882 | | 4.5656 | 4.5082 | 4.5082 | 2.7133 | 2.2021 | 1.0876 | 1.0904 | 1.0904 | Cl5 | 1.7844 | 2.7573 | 3.0867 | 4.5656 | | 2.3341 | 2.3341 | 3.7315 | 2.6244 | 5.1513 | 5.0136 | 5.0136 | H6 | 1.0875 | 2.1330 | 3.2221 | 4.5082 | 2.3341 | | 1.7567 | 2.5115 | 3.5032 | 4.6232 | 4.9929 | 5.2915 | H7 | 1.0875 | 2.1330 | 3.2221 | 4.5082 | 2.3341 | 1.7567 | | 2.5115 | 3.5032 | 4.6232 | 5.2915 | 4.9929 | H8 | 2.1586 | 1.0860 | 2.0824 | 2.7133 | 3.7315 | 2.5115 | 2.5115 | | 3.0371 | 2.4067 | 3.4757 | 3.4757 | H9 | 2.7520 | 2.0696 | 1.0827 | 2.2021 | 2.6244 | 3.5032 | 3.5032 | 3.0371 | | 3.0946 | 2.5633 | 2.5633 | H10 | 4.1285 | 2.6414 | 2.1440 | 1.0876 | 5.1513 | 4.6232 | 4.6232 | 2.4067 | 3.0946 | | 1.7648 | 1.7648 | H11 | 4.5300 | 3.2032 | 2.1360 | 1.0904 | 5.0136 | 4.9929 | 5.2915 | 3.4757 | 2.5633 | 1.7648 | | 1.7482 | H12 | 4.5300 | 3.2032 | 2.1360 | 1.0904 | 5.0136 | 5.2915 | 4.9929 | 3.4757 | 2.5633 | 1.7648 | 1.7482 | |
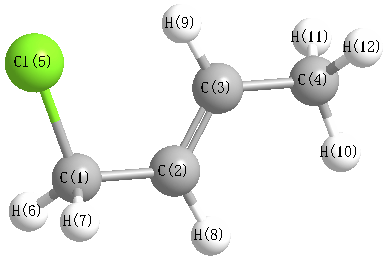 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.460 |
|
C1 |
C2 |
H8 |
113.109 |
| C2 |
C1 |
Cl5 |
114.561 |
|
C2 |
C1 |
H6 |
110.909 |
| C2 |
C1 |
H7 |
110.909 |
|
C2 |
C3 |
C4 |
124.533 |
| C2 |
C3 |
H9 |
118.478 |
|
C3 |
C2 |
H8 |
119.431 |
| C3 |
C4 |
H10 |
111.740 |
|
C3 |
C4 |
H11 |
110.916 |
| C3 |
C4 |
H12 |
110.916 |
|
C4 |
C3 |
H9 |
116.989 |
| Cl5 |
C1 |
H6 |
106.177 |
|
Cl5 |
C1 |
H7 |
106.177 |
| H6 |
C1 |
H7 |
107.731 |
|
H10 |
C4 |
H11 |
108.250 |
| H10 |
C4 |
H12 |
108.250 |
|
H11 |
C4 |
H12 |
106.574 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.102 |
|
|
|
| 2 |
C |
-0.202 |
|
|
|
| 3 |
C |
-0.102 |
|
|
|
| 4 |
C |
-0.331 |
|
|
|
| 5 |
Cl |
-0.168 |
|
|
|
| 6 |
H |
0.137 |
|
|
|
| 7 |
H |
0.137 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
| 10 |
H |
0.109 |
|
|
|
| 11 |
H |
0.120 |
|
|
|
| 12 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.302 |
1.888 |
0.000 |
1.912 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.499 |
-2.503 |
0.000 |
| y |
-2.503 |
-38.354 |
0.000 |
| z |
0.000 |
0.000 |
-39.184 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.270 |
-2.503 |
0.000 |
| y |
-2.503 |
-1.013 |
0.000 |
| z |
0.000 |
0.000 |
-2.257 |
|
| Polar |
|---|
| 3z2-r2 | -4.515 |
|---|
| x2-y2 | 2.855 |
|---|
| xy | -2.503 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.033 |
0.591 |
0.000 |
| y |
0.591 |
9.377 |
0.000 |
| z |
0.000 |
0.000 |
6.124 |
<r2> (average value of r
2) Å
2
| <r2> |
206.690 |
| (<r2>)1/2 |
14.377 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -616.859952 |
| Energy at 298.15K | -616.867017 |
| HF Energy | -616.859952 |
| Nuclear repulsion energy | 202.361833 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3041 |
16.53 |
|
|
|
| 2 |
A |
3161 |
3025 |
1.58 |
|
|
|
| 3 |
A |
3153 |
3017 |
25.78 |
|
|
|
| 4 |
A |
3134 |
3000 |
2.64 |
|
|
|
| 5 |
A |
3104 |
2971 |
14.13 |
|
|
|
| 6 |
A |
3100 |
2967 |
15.15 |
|
|
|
| 7 |
A |
3047 |
2916 |
22.91 |
|
|
|
| 8 |
A |
1757 |
1682 |
12.36 |
|
|
|
| 9 |
A |
1489 |
1425 |
15.37 |
|
|
|
| 10 |
A |
1480 |
1416 |
3.14 |
|
|
|
| 11 |
A |
1477 |
1414 |
7.92 |
|
|
|
| 12 |
A |
1411 |
1350 |
2.62 |
|
|
|
| 13 |
A |
1337 |
1280 |
3.12 |
|
|
|
| 14 |
A |
1332 |
1275 |
0.64 |
|
|
|
| 15 |
A |
1271 |
1217 |
35.83 |
|
|
|
| 16 |
A |
1205 |
1153 |
2.46 |
|
|
|
| 17 |
A |
1123 |
1075 |
0.61 |
|
|
|
| 18 |
A |
1110 |
1063 |
1.54 |
|
|
|
| 19 |
A |
1069 |
1023 |
1.03 |
|
|
|
| 20 |
A |
1005 |
962 |
35.77 |
|
|
|
| 21 |
A |
951 |
910 |
11.24 |
|
|
|
| 22 |
A |
896 |
858 |
2.79 |
|
|
|
| 23 |
A |
808 |
773 |
13.11 |
|
|
|
| 24 |
A |
701 |
671 |
56.00 |
|
|
|
| 25 |
A |
488 |
467 |
1.41 |
|
|
|
| 26 |
A |
355 |
339 |
3.85 |
|
|
|
| 27 |
A |
274 |
262 |
2.20 |
|
|
|
| 28 |
A |
215 |
206 |
0.95 |
|
|
|
| 29 |
A |
158 |
152 |
2.64 |
|
|
|
| 30 |
A |
90 |
87 |
0.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21938.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20997.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.769 |
0.822 |
0.114 |
| C2 |
0.515 |
0.169 |
0.449 |
| C3 |
1.590 |
0.240 |
-0.321 |
| C4 |
2.905 |
-0.382 |
-0.012 |
| Cl5 |
-2.086 |
-0.395 |
-0.105 |
| H6 |
-1.120 |
1.474 |
0.907 |
| H7 |
-0.708 |
1.381 |
-0.813 |
| H8 |
0.547 |
-0.385 |
1.380 |
| H9 |
1.522 |
0.791 |
-1.254 |
| H10 |
2.882 |
-0.914 |
0.936 |
| H11 |
3.194 |
-1.086 |
-0.793 |
| H12 |
3.693 |
0.370 |
0.037 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4786 | 2.4690 | 3.8685 | 1.8060 | 1.0856 | 1.0844 | 2.1883 | 2.6688 | 4.1250 | 4.4911 | 4.4855 |
C2 | 1.4786 | | 1.3250 | 2.4961 | 2.7178 | 2.1417 | 2.1344 | 1.0834 | 2.0739 | 2.6480 | 3.2089 | 3.2112 | C3 | 2.4690 | 1.3250 | | 1.4868 | 3.7368 | 3.2219 | 2.6127 | 2.0910 | 1.0853 | 2.1396 | 2.1335 | 2.1367 | C4 | 3.8685 | 2.4961 | 1.4868 | | 4.9917 | 4.5270 | 4.0991 | 2.7384 | 2.1976 | 1.0871 | 1.0904 | 1.0904 | Cl5 | 1.8060 | 2.7178 | 3.7368 | 4.9917 | | 2.3351 | 2.3567 | 3.0222 | 3.9677 | 5.1016 | 5.3692 | 5.8308 | H6 | 1.0856 | 2.1417 | 3.2219 | 4.5270 | 2.3351 | | 1.7714 | 2.5413 | 3.4812 | 4.6603 | 5.2971 | 5.0141 | H7 | 1.0844 | 2.1344 | 2.6127 | 4.0991 | 2.3567 | 1.7714 | | 3.0820 | 2.3485 | 4.6052 | 4.6164 | 4.5944 | H8 | 2.1883 | 1.0834 | 2.0910 | 2.7384 | 3.0222 | 2.5413 | 3.0820 | | 3.0443 | 2.4349 | 3.4959 | 3.5032 | H9 | 2.6688 | 2.0739 | 1.0853 | 2.1976 | 3.9677 | 3.4812 | 2.3485 | 3.0443 | | 3.0900 | 2.5552 | 2.5600 | H10 | 4.1250 | 2.6480 | 2.1396 | 1.0871 | 5.1016 | 4.6603 | 4.6052 | 2.4349 | 3.0900 | | 1.7652 | 1.7654 | H11 | 4.4911 | 3.2089 | 2.1335 | 1.0904 | 5.3692 | 5.2971 | 4.6164 | 3.4959 | 2.5552 | 1.7652 | | 1.7486 | H12 | 4.4855 | 3.2112 | 2.1367 | 1.0904 | 5.8308 | 5.0141 | 4.5944 | 3.5032 | 2.5600 | 1.7654 | 1.7486 | |
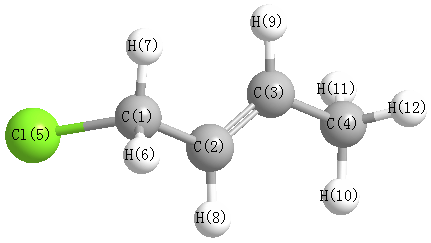 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.346 |
|
C1 |
C2 |
H8 |
116.482 |
| C2 |
C1 |
Cl5 |
111.280 |
|
C2 |
C1 |
H6 |
112.373 |
| C2 |
C1 |
H7 |
111.850 |
|
C2 |
C3 |
C4 |
125.081 |
| C2 |
C3 |
H9 |
118.395 |
|
C3 |
C2 |
H8 |
120.172 |
| C3 |
C4 |
H10 |
111.518 |
|
C3 |
C4 |
H11 |
110.814 |
| C3 |
C4 |
H12 |
111.075 |
|
C4 |
C3 |
H9 |
116.524 |
| Cl5 |
C1 |
H6 |
104.963 |
|
Cl5 |
C1 |
H7 |
106.562 |
| H6 |
C1 |
H7 |
109.435 |
|
H10 |
C4 |
H11 |
108.317 |
| H10 |
C4 |
H12 |
108.332 |
|
H11 |
C4 |
H12 |
106.604 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.160 |
|
|
|
| 2 |
C |
-0.142 |
|
|
|
| 3 |
C |
-0.112 |
|
|
|
| 4 |
C |
-0.327 |
|
|
|
| 5 |
Cl |
-0.172 |
|
|
|
| 6 |
H |
0.146 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
| 9 |
H |
0.129 |
|
|
|
| 10 |
H |
0.115 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.153 |
1.148 |
0.144 |
2.444 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.483 |
-1.900 |
-1.016 |
| y |
-1.900 |
-37.668 |
-0.818 |
| z |
-1.016 |
-0.818 |
-37.102 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.098 |
-1.900 |
-1.016 |
| y |
-1.900 |
0.624 |
-0.818 |
| z |
-1.016 |
-0.818 |
1.474 |
|
| Polar |
|---|
| 3z2-r2 | 2.948 |
|---|
| x2-y2 | -1.815 |
|---|
| xy | -1.900 |
|---|
| xz | -1.016 |
|---|
| yz | -0.818 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.761 |
0.200 |
-0.701 |
| y |
0.200 |
7.318 |
-0.268 |
| z |
-0.701 |
-0.268 |
7.325 |
<r2> (average value of r
2) Å
2
| <r2> |
228.869 |
| (<r2>)1/2 |
15.128 |
