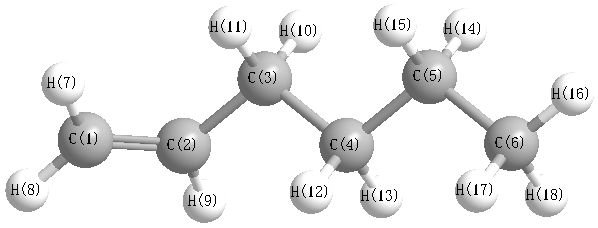
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3241 |
3102 |
17.64 |
|
|
|
| 2 |
A |
3155 |
3020 |
4.33 |
|
|
|
| 3 |
A |
3144 |
3009 |
26.36 |
|
|
|
| 4 |
A |
3126 |
2992 |
37.35 |
|
|
|
| 5 |
A |
3121 |
2988 |
56.50 |
|
|
|
| 6 |
A |
3095 |
2962 |
43.59 |
|
|
|
| 7 |
A |
3077 |
2945 |
3.02 |
|
|
|
| 8 |
A |
3061 |
2930 |
7.96 |
|
|
|
| 9 |
A |
3051 |
2920 |
41.59 |
|
|
|
| 10 |
A |
3045 |
2914 |
53.90 |
|
|
|
| 11 |
A |
3036 |
2906 |
1.17 |
|
|
|
| 12 |
A |
3026 |
2896 |
13.11 |
|
|
|
| 13 |
A |
1730 |
1655 |
16.04 |
|
|
|
| 14 |
A |
1509 |
1444 |
7.75 |
|
|
|
| 15 |
A |
1498 |
1433 |
6.32 |
|
|
|
| 16 |
A |
1496 |
1432 |
2.53 |
|
|
|
| 17 |
A |
1485 |
1421 |
0.52 |
|
|
|
| 18 |
A |
1477 |
1414 |
2.86 |
|
|
|
| 19 |
A |
1452 |
1390 |
1.29 |
|
|
|
| 20 |
A |
1408 |
1348 |
2.96 |
|
|
|
| 21 |
A |
1395 |
1335 |
0.35 |
|
|
|
| 22 |
A |
1350 |
1292 |
0.72 |
|
|
|
| 23 |
A |
1330 |
1273 |
0.74 |
|
|
|
| 24 |
A |
1322 |
1265 |
0.11 |
|
|
|
| 25 |
A |
1307 |
1251 |
0.37 |
|
|
|
| 26 |
A |
1265 |
1210 |
2.04 |
|
|
|
| 27 |
A |
1245 |
1192 |
0.14 |
|
|
|
| 28 |
A |
1198 |
1147 |
0.37 |
|
|
|
| 29 |
A |
1130 |
1082 |
2.44 |
|
|
|
| 30 |
A |
1083 |
1036 |
0.17 |
|
|
|
| 31 |
A |
1055 |
1010 |
8.03 |
|
|
|
| 32 |
A |
1041 |
996 |
5.03 |
|
|
|
| 33 |
A |
1034 |
990 |
4.21 |
|
|
|
| 34 |
A |
956 |
915 |
32.96 |
|
|
|
| 35 |
A |
945 |
904 |
9.96 |
|
|
|
| 36 |
A |
922 |
882 |
2.26 |
|
|
|
| 37 |
A |
908 |
869 |
1.36 |
|
|
|
| 38 |
A |
790 |
756 |
1.12 |
|
|
|
| 39 |
A |
730 |
699 |
3.47 |
|
|
|
| 40 |
A |
654 |
626 |
10.91 |
|
|
|
| 41 |
A |
450 |
431 |
1.21 |
|
|
|
| 42 |
A |
356 |
341 |
0.07 |
|
|
|
| 43 |
A |
344 |
329 |
0.54 |
|
|
|
| 44 |
A |
239 |
229 |
0.00 |
|
|
|
| 45 |
A |
169 |
162 |
0.04 |
|
|
|
| 46 |
A |
122 |
117 |
0.05 |
|
|
|
| 47 |
A |
86 |
82 |
0.03 |
|
|
|
| 48 |
A |
72 |
69 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36364.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34804.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.334 |
|
|
|
| 2 |
C |
-0.105 |
|
|
|
| 3 |
C |
-0.195 |
|
|
|
| 4 |
C |
-0.206 |
|
|
|
| 5 |
C |
-0.207 |
|
|
|
| 6 |
C |
-0.363 |
|
|
|
| 7 |
H |
0.124 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.119 |
|
|
|
| 11 |
H |
0.109 |
|
|
|
| 12 |
H |
0.118 |
|
|
|
| 13 |
H |
0.111 |
|
|
|
| 14 |
H |
0.110 |
|
|
|
| 15 |
H |
0.112 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
H |
0.113 |
|
|
|
| 18 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.425 |
0.092 |
0.114 |
0.449 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.224 |
-0.868 |
-0.478 |
| y |
-0.868 |
-40.668 |
-1.146 |
| z |
-0.478 |
-1.146 |
-39.203 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.289 |
-0.868 |
-0.478 |
| y |
-0.868 |
-0.454 |
-1.146 |
| z |
-0.478 |
-1.146 |
1.743 |
|
| Polar |
|---|
| 3z2-r2 | 3.486 |
|---|
| x2-y2 | -0.556 |
|---|
| xy | -0.868 |
|---|
| xz | -0.478 |
|---|
| yz | -1.146 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.969 |
-0.257 |
-0.964 |
| y |
-0.257 |
9.239 |
-0.342 |
| z |
-0.964 |
-0.342 |
9.491 |
<r2> (average value of r
2) Å
2
| <r2> |
282.264 |
| (<r2>)1/2 |
16.801 |