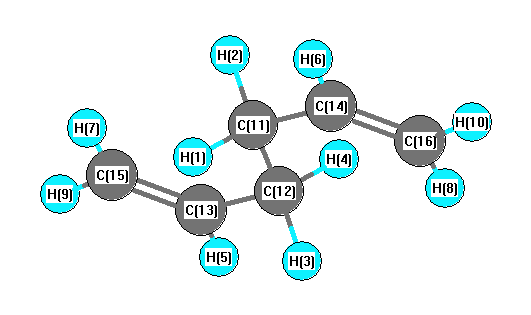
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3242 |
3103 |
7.35 |
|
|
|
| 2 |
A |
3156 |
3020 |
0.11 |
|
|
|
| 3 |
A |
3144 |
3009 |
37.97 |
|
|
|
| 4 |
A |
3104 |
2971 |
19.74 |
|
|
|
| 5 |
A |
3033 |
2903 |
7.34 |
|
|
|
| 6 |
A |
1730 |
1656 |
6.34 |
|
|
|
| 7 |
A |
1477 |
1414 |
3.95 |
|
|
|
| 8 |
A |
1449 |
1387 |
2.63 |
|
|
|
| 9 |
A |
1362 |
1303 |
0.18 |
|
|
|
| 10 |
A |
1323 |
1266 |
1.05 |
|
|
|
| 11 |
A |
1243 |
1190 |
0.01 |
|
|
|
| 12 |
A |
1095 |
1048 |
0.02 |
|
|
|
| 13 |
A |
1051 |
1006 |
1.26 |
|
|
|
| 14 |
A |
1038 |
993 |
8.55 |
|
|
|
| 15 |
A |
960 |
919 |
12.70 |
|
|
|
| 16 |
A |
949 |
909 |
8.42 |
|
|
|
| 17 |
A |
782 |
749 |
0.38 |
|
|
|
| 18 |
A |
640 |
613 |
8.13 |
|
|
|
| 19 |
A |
410 |
392 |
0.01 |
|
|
|
| 20 |
A |
346 |
332 |
0.11 |
|
|
|
| 21 |
A |
101 |
97 |
0.04 |
|
|
|
| 22 |
A |
73 |
70 |
0.05 |
|
|
|
| 23 |
B |
3242 |
3103 |
25.51 |
|
|
|
| 24 |
B |
3156 |
3020 |
8.18 |
|
|
|
| 25 |
B |
3144 |
3009 |
12.75 |
|
|
|
| 26 |
B |
3086 |
2954 |
7.11 |
|
|
|
| 27 |
B |
3037 |
2907 |
34.71 |
|
|
|
| 28 |
B |
1733 |
1659 |
23.49 |
|
|
|
| 29 |
B |
1492 |
1428 |
9.38 |
|
|
|
| 30 |
B |
1452 |
1390 |
0.24 |
|
|
|
| 31 |
B |
1329 |
1272 |
0.84 |
|
|
|
| 32 |
B |
1309 |
1252 |
0.26 |
|
|
|
| 33 |
B |
1267 |
1213 |
2.33 |
|
|
|
| 34 |
B |
1183 |
1132 |
2.07 |
|
|
|
| 35 |
B |
1042 |
997 |
19.32 |
|
|
|
| 36 |
B |
965 |
923 |
9.25 |
|
|
|
| 37 |
B |
955 |
914 |
60.43 |
|
|
|
| 38 |
B |
941 |
900 |
1.43 |
|
|
|
| 39 |
B |
668 |
639 |
19.38 |
|
|
|
| 40 |
B |
434 |
415 |
2.11 |
|
|
|
| 41 |
B |
212 |
203 |
0.23 |
|
|
|
| 42 |
B |
117 |
112 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31234.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29894.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.196 |
|
|
|
| 2 |
C |
-0.196 |
|
|
|
| 3 |
C |
-0.106 |
|
|
|
| 4 |
C |
-0.106 |
|
|
|
| 5 |
C |
-0.331 |
|
|
|
| 6 |
C |
-0.331 |
|
|
|
| 7 |
H |
0.122 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
H |
0.122 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.134 |
|
|
|
| 12 |
H |
0.134 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
| 14 |
H |
0.126 |
|
|
|
| 15 |
H |
0.132 |
|
|
|
| 16 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.269 |
0.269 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.229 |
1.423 |
0.000 |
| y |
1.423 |
-39.663 |
0.000 |
| z |
0.000 |
0.000 |
-37.130 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.833 |
1.423 |
0.000 |
| y |
1.423 |
-1.484 |
0.000 |
| z |
0.000 |
0.000 |
2.316 |
|
| Polar |
|---|
| 3z2-r2 | 4.633 |
|---|
| x2-y2 | 0.434 |
|---|
| xy | 1.423 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.285 |
1.321 |
0.000 |
| y |
1.321 |
13.956 |
0.000 |
| z |
0.000 |
0.000 |
9.819 |
<r2> (average value of r
2) Å
2
| <r2> |
251.660 |
| (<r2>)1/2 |
15.864 |