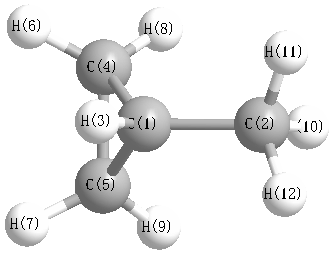
All results from a given calculation for C4H8 (methylcyclopropane)
using model chemistry: CCSD(T)/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -156.912708 |
| Energy at 298.15K | |
| HF Energy | -156.150764 |
| Nuclear repulsion energy | 124.283380 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.351 |
0.410 |
0.000 |
| C2 |
-0.891 |
1.269 |
0.000 |
| H3 |
1.290 |
0.952 |
0.000 |
| C4 |
0.351 |
-0.894 |
0.757 |
| C5 |
0.351 |
-0.894 |
-0.757 |
| H6 |
1.258 |
-1.199 |
1.263 |
| H7 |
1.258 |
-1.199 |
-1.263 |
| H8 |
-0.567 |
-1.177 |
1.258 |
| H9 |
-0.567 |
-1.177 |
-1.258 |
| H10 |
-1.785 |
0.639 |
0.000 |
| H11 |
-0.933 |
1.908 |
0.885 |
| H12 |
-0.933 |
1.908 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5101 | 1.0840 | 1.5084 | 1.5084 | 2.2373 | 2.2373 | 2.2232 | 2.2232 | 2.1483 | 2.1626 | 2.1626 |
C2 | 1.5101 | | 2.2039 | 2.6069 | 2.6069 | 3.5073 | 3.5073 | 2.7691 | 2.7691 | 1.0936 | 1.0930 | 1.0930 | H3 | 1.0840 | 2.2039 | | 2.2055 | 2.2055 | 2.4945 | 2.4945 | 3.0921 | 3.0921 | 3.0908 | 2.5765 | 2.5765 | C4 | 1.5084 | 2.6069 | 2.2055 | | 1.5147 | 1.0816 | 2.2353 | 1.0829 | 2.2323 | 2.7364 | 3.0855 | 3.4932 | C5 | 1.5084 | 2.6069 | 2.2055 | 1.5147 | | 2.2353 | 1.0816 | 2.2323 | 1.0829 | 2.7364 | 3.4932 | 3.0855 | H6 | 2.2373 | 3.5073 | 2.4945 | 1.0816 | 2.2353 | | 2.5262 | 1.8243 | 3.1118 | 3.7722 | 3.8203 | 4.3666 | H7 | 2.2373 | 3.5073 | 2.4945 | 2.2353 | 1.0816 | 2.5262 | | 3.1118 | 1.8243 | 3.7722 | 4.3666 | 3.8203 | H8 | 2.2232 | 2.7691 | 3.0921 | 1.0829 | 2.2323 | 1.8243 | 3.1118 | | 2.5156 | 2.5225 | 3.1290 | 3.7742 | H9 | 2.2232 | 2.7691 | 3.0921 | 2.2323 | 1.0829 | 3.1118 | 1.8243 | 2.5156 | | 2.5225 | 3.7742 | 3.1290 | H10 | 2.1483 | 1.0936 | 3.0908 | 2.7364 | 2.7364 | 3.7722 | 3.7722 | 2.5225 | 2.5225 | | 1.7668 | 1.7668 | H11 | 2.1626 | 1.0930 | 2.5765 | 3.0855 | 3.4932 | 3.8203 | 4.3666 | 3.1290 | 3.7742 | 1.7668 | | 1.7706 | H12 | 2.1626 | 1.0930 | 2.5765 | 3.4932 | 3.0855 | 4.3666 | 3.8203 | 3.7742 | 3.1290 | 1.7668 | 1.7706 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H10 |
110.174 |
|
C1 |
C2 |
H11 |
111.355 |
| C1 |
C2 |
H12 |
111.355 |
|
C1 |
C4 |
H5 |
59.861 |
| C1 |
C4 |
H6 |
118.566 |
|
C1 |
C4 |
H8 |
117.227 |
| C1 |
H5 |
C4 |
59.861 |
|
C1 |
H5 |
H7 |
118.566 |
| C1 |
H5 |
H9 |
117.227 |
|
C2 |
C1 |
C3 |
115.345 |
| C2 |
C1 |
C4 |
119.459 |
|
C2 |
C1 |
H5 |
119.459 |
| C3 |
C1 |
C4 |
115.618 |
|
C3 |
C1 |
H5 |
115.618 |
| C4 |
C1 |
H5 |
60.278 |
|
C4 |
H5 |
H7 |
117.878 |
| C4 |
H5 |
H9 |
117.524 |
|
H5 |
C4 |
H6 |
117.878 |
| H5 |
C4 |
H8 |
117.524 |
|
H6 |
C4 |
H8 |
114.885 |
| H7 |
H5 |
H9 |
114.885 |
|
H10 |
C2 |
H11 |
107.811 |
| H10 |
C2 |
H12 |
107.811 |
|
H11 |
C2 |
H12 |
108.193 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
