return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8 (methylcyclopropane)

using model chemistry: MP4=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4=FULL/cc-pVTZ
 hartrees
Energy at 0K-156.980066
Energy at 298.15K-156.988696
HF Energy-156.151191
Nuclear repulsion energy124.814776
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3242 3122        
2 A' 3175 3058        
3 A' 3165 3047        
4 A' 3102 2987        
5 A' 3045 2932        
6 A' 1529 1473        
7 A' 1517 1461        
8 A' 1439 1386        
9 A' 1403 1351        
10 A' 1245 1198        
11 A' 1223 1177        
12 A' 1054 1015        
13 A' 1012 975        
14 A' 950 915        
15 A' 803 773        
16 A' 779 750        
17 A' 362 349        
18 A" 3228 3108        
19 A" 3158 3041        
20 A" 3107 2992        
21 A" 1508 1452        
22 A" 1481 1426        
23 A" 1205 1160        
24 A" 1157 1114        
25 A" 1118 1077        
26 A" 1073 1033        
27 A" 886 853        
28 A" 829 798        
29 A" 343 330        
30 A" 236 227        

Unscaled Zero Point Vibrational Energy (zpe) 24186.7 cm-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 23289.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/cc-pVTZ
ABC
0.52208 0.21473 0.18854

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.351 0.410 0.000
C2 -0.889 1.260 0.000
H3 1.283 0.954 0.000
C4 0.351 -0.889 0.755
C5 0.351 -0.889 -0.755
H6 1.254 -1.194 1.256
H7 1.254 -1.194 -1.256
H8 -0.563 -1.168 1.255
H9 -0.563 -1.168 -1.255
H10 -1.777 0.627 0.000
H11 -0.934 1.896 0.883
H12 -0.934 1.896 -0.883

Atom - Atom Distances (Å)
  C1 C2 H3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.50351.07911.50201.50202.22832.22832.21342.21342.13862.15442.1544
C21.50352.19342.59302.59303.49183.49182.75202.75201.09011.08941.0894
H31.07912.19342.19862.19862.48832.48833.07973.07973.07692.56572.5657
C41.50202.59302.19861.50961.07732.22521.07882.22552.71903.07003.4772
C51.50202.59302.19861.50962.22521.07732.22551.07882.71903.47723.0700
H62.22833.49182.48831.07732.22522.51141.81803.09983.75233.80534.3491
H72.22833.49182.48832.22521.07732.51143.09981.81803.75234.34913.8053
H82.21342.75203.07971.07882.22551.81803.09982.51002.50343.10873.7545
H92.21342.75203.07972.22551.07883.09981.81802.51002.50343.75453.1087
H102.13861.09013.07692.71902.71903.75233.75232.50342.50341.76101.7610
H112.15441.08942.56573.07003.47723.80534.34913.10873.75451.76101.7658
H122.15441.08942.56573.47723.07004.34913.80533.75453.10871.76101.7658

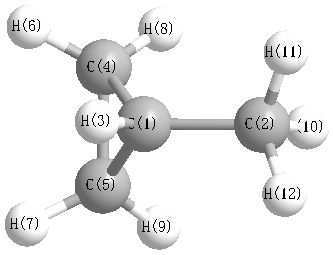
picture of methylcyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H10 110.071 C1 C2 H11 111.381
C1 C2 H12 111.381 C1 C4 H5 59.832
C1 C4 H6 118.594 C1 C4 H8 117.165
C1 H5 C4 59.832 C1 H5 H7 118.594
C1 H5 H9 117.165 C2 C1 C3 115.285
C2 C1 C4 119.259 C2 C1 H5 119.259
C3 C1 C4 115.852 C3 C1 H5 115.852
C4 C1 H5 60.337 C4 H5 H7 117.707
C4 H5 H9 117.627 H5 C4 H6 117.707
H5 C4 H8 117.627 H6 C4 H8 114.958
H7 H5 H9 114.958 H10 C2 H11 107.795
H10 C2 H12 107.795 H11 C2 H12 108.276
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability