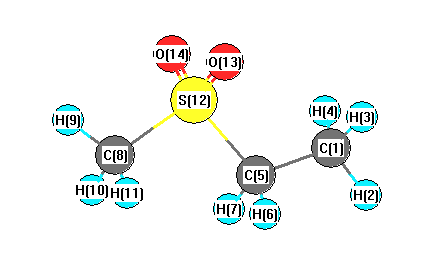
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3200 |
3063 |
1.06 |
|
|
|
| 2 |
A |
3190 |
3053 |
0.96 |
|
|
|
| 3 |
A |
3164 |
3028 |
7.65 |
|
|
|
| 4 |
A |
3146 |
3011 |
7.88 |
|
|
|
| 5 |
A |
3136 |
3002 |
6.69 |
|
|
|
| 6 |
A |
3088 |
2956 |
3.54 |
|
|
|
| 7 |
A |
3086 |
2954 |
2.76 |
|
|
|
| 8 |
A |
3073 |
2941 |
15.05 |
|
|
|
| 9 |
A |
1508 |
1443 |
6.63 |
|
|
|
| 10 |
A |
1501 |
1437 |
9.54 |
|
|
|
| 11 |
A |
1459 |
1397 |
6.26 |
|
|
|
| 12 |
A |
1454 |
1392 |
1.59 |
|
|
|
| 13 |
A |
1448 |
1385 |
4.74 |
|
|
|
| 14 |
A |
1413 |
1352 |
1.57 |
|
|
|
| 15 |
A |
1367 |
1309 |
209.96 |
|
|
|
| 16 |
A |
1337 |
1280 |
26.01 |
|
|
|
| 17 |
A |
1301 |
1245 |
12.15 |
|
|
|
| 18 |
A |
1260 |
1206 |
5.95 |
|
|
|
| 19 |
A |
1169 |
1119 |
144.95 |
|
|
|
| 20 |
A |
1087 |
1040 |
1.68 |
|
|
|
| 21 |
A |
1059 |
1014 |
3.24 |
|
|
|
| 22 |
A |
994 |
952 |
2.34 |
|
|
|
| 23 |
A |
964 |
922 |
30.20 |
|
|
|
| 24 |
A |
952 |
911 |
13.53 |
|
|
|
| 25 |
A |
801 |
766 |
59.84 |
|
|
|
| 26 |
A |
728 |
697 |
15.90 |
|
|
|
| 27 |
A |
660 |
632 |
13.32 |
|
|
|
| 28 |
A |
498 |
477 |
20.93 |
|
|
|
| 29 |
A |
443 |
424 |
31.03 |
|
|
|
| 30 |
A |
397 |
380 |
5.74 |
|
|
|
| 31 |
A |
316 |
303 |
0.46 |
|
|
|
| 32 |
A |
277 |
265 |
1.36 |
|
|
|
| 33 |
A |
248 |
237 |
3.30 |
|
|
|
| 34 |
A |
206 |
197 |
0.73 |
|
|
|
| 35 |
A |
191 |
183 |
0.20 |
|
|
|
| 36 |
A |
86 |
82 |
2.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25103.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24026.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.376 |
|
|
|
| 2 |
H |
0.152 |
|
|
|
| 3 |
H |
0.132 |
|
|
|
| 4 |
H |
0.121 |
|
|
|
| 5 |
C |
-0.269 |
|
|
|
| 6 |
H |
0.153 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
C |
-0.376 |
|
|
|
| 9 |
H |
0.138 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
| 12 |
S |
0.793 |
|
|
|
| 13 |
O |
-0.455 |
|
|
|
| 14 |
O |
-0.451 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.051 |
2.760 |
-2.108 |
4.623 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.855 |
-5.055 |
-1.335 |
| y |
-5.055 |
-41.485 |
-0.543 |
| z |
-1.335 |
-0.543 |
-47.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.745 |
-5.055 |
-1.335 |
| y |
-5.055 |
4.300 |
-0.543 |
| z |
-1.335 |
-0.543 |
-5.045 |
|
| Polar |
|---|
| 3z2-r2 | -10.090 |
|---|
| x2-y2 | -2.370 |
|---|
| xy | -5.055 |
|---|
| xz | -1.335 |
|---|
| yz | -0.543 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.756 |
-0.211 |
0.092 |
| y |
-0.211 |
8.779 |
0.060 |
| z |
0.092 |
0.060 |
8.251 |
<r2> (average value of r
2) Å
2
| <r2> |
188.689 |
| (<r2>)1/2 |
13.736 |