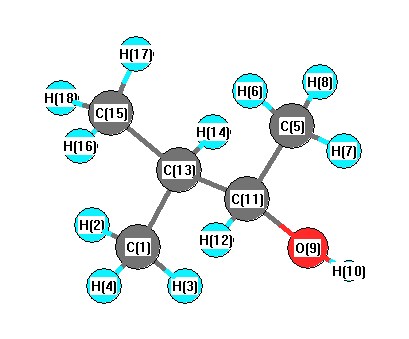
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3877 |
3711 |
22.32 |
|
|
|
| 2 |
A |
3159 |
3024 |
15.31 |
|
|
|
| 3 |
A |
3146 |
3011 |
21.68 |
|
|
|
| 4 |
A |
3134 |
3000 |
32.96 |
|
|
|
| 5 |
A |
3119 |
2985 |
87.24 |
|
|
|
| 6 |
A |
3117 |
2983 |
7.44 |
|
|
|
| 7 |
A |
3113 |
2980 |
7.74 |
|
|
|
| 8 |
A |
3052 |
2921 |
33.56 |
|
|
|
| 9 |
A |
3050 |
2919 |
38.64 |
|
|
|
| 10 |
A |
3047 |
2916 |
18.82 |
|
|
|
| 11 |
A |
3039 |
2909 |
6.69 |
|
|
|
| 12 |
A |
2969 |
2842 |
44.86 |
|
|
|
| 13 |
A |
1527 |
1462 |
5.54 |
|
|
|
| 14 |
A |
1509 |
1444 |
7.19 |
|
|
|
| 15 |
A |
1505 |
1440 |
7.06 |
|
|
|
| 16 |
A |
1494 |
1430 |
5.70 |
|
|
|
| 17 |
A |
1490 |
1426 |
6.44 |
|
|
|
| 18 |
A |
1486 |
1423 |
3.47 |
|
|
|
| 19 |
A |
1427 |
1365 |
11.60 |
|
|
|
| 20 |
A |
1416 |
1355 |
5.60 |
|
|
|
| 21 |
A |
1405 |
1344 |
15.41 |
|
|
|
| 22 |
A |
1402 |
1341 |
1.94 |
|
|
|
| 23 |
A |
1371 |
1312 |
5.69 |
|
|
|
| 24 |
A |
1344 |
1286 |
1.04 |
|
|
|
| 25 |
A |
1331 |
1274 |
15.08 |
|
|
|
| 26 |
A |
1270 |
1215 |
23.94 |
|
|
|
| 27 |
A |
1198 |
1147 |
6.65 |
|
|
|
| 28 |
A |
1185 |
1134 |
13.33 |
|
|
|
| 29 |
A |
1158 |
1109 |
6.43 |
|
|
|
| 30 |
A |
1110 |
1063 |
85.89 |
|
|
|
| 31 |
A |
1089 |
1042 |
8.70 |
|
|
|
| 32 |
A |
990 |
948 |
3.30 |
|
|
|
| 33 |
A |
977 |
935 |
5.48 |
|
|
|
| 34 |
A |
965 |
923 |
9.81 |
|
|
|
| 35 |
A |
930 |
890 |
0.03 |
|
|
|
| 36 |
A |
908 |
869 |
8.64 |
|
|
|
| 37 |
A |
792 |
758 |
1.52 |
|
|
|
| 38 |
A |
521 |
499 |
3.46 |
|
|
|
| 39 |
A |
463 |
443 |
8.28 |
|
|
|
| 40 |
A |
408 |
391 |
1.62 |
|
|
|
| 41 |
A |
392 |
375 |
0.46 |
|
|
|
| 42 |
A |
353 |
338 |
4.02 |
|
|
|
| 43 |
A |
320 |
306 |
3.02 |
|
|
|
| 44 |
A |
257 |
246 |
86.22 |
|
|
|
| 45 |
A |
240 |
230 |
12.72 |
|
|
|
| 46 |
A |
221 |
212 |
0.14 |
|
|
|
| 47 |
A |
205 |
197 |
0.48 |
|
|
|
| 48 |
A |
82 |
78 |
1.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36281.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34724.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.368 |
|
|
|
| 2 |
H |
0.111 |
|
|
|
| 3 |
H |
0.132 |
|
|
|
| 4 |
H |
0.105 |
|
|
|
| 5 |
C |
-0.371 |
|
|
|
| 6 |
H |
0.114 |
|
|
|
| 7 |
H |
0.102 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
O |
-0.338 |
|
|
|
| 10 |
H |
0.202 |
|
|
|
| 11 |
C |
0.074 |
|
|
|
| 12 |
H |
0.091 |
|
|
|
| 13 |
C |
-0.071 |
|
|
|
| 14 |
H |
0.124 |
|
|
|
| 15 |
C |
-0.391 |
|
|
|
| 16 |
H |
0.116 |
|
|
|
| 17 |
H |
0.122 |
|
|
|
| 18 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.801 |
0.933 |
0.739 |
1.435 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.449 |
-0.818 |
1.883 |
| y |
-0.818 |
-41.883 |
-1.119 |
| z |
1.883 |
-1.119 |
-39.804 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.394 |
-0.818 |
1.883 |
| y |
-0.818 |
-3.756 |
-1.119 |
| z |
1.883 |
-1.119 |
-0.638 |
|
| Polar |
|---|
| 3z2-r2 | -1.276 |
|---|
| x2-y2 | 5.433 |
|---|
| xy | -0.818 |
|---|
| xz | 1.883 |
|---|
| yz | -1.119 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.680 |
0.145 |
0.051 |
| y |
0.145 |
9.948 |
-0.012 |
| z |
0.051 |
-0.012 |
8.602 |
<r2> (average value of r
2) Å
2
| <r2> |
192.891 |
| (<r2>)1/2 |
13.889 |