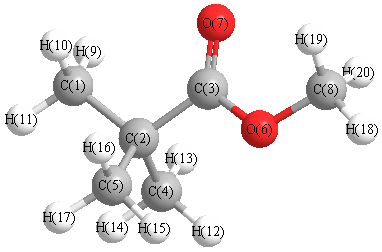
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3046 |
14.23 |
|
|
|
| 2 |
A |
3149 |
3014 |
28.39 |
|
|
|
| 3 |
A |
3149 |
3014 |
33.67 |
|
|
|
| 4 |
A |
3149 |
3014 |
12.13 |
|
|
|
| 5 |
A |
3148 |
3013 |
0.00 |
|
|
|
| 6 |
A |
3141 |
3006 |
26.92 |
|
|
|
| 7 |
A |
3127 |
2993 |
27.44 |
|
|
|
| 8 |
A |
3126 |
2992 |
10.69 |
|
|
|
| 9 |
A |
3073 |
2942 |
32.20 |
|
|
|
| 10 |
A |
3065 |
2933 |
24.73 |
|
|
|
| 11 |
A |
3057 |
2925 |
13.71 |
|
|
|
| 12 |
A |
3055 |
2924 |
29.60 |
|
|
|
| 13 |
A |
1827 |
1749 |
225.07 |
|
|
|
| 14 |
A |
1522 |
1456 |
34.91 |
|
|
|
| 15 |
A |
1504 |
1439 |
15.82 |
|
|
|
| 16 |
A |
1501 |
1437 |
1.91 |
|
|
|
| 17 |
A |
1501 |
1436 |
6.97 |
|
|
|
| 18 |
A |
1488 |
1425 |
0.42 |
|
|
|
| 19 |
A |
1487 |
1424 |
8.69 |
|
|
|
| 20 |
A |
1484 |
1421 |
0.37 |
|
|
|
| 21 |
A |
1478 |
1414 |
0.02 |
|
|
|
| 22 |
A |
1475 |
1412 |
10.34 |
|
|
|
| 23 |
A |
1427 |
1366 |
12.86 |
|
|
|
| 24 |
A |
1401 |
1341 |
5.88 |
|
|
|
| 25 |
A |
1392 |
1332 |
7.27 |
|
|
|
| 26 |
A |
1327 |
1270 |
103.26 |
|
|
|
| 27 |
A |
1267 |
1213 |
8.53 |
|
|
|
| 28 |
A |
1241 |
1188 |
1.74 |
|
|
|
| 29 |
A |
1228 |
1175 |
171.99 |
|
|
|
| 30 |
A |
1196 |
1145 |
134.53 |
|
|
|
| 31 |
A |
1182 |
1131 |
0.88 |
|
|
|
| 32 |
A |
1068 |
1022 |
12.11 |
|
|
|
| 33 |
A |
1050 |
1005 |
0.30 |
|
|
|
| 34 |
A |
1030 |
985 |
4.04 |
|
|
|
| 35 |
A |
966 |
924 |
0.01 |
|
|
|
| 36 |
A |
957 |
916 |
3.04 |
|
|
|
| 37 |
A |
953 |
912 |
1.34 |
|
|
|
| 38 |
A |
888 |
850 |
12.76 |
|
|
|
| 39 |
A |
809 |
775 |
1.87 |
|
|
|
| 40 |
A |
789 |
755 |
4.81 |
|
|
|
| 41 |
A |
588 |
563 |
2.77 |
|
|
|
| 42 |
A |
496 |
475 |
2.53 |
|
|
|
| 43 |
A |
365 |
349 |
1.07 |
|
|
|
| 44 |
A |
362 |
347 |
13.17 |
|
|
|
| 45 |
A |
339 |
325 |
0.14 |
|
|
|
| 46 |
A |
317 |
303 |
4.14 |
|
|
|
| 47 |
A |
295 |
283 |
2.01 |
|
|
|
| 48 |
A |
262 |
251 |
1.22 |
|
|
|
| 49 |
A |
259 |
248 |
0.00 |
|
|
|
| 50 |
A |
217 |
208 |
0.70 |
|
|
|
| 51 |
A |
206 |
197 |
1.89 |
|
|
|
| 52 |
A |
169 |
162 |
0.00 |
|
|
|
| 53 |
A |
127 |
122 |
2.01 |
|
|
|
| 54 |
A |
41 |
40 |
0.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38450.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 36800.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.364 |
|
|
|
| 2 |
C |
-0.021 |
|
|
|
| 3 |
C |
0.338 |
|
|
|
| 4 |
C |
-0.362 |
|
|
|
| 5 |
C |
-0.362 |
|
|
|
| 6 |
O |
-0.224 |
|
|
|
| 7 |
O |
-0.316 |
|
|
|
| 8 |
C |
-0.176 |
|
|
|
| 9 |
H |
0.131 |
|
|
|
| 10 |
H |
0.131 |
|
|
|
| 11 |
H |
0.116 |
|
|
|
| 12 |
H |
0.135 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
H |
0.120 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
H |
0.126 |
|
|
|
| 17 |
H |
0.120 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
| 19 |
H |
0.114 |
|
|
|
| 20 |
H |
0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.102 |
-1.781 |
0.002 |
1.784 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.922 |
-1.812 |
0.001 |
| y |
-1.812 |
-54.877 |
0.001 |
| z |
0.001 |
0.001 |
-48.905 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.969 |
-1.812 |
0.001 |
| y |
-1.812 |
-8.464 |
0.001 |
| z |
0.001 |
0.001 |
0.495 |
|
| Polar |
|---|
| 3z2-r2 | 0.989 |
|---|
| x2-y2 | 10.955 |
|---|
| xy | -1.812 |
|---|
| xz | 0.001 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.845 |
-0.151 |
-0.000 |
| y |
-0.151 |
11.267 |
-0.000 |
| z |
-0.000 |
-0.000 |
10.268 |
<r2> (average value of r
2) Å
2
| <r2> |
293.096 |
| (<r2>)1/2 |
17.120 |