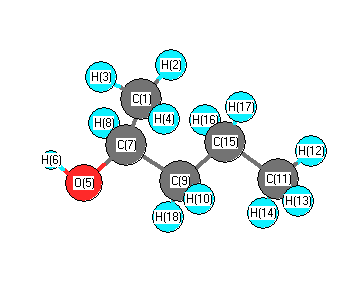
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3873 |
3707 |
18.70 |
|
|
|
| 2 |
A |
3139 |
3004 |
26.02 |
|
|
|
| 3 |
A |
3127 |
2993 |
34.54 |
|
|
|
| 4 |
A |
3123 |
2989 |
65.02 |
|
|
|
| 5 |
A |
3118 |
2984 |
26.63 |
|
|
|
| 6 |
A |
3098 |
2965 |
26.60 |
|
|
|
| 7 |
A |
3072 |
2940 |
1.58 |
|
|
|
| 8 |
A |
3053 |
2922 |
55.88 |
|
|
|
| 9 |
A |
3050 |
2919 |
23.79 |
|
|
|
| 10 |
A |
3049 |
2918 |
23.82 |
|
|
|
| 11 |
A |
3040 |
2909 |
3.88 |
|
|
|
| 12 |
A |
2980 |
2853 |
47.50 |
|
|
|
| 13 |
A |
1510 |
1445 |
6.91 |
|
|
|
| 14 |
A |
1499 |
1435 |
6.41 |
|
|
|
| 15 |
A |
1498 |
1434 |
2.16 |
|
|
|
| 16 |
A |
1495 |
1431 |
4.15 |
|
|
|
| 17 |
A |
1491 |
1427 |
4.12 |
|
|
|
| 18 |
A |
1482 |
1418 |
0.51 |
|
|
|
| 19 |
A |
1427 |
1366 |
19.57 |
|
|
|
| 20 |
A |
1412 |
1351 |
4.29 |
|
|
|
| 21 |
A |
1406 |
1346 |
23.08 |
|
|
|
| 22 |
A |
1388 |
1328 |
3.58 |
|
|
|
| 23 |
A |
1373 |
1314 |
1.78 |
|
|
|
| 24 |
A |
1325 |
1268 |
0.51 |
|
|
|
| 25 |
A |
1307 |
1251 |
21.56 |
|
|
|
| 26 |
A |
1282 |
1227 |
6.50 |
|
|
|
| 27 |
A |
1246 |
1193 |
13.36 |
|
|
|
| 28 |
A |
1187 |
1136 |
12.90 |
|
|
|
| 29 |
A |
1155 |
1105 |
30.68 |
|
|
|
| 30 |
A |
1104 |
1057 |
35.40 |
|
|
|
| 31 |
A |
1102 |
1054 |
25.40 |
|
|
|
| 32 |
A |
1057 |
1012 |
3.38 |
|
|
|
| 33 |
A |
1006 |
963 |
4.56 |
|
|
|
| 34 |
A |
967 |
926 |
7.54 |
|
|
|
| 35 |
A |
920 |
881 |
19.56 |
|
|
|
| 36 |
A |
874 |
837 |
0.48 |
|
|
|
| 37 |
A |
844 |
808 |
0.09 |
|
|
|
| 38 |
A |
744 |
712 |
3.05 |
|
|
|
| 39 |
A |
483 |
462 |
6.74 |
|
|
|
| 40 |
A |
439 |
420 |
1.26 |
|
|
|
| 41 |
A |
408 |
391 |
9.98 |
|
|
|
| 42 |
A |
311 |
298 |
2.77 |
|
|
|
| 43 |
A |
264 |
253 |
99.75 |
|
|
|
| 44 |
A |
255 |
245 |
1.74 |
|
|
|
| 45 |
A |
237 |
227 |
1.63 |
|
|
|
| 46 |
A |
174 |
167 |
1.48 |
|
|
|
| 47 |
A |
113 |
108 |
0.01 |
|
|
|
| 48 |
A |
79 |
76 |
0.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36293.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34736.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.364 |
|
|
|
| 2 |
H |
0.111 |
|
|
|
| 3 |
H |
0.103 |
|
|
|
| 4 |
H |
0.124 |
|
|
|
| 5 |
O |
-0.337 |
|
|
|
| 6 |
H |
0.200 |
|
|
|
| 7 |
C |
0.066 |
|
|
|
| 8 |
H |
0.086 |
|
|
|
| 9 |
C |
-0.208 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
C |
-0.364 |
|
|
|
| 12 |
H |
0.118 |
|
|
|
| 13 |
H |
0.113 |
|
|
|
| 14 |
H |
0.114 |
|
|
|
| 15 |
C |
-0.228 |
|
|
|
| 16 |
H |
0.112 |
|
|
|
| 17 |
H |
0.116 |
|
|
|
| 18 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.334 |
1.300 |
-0.687 |
1.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.184 |
-1.757 |
1.567 |
| y |
-1.757 |
-42.298 |
0.721 |
| z |
1.567 |
0.721 |
-39.755 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.842 |
-1.757 |
1.567 |
| y |
-1.757 |
-4.328 |
0.721 |
| z |
1.567 |
0.721 |
-0.514 |
|
| Polar |
|---|
| 3z2-r2 | -1.027 |
|---|
| x2-y2 | 6.113 |
|---|
| xy | -1.757 |
|---|
| xz | 1.567 |
|---|
| yz | 0.721 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.393 |
-0.197 |
0.053 |
| y |
-0.197 |
9.291 |
-0.089 |
| z |
0.053 |
-0.089 |
8.655 |
<r2> (average value of r
2) Å
2
| <r2> |
236.406 |
| (<r2>)1/2 |
15.375 |