Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -190.339390 |
| Energy at 298.15K | |
| HF Energy | -189.604159 |
| Nuclear repulsion energy | 88.012978 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3096 |
2937 |
|
|
|
|
| 2 |
A1 |
2278 |
2161 |
|
|
|
|
| 3 |
A1 |
1748 |
1658 |
|
|
|
|
| 4 |
A1 |
1488 |
1411 |
|
|
|
|
| 5 |
A1 |
904 |
857 |
|
|
|
|
| 6 |
B1 |
995 |
944 |
|
|
|
|
| 7 |
B1 |
618 |
587 |
|
|
|
|
| 8 |
B1 |
210 |
199 |
|
|
|
|
| 9 |
B2 |
3172 |
3009 |
|
|
|
|
| 10 |
B2 |
1065 |
1011 |
|
|
|
|
| 11 |
B2 |
432 |
409 |
|
|
|
|
| 12 |
B2 |
178i |
169i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7912.8 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 7506.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.884 |
| C2 |
0.000 |
0.000 |
-0.559 |
| C3 |
0.000 |
0.000 |
0.723 |
| O4 |
0.000 |
0.000 |
1.904 |
| H5 |
0.000 |
0.925 |
-2.456 |
| H6 |
0.000 |
-0.925 |
-2.456 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3258 | 2.6074 | 3.7886 | 1.0874 | 1.0874 |
C2 | 1.3258 | | 1.2817 | 2.4628 | 2.1107 | 2.1107 | C3 | 2.6074 | 1.2817 | | 1.1812 | 3.3108 | 3.3108 | O4 | 3.7886 | 2.4628 | 1.1812 | | 4.4571 | 4.4571 | H5 | 1.0874 | 2.1107 | 3.3108 | 4.4571 | | 1.8502 | H6 | 1.0874 | 2.1107 | 3.3108 | 4.4571 | 1.8502 | |
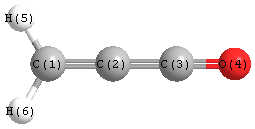 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.704 |
| C2 |
C1 |
H6 |
121.704 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.593 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -190.341737 |
| Energy at 298.15K | |
| HF Energy | -189.603224 |
| Nuclear repulsion energy | 88.327311 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3192 |
3028 |
|
|
|
|
| 2 |
A' |
3110 |
2950 |
|
|
|
|
| 3 |
A' |
2158 |
2047 |
|
|
|
|
| 4 |
A' |
1704 |
1617 |
|
|
|
|
| 5 |
A' |
1487 |
1411 |
|
|
|
|
| 6 |
A' |
1076 |
1021 |
|
|
|
|
| 7 |
A' |
930 |
882 |
|
|
|
|
| 8 |
A' |
500 |
474 |
|
|
|
|
| 9 |
A' |
188 |
178 |
|
|
|
|
| 10 |
A" |
1006 |
954 |
|
|
|
|
| 11 |
A" |
702 |
666 |
|
|
|
|
| 12 |
A" |
272 |
258 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8162.3 cm
-1
Scaled (by 0.9486) Zero Point Vibrational Energy (zpe) 7742.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.103 |
-1.440 |
0.000 |
| C2 |
0.000 |
-0.684 |
0.000 |
| C3 |
-0.370 |
0.579 |
0.000 |
| O4 |
-0.942 |
1.602 |
0.000 |
| H5 |
2.106 |
-1.020 |
0.000 |
| H6 |
1.033 |
-2.523 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3374 | 2.4992 | 3.6657 | 1.0869 | 1.0853 |
C2 | 1.3374 | | 1.3160 | 2.4727 | 2.1327 | 2.1089 | C3 | 2.4992 | 1.3160 | | 1.1724 | 2.9476 | 3.4041 | O4 | 3.6657 | 2.4727 | 1.1724 | | 4.0209 | 4.5733 | H5 | 1.0869 | 2.1327 | 2.9476 | 4.0209 | | 1.8465 | H6 | 1.0853 | 2.1089 | 3.4041 | 4.5733 | 1.8465 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
140.751 |
|
C2 |
C1 |
H5 |
122.881 |
| C2 |
C1 |
H6 |
120.677 |
|
C2 |
C3 |
O4 |
167.150 |
| H5 |
C1 |
H6 |
116.442 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
