Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -190.349976 |
| Energy at 298.15K | |
| HF Energy | -189.605648 |
| Nuclear repulsion energy | 88.499026 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3141 |
2982 |
26.57 |
204.22 |
0.12 |
0.21 |
| 2 |
A1 |
2334 |
2216 |
756.85 |
40.88 |
0.38 |
0.55 |
| 3 |
A1 |
1785 |
1695 |
16.93 |
9.72 |
0.44 |
0.61 |
| 4 |
A1 |
1507 |
1430 |
2.01 |
14.56 |
0.54 |
0.70 |
| 5 |
A1 |
920 |
874 |
2.18 |
39.72 |
0.24 |
0.38 |
| 6 |
B1 |
1040 |
987 |
23.58 |
0.57 |
0.75 |
0.86 |
| 7 |
B1 |
638 |
606 |
17.29 |
0.30 |
0.75 |
0.86 |
| 8 |
B1 |
219 |
208 |
0.60 |
2.20 |
0.75 |
0.86 |
| 9 |
B2 |
3203 |
3041 |
5.32 |
114.92 |
0.75 |
0.86 |
| 10 |
B2 |
1082 |
1028 |
2.09 |
0.08 |
0.75 |
0.86 |
| 11 |
B2 |
461 |
437 |
8.98 |
1.73 |
0.75 |
0.86 |
| 12 |
B2 |
193i |
183i |
18.85 |
0.24 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8068.0 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 7659.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.872 |
| C2 |
0.000 |
0.000 |
-0.557 |
| C3 |
0.000 |
0.000 |
0.717 |
| O4 |
0.000 |
0.000 |
1.894 |
| H5 |
0.000 |
0.919 |
-2.443 |
| H6 |
0.000 |
-0.919 |
-2.443 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3153 | 2.5894 | 3.7666 | 1.0820 | 1.0820 |
C2 | 1.3153 | | 1.2741 | 2.4513 | 2.0980 | 2.0980 | C3 | 2.5894 | 1.2741 | | 1.1772 | 3.2910 | 3.2910 | O4 | 3.7666 | 2.4513 | 1.1772 | | 4.4335 | 4.4335 | H5 | 1.0820 | 2.0980 | 3.2910 | 4.4335 | | 1.8386 | H6 | 1.0820 | 2.0980 | 3.2910 | 4.4335 | 1.8386 | |
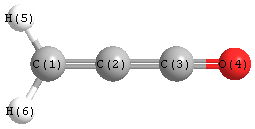 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.828 |
| C2 |
C1 |
H6 |
121.828 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.345 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -190.352364 |
| Energy at 298.15K | |
| HF Energy | -189.604840 |
| Nuclear repulsion energy | 88.823096 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3224 |
3061 |
4.74 |
101.55 |
0.67 |
0.80 |
| 2 |
A' |
3154 |
2994 |
24.53 |
153.59 |
0.12 |
0.22 |
| 3 |
A' |
2236 |
2123 |
738.67 |
52.94 |
0.43 |
0.61 |
| 4 |
A' |
1746 |
1657 |
5.30 |
3.01 |
0.66 |
0.80 |
| 5 |
A' |
1511 |
1434 |
1.23 |
13.60 |
0.57 |
0.72 |
| 6 |
A' |
1098 |
1043 |
21.64 |
5.02 |
0.07 |
0.13 |
| 7 |
A' |
955 |
906 |
1.84 |
36.66 |
0.16 |
0.28 |
| 8 |
A' |
530 |
503 |
10.98 |
4.60 |
0.70 |
0.82 |
| 9 |
A' |
188 |
178 |
20.26 |
4.86 |
0.75 |
0.86 |
| 10 |
A" |
1058 |
1005 |
23.48 |
0.73 |
0.75 |
0.86 |
| 11 |
A" |
720 |
683 |
9.71 |
0.67 |
0.75 |
0.86 |
| 12 |
A" |
291 |
276 |
2.71 |
3.15 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8354.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 7932.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.078 |
-1.449 |
0.000 |
| C2 |
0.000 |
-0.675 |
0.000 |
| C3 |
-0.372 |
0.573 |
0.000 |
| O4 |
-0.913 |
1.610 |
0.000 |
| H5 |
2.084 |
-1.051 |
0.000 |
| H6 |
0.987 |
-2.524 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3265 | 2.4874 | 3.6495 | 1.0818 | 1.0797 |
C2 | 1.3265 | | 1.3021 | 2.4609 | 2.1172 | 2.0964 | C3 | 2.4874 | 1.3021 | | 1.1701 | 2.9437 | 3.3824 | O4 | 3.6495 | 2.4609 | 1.1701 | | 4.0075 | 4.5504 | H5 | 1.0818 | 2.1172 | 2.9437 | 4.0075 | | 1.8369 | H6 | 1.0797 | 2.0964 | 3.3824 | 4.5504 | 1.8369 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
142.268 |
|
C2 |
C1 |
H5 |
122.748 |
| C2 |
C1 |
H6 |
120.868 |
|
C2 |
C3 |
O4 |
169.051 |
| H5 |
C1 |
H6 |
116.384 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
