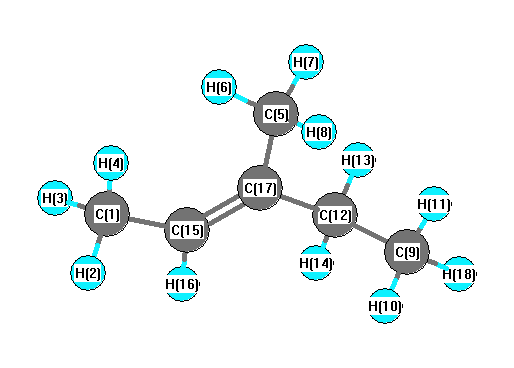
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3162 |
3026 |
34.87 |
|
|
|
| 2 |
A |
3151 |
3016 |
29.20 |
|
|
|
| 3 |
A |
3134 |
3000 |
20.09 |
|
|
|
| 4 |
A |
3133 |
2998 |
20.18 |
|
|
|
| 5 |
A |
3128 |
2994 |
33.98 |
|
|
|
| 6 |
A |
3095 |
2963 |
21.65 |
|
|
|
| 7 |
A |
3089 |
2956 |
19.81 |
|
|
|
| 8 |
A |
3087 |
2954 |
11.47 |
|
|
|
| 9 |
A |
3057 |
2925 |
30.64 |
|
|
|
| 10 |
A |
3047 |
2917 |
17.64 |
|
|
|
| 11 |
A |
3043 |
2913 |
46.26 |
|
|
|
| 12 |
A |
3027 |
2898 |
32.25 |
|
|
|
| 13 |
A |
1762 |
1687 |
2.14 |
|
|
|
| 14 |
A |
1509 |
1444 |
1.62 |
|
|
|
| 15 |
A |
1503 |
1439 |
10.54 |
|
|
|
| 16 |
A |
1496 |
1432 |
3.93 |
|
|
|
| 17 |
A |
1489 |
1425 |
12.92 |
|
|
|
| 18 |
A |
1486 |
1422 |
6.01 |
|
|
|
| 19 |
A |
1480 |
1417 |
4.37 |
|
|
|
| 20 |
A |
1473 |
1410 |
0.34 |
|
|
|
| 21 |
A |
1423 |
1362 |
3.59 |
|
|
|
| 22 |
A |
1415 |
1354 |
4.12 |
|
|
|
| 23 |
A |
1403 |
1343 |
2.45 |
|
|
|
| 24 |
A |
1375 |
1316 |
1.62 |
|
|
|
| 25 |
A |
1357 |
1298 |
2.62 |
|
|
|
| 26 |
A |
1286 |
1231 |
0.21 |
|
|
|
| 27 |
A |
1243 |
1190 |
3.64 |
|
|
|
| 28 |
A |
1146 |
1097 |
7.74 |
|
|
|
| 29 |
A |
1101 |
1054 |
3.67 |
|
|
|
| 30 |
A |
1092 |
1045 |
3.58 |
|
|
|
| 31 |
A |
1061 |
1015 |
1.43 |
|
|
|
| 32 |
A |
1043 |
998 |
1.46 |
|
|
|
| 33 |
A |
1018 |
974 |
3.81 |
|
|
|
| 34 |
A |
995 |
952 |
8.70 |
|
|
|
| 35 |
A |
937 |
897 |
1.61 |
|
|
|
| 36 |
A |
857 |
820 |
11.87 |
|
|
|
| 37 |
A |
788 |
754 |
1.98 |
|
|
|
| 38 |
A |
763 |
730 |
1.28 |
|
|
|
| 39 |
A |
551 |
527 |
4.83 |
|
|
|
| 40 |
A |
493 |
472 |
2.84 |
|
|
|
| 41 |
A |
398 |
381 |
0.43 |
|
|
|
| 42 |
A |
321 |
307 |
0.47 |
|
|
|
| 43 |
A |
312 |
298 |
0.51 |
|
|
|
| 44 |
A |
250 |
239 |
0.15 |
|
|
|
| 45 |
A |
189 |
181 |
0.80 |
|
|
|
| 46 |
A |
130 |
125 |
0.19 |
|
|
|
| 47 |
A |
119 |
114 |
0.74 |
|
|
|
| 48 |
A |
66 |
63 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36239.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34684.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.346 |
|
|
|
| 2 |
H |
0.118 |
|
|
|
| 3 |
H |
0.118 |
|
|
|
| 4 |
H |
0.109 |
|
|
|
| 5 |
C |
-0.374 |
|
|
|
| 6 |
H |
0.118 |
|
|
|
| 7 |
H |
0.119 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
C |
-0.362 |
|
|
|
| 10 |
H |
0.119 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
C |
-0.211 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.107 |
|
|
|
| 15 |
C |
-0.165 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
C |
0.057 |
|
|
|
| 18 |
H |
0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.030 |
0.172 |
0.018 |
0.175 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.980 |
0.160 |
-0.668 |
| y |
0.160 |
-38.881 |
0.024 |
| z |
-0.668 |
0.024 |
-40.946 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.934 |
0.160 |
-0.668 |
| y |
0.160 |
1.082 |
0.024 |
| z |
-0.668 |
0.024 |
-2.015 |
|
| Polar |
|---|
| 3z2-r2 | -4.031 |
|---|
| x2-y2 | -0.099 |
|---|
| xy | 0.160 |
|---|
| xz | -0.668 |
|---|
| yz | 0.024 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.655 |
0.932 |
-0.359 |
| y |
0.932 |
10.648 |
0.004 |
| z |
-0.359 |
0.004 |
8.590 |
<r2> (average value of r
2) Å
2
| <r2> |
220.612 |
| (<r2>)1/2 |
14.853 |