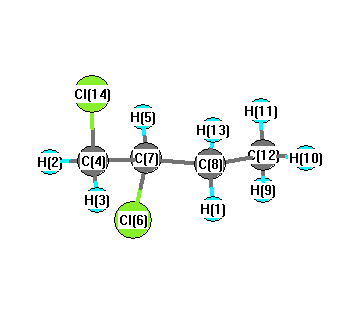
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3188 |
3051 |
2.14 |
|
|
|
| 2 |
A |
3146 |
3011 |
18.23 |
|
|
|
| 3 |
A |
3135 |
3000 |
32.53 |
|
|
|
| 4 |
A |
3124 |
2990 |
8.93 |
|
|
|
| 5 |
A |
3116 |
2982 |
0.47 |
|
|
|
| 6 |
A |
3099 |
2966 |
1.45 |
|
|
|
| 7 |
A |
3063 |
2931 |
30.93 |
|
|
|
| 8 |
A |
3060 |
2929 |
4.15 |
|
|
|
| 9 |
A |
1506 |
1442 |
7.24 |
|
|
|
| 10 |
A |
1496 |
1432 |
7.68 |
|
|
|
| 11 |
A |
1483 |
1419 |
5.41 |
|
|
|
| 12 |
A |
1475 |
1412 |
4.38 |
|
|
|
| 13 |
A |
1412 |
1352 |
3.95 |
|
|
|
| 14 |
A |
1402 |
1342 |
0.52 |
|
|
|
| 15 |
A |
1337 |
1279 |
6.34 |
|
|
|
| 16 |
A |
1324 |
1267 |
1.59 |
|
|
|
| 17 |
A |
1291 |
1235 |
5.86 |
|
|
|
| 18 |
A |
1242 |
1189 |
6.95 |
|
|
|
| 19 |
A |
1201 |
1149 |
13.63 |
|
|
|
| 20 |
A |
1140 |
1091 |
1.00 |
|
|
|
| 21 |
A |
1098 |
1051 |
1.64 |
|
|
|
| 22 |
A |
1073 |
1027 |
3.16 |
|
|
|
| 23 |
A |
1051 |
1006 |
1.34 |
|
|
|
| 24 |
A |
950 |
909 |
9.04 |
|
|
|
| 25 |
A |
842 |
806 |
6.62 |
|
|
|
| 26 |
A |
814 |
779 |
7.96 |
|
|
|
| 27 |
A |
753 |
721 |
54.52 |
|
|
|
| 28 |
A |
673 |
645 |
32.66 |
|
|
|
| 29 |
A |
457 |
438 |
1.77 |
|
|
|
| 30 |
A |
380 |
364 |
1.45 |
|
|
|
| 31 |
A |
295 |
282 |
0.03 |
|
|
|
| 32 |
A |
235 |
225 |
0.55 |
|
|
|
| 33 |
A |
212 |
203 |
2.89 |
|
|
|
| 34 |
A |
188 |
180 |
3.24 |
|
|
|
| 35 |
A |
119 |
114 |
3.39 |
|
|
|
| 36 |
A |
93 |
89 |
1.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25234.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24152.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.132 |
|
|
|
| 2 |
H |
0.168 |
|
|
|
| 3 |
H |
0.161 |
|
|
|
| 4 |
C |
-0.164 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
Cl |
-0.163 |
|
|
|
| 7 |
C |
-0.084 |
|
|
|
| 8 |
C |
-0.207 |
|
|
|
| 9 |
H |
0.130 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
C |
-0.360 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
Cl |
-0.159 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.271 |
0.594 |
0.229 |
0.692 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-57.028 |
3.154 |
-0.194 |
| y |
3.154 |
-50.789 |
-0.167 |
| z |
-0.194 |
-0.167 |
-50.311 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.478 |
3.154 |
-0.194 |
| y |
3.154 |
2.881 |
-0.167 |
| z |
-0.194 |
-0.167 |
3.597 |
|
| Polar |
|---|
| 3z2-r2 | 7.195 |
|---|
| x2-y2 | -6.239 |
|---|
| xy | 3.154 |
|---|
| xz | -0.194 |
|---|
| yz | -0.167 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.717 |
-0.791 |
0.138 |
| y |
-0.791 |
10.927 |
-0.173 |
| z |
0.138 |
-0.173 |
8.400 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |