return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H6O3S (Sulfurous acid, dimethyl ester)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-702.776466
Energy at 298.15K-702.784491
HF Energy-701.417244
Nuclear repulsion energy352.446871
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3224 3061 6.93      
2 A 3210 3048 6.60      
3 A 3205 3042 6.57      
4 A 3169 3009 15.16      
5 A 3119 2961 26.80      
6 A 3087 2931 40.47      
7 A 1540 1462 13.58      
8 A 1539 1461 6.77      
9 A 1520 1443 12.31      
10 A 1520 1443 2.83      
11 A 1482 1407 0.57      
12 A 1474 1399 0.57      
13 A 1254 1191 137.30      
14 A 1203 1142 4.93      
15 A 1201 1140 3.61      
16 A 1189 1128 2.05      
17 A 1185 1125 0.92      
18 A 1067 1013 115.44      
19 A 1036 983 254.96      
20 A 763 724 102.71      
21 A 703 668 222.44      
22 A 580 550 14.31      
23 A 466 442 2.48      
24 A 397 377 7.00      
25 A 296 281 5.94      
26 A 233 221 6.62      
27 A 152 144 6.25      
28 A 126 120 0.92      
29 A 90 85 4.18      
30 A 66 63 2.04      

Unscaled Zero Point Vibrational Energy (zpe) 20047.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19032.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.16778 0.07995 0.06829

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -0.024 0.595 -0.481
O2 -0.058 1.333 0.772
O3 -0.860 -0.786 -0.212
O4 1.374 -0.161 -0.625
C5 -2.197 -0.570 0.235
C6 1.900 -0.791 0.551
H7 -2.649 -1.549 0.307
H8 2.898 -1.108 0.286
H9 -2.203 -0.085 1.204
H10 -2.758 0.031 -0.477
H11 1.935 -0.082 1.370
H12 1.293 -1.646 0.822

Atom - Atom Distances (Å)
  S1 O2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12
S11.45461.63691.59642.56722.58623.47993.46782.83712.79132.77922.9074
O21.45462.47082.49752.91322.89743.90393.86432.60763.24732.51693.2717
O31.63692.47082.35691.42552.86402.01283.80462.07372.08283.28832.5385
O41.59642.49752.35693.69581.43434.35712.01194.01874.13922.07432.0747
C52.56722.91321.42553.69584.11481.08085.12291.08401.08754.31243.6984
C62.58622.89742.86401.43434.11484.61851.08004.21424.84021.08321.0830
H73.47993.90392.01284.35711.08084.61855.56451.77431.76714.92913.9767
H83.46783.86433.80462.01195.12291.08005.56455.28275.81931.77581.7752
H92.83712.60762.07374.01871.08404.21421.77435.28271.77374.14143.8478
H102.79133.24732.08284.13921.08754.84021.76715.81931.77375.04444.5722
H112.77922.51693.28832.07434.31241.08324.92911.77584.14145.04441.7769
H122.90743.27172.53852.07473.69841.08303.97671.77523.84784.57221.7769

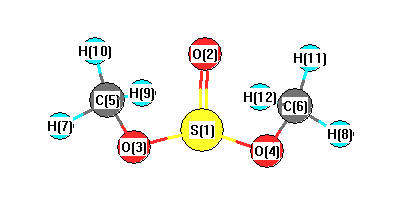
picture of Sulfurous acid, dimethyl ester state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 O3 C5 113.740 S1 O4 C6 117.052
O2 S1 O3 105.962 O2 S1 O4 109.804
O3 S1 O4 93.589 O3 C5 H7 106.038
O3 C5 H9 110.716 O3 C5 H10 111.242
O4 C6 H8 105.431 O4 C6 H11 110.181
O4 C6 H12 110.230 H7 C5 H9 110.091
H7 C5 H10 109.163 H8 C6 H11 110.356
H8 C6 H12 110.321 H9 C5 H10 109.530
H11 C6 H12 110.227
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability