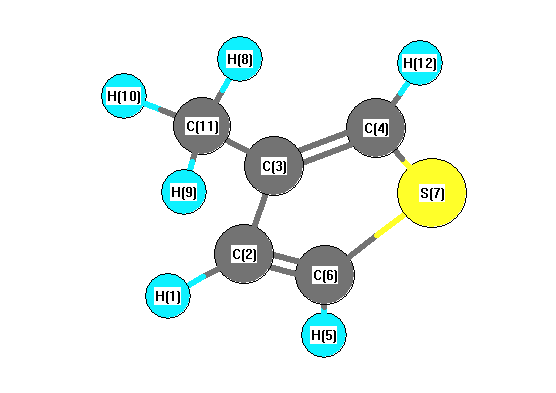
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3274 |
3133 |
0.75 |
|
|
|
| 2 |
A' |
3262 |
3122 |
0.86 |
|
|
|
| 3 |
A' |
3221 |
3083 |
4.89 |
|
|
|
| 4 |
A' |
3143 |
3009 |
14.98 |
|
|
|
| 5 |
A' |
3054 |
2923 |
30.77 |
|
|
|
| 6 |
A' |
1616 |
1546 |
4.08 |
|
|
|
| 7 |
A' |
1501 |
1436 |
12.33 |
|
|
|
| 8 |
A' |
1470 |
1407 |
1.71 |
|
|
|
| 9 |
A' |
1433 |
1372 |
5.53 |
|
|
|
| 10 |
A' |
1408 |
1347 |
0.07 |
|
|
|
| 11 |
A' |
1262 |
1208 |
2.96 |
|
|
|
| 12 |
A' |
1185 |
1134 |
2.26 |
|
|
|
| 13 |
A' |
1104 |
1057 |
3.48 |
|
|
|
| 14 |
A' |
1009 |
966 |
8.67 |
|
|
|
| 15 |
A' |
952 |
911 |
2.23 |
|
|
|
| 16 |
A' |
888 |
850 |
13.68 |
|
|
|
| 17 |
A' |
862 |
825 |
7.66 |
|
|
|
| 18 |
A' |
671 |
643 |
0.25 |
|
|
|
| 19 |
A' |
550 |
527 |
0.21 |
|
|
|
| 20 |
A' |
316 |
303 |
0.79 |
|
|
|
| 21 |
A" |
3111 |
2978 |
15.64 |
|
|
|
| 22 |
A" |
1483 |
1420 |
6.37 |
|
|
|
| 23 |
A" |
1058 |
1012 |
2.43 |
|
|
|
| 24 |
A" |
909 |
870 |
0.04 |
|
|
|
| 25 |
A" |
790 |
756 |
67.54 |
|
|
|
| 26 |
A" |
702 |
672 |
5.71 |
|
|
|
| 27 |
A" |
613 |
587 |
11.61 |
|
|
|
| 28 |
A" |
470 |
450 |
0.01 |
|
|
|
| 29 |
A" |
236 |
225 |
3.17 |
|
|
|
| 30 |
A" |
112 |
107 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20832.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 19938.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.137 |
|
|
|
| 2 |
C |
-0.186 |
|
|
|
| 3 |
C |
0.100 |
|
|
|
| 4 |
C |
-0.239 |
|
|
|
| 5 |
H |
0.147 |
|
|
|
| 6 |
C |
-0.193 |
|
|
|
| 7 |
S |
0.091 |
|
|
|
| 8 |
H |
0.114 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
C |
-0.360 |
|
|
|
| 12 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.297 |
0.840 |
0.000 |
0.891 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.674 |
1.464 |
0.000 |
| y |
1.464 |
-40.404 |
0.000 |
| z |
0.000 |
0.000 |
-45.881 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.469 |
1.464 |
0.000 |
| y |
1.464 |
1.874 |
0.000 |
| z |
0.000 |
0.000 |
-6.342 |
|
| Polar |
|---|
| 3z2-r2 | -12.685 |
|---|
| x2-y2 | 1.730 |
|---|
| xy | 1.464 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.173 |
-0.404 |
0.000 |
| y |
-0.404 |
13.551 |
0.000 |
| z |
0.000 |
0.000 |
6.327 |
<r2> (average value of r
2) Å
2
| <r2> |
185.028 |
| (<r2>)1/2 |
13.602 |