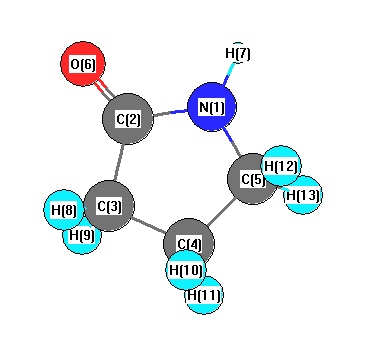
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3675 |
3525 |
33.95 |
|
|
|
| 2 |
A |
3148 |
3020 |
13.51 |
|
|
|
| 3 |
A |
3136 |
3008 |
17.42 |
|
|
|
| 4 |
A |
3096 |
2970 |
42.91 |
|
|
|
| 5 |
A |
3080 |
2955 |
12.35 |
|
|
|
| 6 |
A |
3073 |
2947 |
10.77 |
|
|
|
| 7 |
A |
3018 |
2895 |
62.41 |
|
|
|
| 8 |
A |
1836 |
1761 |
478.31 |
|
|
|
| 9 |
A |
1541 |
1478 |
3.69 |
|
|
|
| 10 |
A |
1500 |
1439 |
10.28 |
|
|
|
| 11 |
A |
1470 |
1410 |
3.59 |
|
|
|
| 12 |
A |
1460 |
1400 |
41.54 |
|
|
|
| 13 |
A |
1380 |
1323 |
9.29 |
|
|
|
| 14 |
A |
1344 |
1289 |
16.17 |
|
|
|
| 15 |
A |
1312 |
1259 |
41.88 |
|
|
|
| 16 |
A |
1280 |
1228 |
61.74 |
|
|
|
| 17 |
A |
1255 |
1204 |
7.74 |
|
|
|
| 18 |
A |
1220 |
1171 |
1.39 |
|
|
|
| 19 |
A |
1193 |
1144 |
4.17 |
|
|
|
| 20 |
A |
1101 |
1056 |
9.09 |
|
|
|
| 21 |
A |
1091 |
1047 |
7.44 |
|
|
|
| 22 |
A |
1018 |
977 |
11.22 |
|
|
|
| 23 |
A |
937 |
899 |
0.46 |
|
|
|
| 24 |
A |
914 |
876 |
1.46 |
|
|
|
| 25 |
A |
901 |
864 |
4.53 |
|
|
|
| 26 |
A |
823 |
789 |
5.12 |
|
|
|
| 27 |
A |
701 |
672 |
6.68 |
|
|
|
| 28 |
A |
644 |
618 |
19.35 |
|
|
|
| 29 |
A |
573 |
550 |
36.70 |
|
|
|
| 30 |
A |
493 |
473 |
57.61 |
|
|
|
| 31 |
A |
473 |
454 |
11.27 |
|
|
|
| 32 |
A |
193 |
185 |
3.43 |
|
|
|
| 33 |
A |
145 |
139 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24511.4 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 23511.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.126 |
|
|
|
| 2 |
C |
0.179 |
|
|
|
| 3 |
C |
-0.161 |
|
|
|
| 4 |
C |
-0.165 |
|
|
|
| 5 |
C |
-0.047 |
|
|
|
| 6 |
O |
-0.333 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
H |
0.097 |
|
|
|
| 9 |
H |
0.097 |
|
|
|
| 10 |
H |
0.085 |
|
|
|
| 11 |
H |
0.089 |
|
|
|
| 12 |
H |
0.070 |
|
|
|
| 13 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.990 |
-0.650 |
0.329 |
4.056 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.682 |
-0.256 |
0.188 |
| y |
-0.256 |
-31.307 |
-0.149 |
| z |
0.188 |
-0.149 |
-35.276 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.390 |
-0.256 |
0.188 |
| y |
-0.256 |
8.172 |
-0.149 |
| z |
0.188 |
-0.149 |
2.219 |
|
| Polar |
|---|
| 3z2-r2 | 4.437 |
|---|
| x2-y2 | -12.375 |
|---|
| xy | -0.256 |
|---|
| xz | 0.188 |
|---|
| yz | -0.149 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.287 |
0.385 |
-0.068 |
| y |
0.385 |
8.202 |
0.026 |
| z |
-0.068 |
0.026 |
6.194 |
<r2> (average value of r
2) Å
2
| <r2> |
144.727 |
| (<r2>)1/2 |
12.030 |