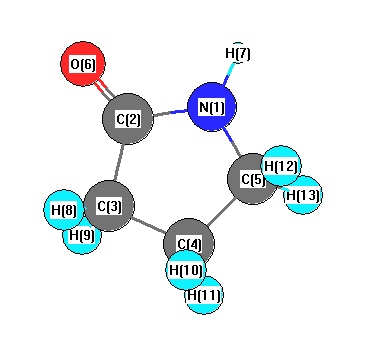
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3671 |
3513 |
33.86 |
|
|
|
| 2 |
A |
3154 |
3019 |
13.38 |
|
|
|
| 3 |
A |
3142 |
3007 |
19.18 |
|
|
|
| 4 |
A |
3103 |
2970 |
42.28 |
|
|
|
| 5 |
A |
3085 |
2953 |
14.25 |
|
|
|
| 6 |
A |
3075 |
2943 |
10.63 |
|
|
|
| 7 |
A |
3021 |
2891 |
63.10 |
|
|
|
| 8 |
A |
1840 |
1761 |
481.59 |
|
|
|
| 9 |
A |
1538 |
1472 |
3.88 |
|
|
|
| 10 |
A |
1498 |
1433 |
10.34 |
|
|
|
| 11 |
A |
1467 |
1404 |
4.04 |
|
|
|
| 12 |
A |
1458 |
1395 |
44.15 |
|
|
|
| 13 |
A |
1377 |
1318 |
10.08 |
|
|
|
| 14 |
A |
1343 |
1286 |
16.65 |
|
|
|
| 15 |
A |
1312 |
1255 |
44.10 |
|
|
|
| 16 |
A |
1278 |
1223 |
55.17 |
|
|
|
| 17 |
A |
1252 |
1199 |
7.87 |
|
|
|
| 18 |
A |
1217 |
1165 |
1.52 |
|
|
|
| 19 |
A |
1189 |
1138 |
4.36 |
|
|
|
| 20 |
A |
1098 |
1051 |
10.09 |
|
|
|
| 21 |
A |
1088 |
1041 |
6.35 |
|
|
|
| 22 |
A |
1018 |
974 |
11.11 |
|
|
|
| 23 |
A |
939 |
899 |
0.38 |
|
|
|
| 24 |
A |
911 |
872 |
2.29 |
|
|
|
| 25 |
A |
902 |
863 |
3.54 |
|
|
|
| 26 |
A |
822 |
786 |
4.88 |
|
|
|
| 27 |
A |
696 |
666 |
8.55 |
|
|
|
| 28 |
A |
641 |
614 |
19.96 |
|
|
|
| 29 |
A |
567 |
543 |
39.80 |
|
|
|
| 30 |
A |
494 |
473 |
52.89 |
|
|
|
| 31 |
A |
471 |
451 |
9.99 |
|
|
|
| 32 |
A |
197 |
189 |
3.26 |
|
|
|
| 33 |
A |
145 |
139 |
0.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24504.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23453.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.139 |
|
|
|
| 2 |
C |
0.181 |
|
|
|
| 3 |
C |
-0.230 |
|
|
|
| 4 |
C |
-0.238 |
|
|
|
| 5 |
C |
-0.114 |
|
|
|
| 6 |
O |
-0.335 |
|
|
|
| 7 |
H |
0.161 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
H |
0.130 |
|
|
|
| 10 |
H |
0.117 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.102 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.981 |
-0.651 |
0.341 |
4.048 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.618 |
-0.250 |
0.200 |
| y |
-0.250 |
-31.320 |
-0.151 |
| z |
0.200 |
-0.151 |
-35.293 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.311 |
-0.250 |
0.200 |
| y |
-0.250 |
8.135 |
-0.151 |
| z |
0.200 |
-0.151 |
2.176 |
|
| Polar |
|---|
| 3z2-r2 | 4.351 |
|---|
| x2-y2 | -12.298 |
|---|
| xy | -0.250 |
|---|
| xz | 0.200 |
|---|
| yz | -0.151 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.287 |
0.381 |
-0.072 |
| y |
0.381 |
8.201 |
0.029 |
| z |
-0.072 |
0.029 |
6.211 |
<r2> (average value of r
2) Å
2
| <r2> |
144.098 |
| (<r2>)1/2 |
12.004 |