return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (E)-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2h 1Ag
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-156.915320
Energy at 298.15K-156.923130
HF Energy-156.167984
Nuclear repulsion energy116.574321
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3191 3030 0.00      
2 A 3164 3004 0.00      
3 A 3081 2925 0.00      
4 A 1753 1664 0.00      
5 A 1516 1439 0.00      
6 A 1423 1351 0.00      
7 A 1339 1271 0.00      
8 A 1186 1126 0.00      
9 A 889 844 0.00      
10 A 503 478 0.00      
11 A 3141 2982 30.11      
12 A 1506 1430 12.86      
13 A 1082 1027 2.62      
14 A 1027 975 33.90      
15 A 253 240 3.22      
16 A 184 175 1.17      
17 A 3142 2983 0.00      
18 A 1507 1430 0.00      
19 A 1084 1029 0.00      
20 A 776 737 0.00      
21 A 238 226 0.00      
22 A 3194 3032 48.03      
23 A 3162 3002 17.24      
24 A 3081 2925 50.52      
25 A 1523 1446 18.82      
26 A 1423 1351 5.20      
27 A 1337 1270 4.06      
28 A 1110 1054 3.11      
29 A 994 944 11.82      
30 A 282 268 0.95      

Unscaled Zero Point Vibrational Energy (zpe) 24045.7 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 22829.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
1.17231 0.12600 0.11870

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.665 0.000
C2 0.000 -0.665 0.000
C3 -1.223 1.514 0.000
C4 1.223 -1.514 0.000
H5 0.951 1.177 0.000
H6 -0.951 -1.177 0.000
H7 -2.120 0.903 0.000
H8 2.120 -0.903 0.000
H9 -1.252 2.160 0.874
H10 -1.252 2.160 -0.874
H11 1.252 -2.160 0.874
H12 1.252 -2.160 -0.874

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.33001.48922.49911.08042.07312.13352.63702.13742.13743.21173.2117
C21.33002.49911.48922.07311.08042.63702.13353.21173.21172.13742.1374
C31.48922.49913.89322.20072.70481.08544.12571.08731.08734.51614.5161
C42.49911.48923.89322.70482.20074.12571.08544.51614.51611.08731.0873
H51.08042.07312.20072.70483.02673.08382.38572.56662.56663.46293.4629
H62.07311.08042.70482.20073.02672.38573.08383.46293.46292.56662.5666
H72.13352.63701.08544.12573.08382.38574.60891.76011.76014.63924.6392
H82.63702.13354.12571.08542.38573.08384.60894.63924.63921.76011.7601
H92.13743.21171.08734.51612.56663.46291.76014.63921.74784.99435.2913
H102.13743.21171.08734.51612.56663.46291.76014.63921.74785.29134.9943
H113.21172.13744.51611.08733.46292.56664.63921.76014.99435.29131.7478
H123.21172.13744.51611.08733.46292.56664.63921.76015.29134.99431.7478

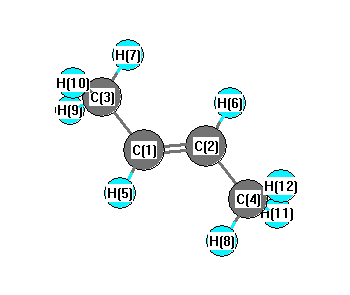
picture of 2-Butene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.769 C1 C2 H6 118.280
C1 C3 H7 110.957 C1 C3 H9 111.153
C1 C3 H10 111.153 C2 C1 C3 124.769
C2 C1 H5 118.280 C2 C4 H8 110.957
C2 C4 H11 111.153 C2 C4 H12 111.153
C3 C1 H5 116.951 C4 C2 H6 116.951
H7 C3 H9 108.220 H7 C3 H10 108.220
H8 C4 H11 108.220 H8 C4 H12 108.220
H9 C3 H10 106.980 H11 C4 H12 106.980
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability