Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.927991 |
| Energy at 298.15K | |
| HF Energy | -277.034316 |
| Nuclear repulsion energy | 125.836604 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3068 |
2991 |
|
|
|
|
| 2 |
Ag |
1526 |
1487 |
|
|
|
|
| 3 |
Ag |
1465 |
1428 |
|
|
|
|
| 4 |
Ag |
1109 |
1081 |
|
|
|
|
| 5 |
Ag |
1089 |
1061 |
|
|
|
|
| 6 |
Ag |
463 |
452 |
|
|
|
|
| 7 |
Au |
3140 |
3061 |
|
|
|
|
| 8 |
Au |
1250 |
1218 |
|
|
|
|
| 9 |
Au |
813 |
792 |
|
|
|
|
| 10 |
Au |
121 |
118 |
|
|
|
|
| 11 |
Bg |
3115 |
3036 |
|
|
|
|
| 12 |
Bg |
1310 |
1277 |
|
|
|
|
| 13 |
Bg |
1189 |
1159 |
|
|
|
|
| 14 |
Bu |
3073 |
2996 |
|
|
|
|
| 15 |
Bu |
1536 |
1497 |
|
|
|
|
| 16 |
Bu |
1371 |
1337 |
|
|
|
|
| 17 |
Bu |
1103 |
1075 |
|
|
|
|
| 18 |
Bu |
277 |
270 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13508.7 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 13168.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.628 |
0.000 |
| C2 |
-0.425 |
-0.628 |
0.000 |
| F3 |
-0.425 |
1.721 |
0.000 |
| F4 |
0.425 |
-1.721 |
0.000 |
| H5 |
1.053 |
0.668 |
0.891 |
| H6 |
1.053 |
0.668 |
-0.891 |
| H7 |
-1.053 |
-0.668 |
0.891 |
| H8 |
-1.053 |
-0.668 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5158 | 1.3844 | 2.3492 | 1.0908 | 1.0908 | 2.1578 | 2.1578 |
C2 | 1.5158 | | 2.3492 | 1.3844 | 2.1578 | 2.1578 | 1.0908 | 1.0908 | F3 | 1.3844 | 2.3492 | | 3.5459 | 2.0210 | 2.0210 | 2.6266 | 2.6266 | F4 | 2.3492 | 1.3844 | 3.5459 | | 2.6266 | 2.6266 | 2.0210 | 2.0210 | H5 | 1.0908 | 2.1578 | 2.0210 | 2.6266 | | 1.7818 | 2.4938 | 3.0649 | H6 | 1.0908 | 2.1578 | 2.0210 | 2.6266 | 1.7818 | | 3.0649 | 2.4938 | H7 | 2.1578 | 1.0908 | 2.6266 | 2.0210 | 2.4938 | 3.0649 | | 1.7818 | H8 | 2.1578 | 1.0908 | 2.6266 | 2.0210 | 3.0649 | 2.4938 | 1.7818 | |
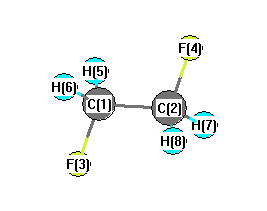 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.106 |
|
C1 |
C2 |
H7 |
110.690 |
| C1 |
C2 |
H8 |
110.690 |
|
C2 |
C1 |
F3 |
108.106 |
| C2 |
C1 |
H5 |
110.690 |
|
C2 |
C1 |
H6 |
110.690 |
| F3 |
C1 |
H5 |
108.891 |
|
F3 |
C1 |
H6 |
108.891 |
| F4 |
C2 |
H7 |
108.891 |
|
F4 |
C2 |
H8 |
108.891 |
| H5 |
C1 |
H6 |
109.523 |
|
H7 |
C2 |
H8 |
109.523 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.928987 |
| Energy at 298.15K | |
| HF Energy | -277.034691 |
| Nuclear repulsion energy | 127.976170 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3109 |
3030 |
|
|
|
|
| 2 |
A |
3060 |
2983 |
|
|
|
|
| 3 |
A |
1504 |
1466 |
|
|
|
|
| 4 |
A |
1456 |
1419 |
|
|
|
|
| 5 |
A |
1314 |
1281 |
|
|
|
|
| 6 |
A |
1146 |
1117 |
|
|
|
|
| 7 |
A |
1142 |
1113 |
|
|
|
|
| 8 |
A |
884 |
862 |
|
|
|
|
| 9 |
A |
323 |
315 |
|
|
|
|
| 10 |
A |
151 |
147 |
|
|
|
|
| 11 |
B |
3122 |
3043 |
|
|
|
|
| 12 |
B |
3050 |
2973 |
|
|
|
|
| 13 |
B |
1501 |
1464 |
|
|
|
|
| 14 |
B |
1418 |
1382 |
|
|
|
|
| 15 |
B |
1276 |
1244 |
|
|
|
|
| 16 |
B |
1114 |
1086 |
|
|
|
|
| 17 |
B |
919 |
896 |
|
|
|
|
| 18 |
B |
499 |
486 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13492.9 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 13152.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.265 |
0.704 |
0.511 |
| C2 |
-0.265 |
-0.704 |
0.511 |
| F3 |
-0.265 |
1.418 |
-0.547 |
| F4 |
0.265 |
-1.418 |
-0.547 |
| H5 |
-0.020 |
1.202 |
1.442 |
| H6 |
1.352 |
0.701 |
0.415 |
| H7 |
0.020 |
-1.202 |
1.442 |
| H8 |
-1.352 |
-0.701 |
0.415 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5048 | 1.3822 | 2.3716 | 1.0932 | 1.0914 | 2.1353 | 2.1443 |
C2 | 1.5048 | | 2.3716 | 1.3822 | 2.1353 | 2.1443 | 1.0932 | 1.0914 | F3 | 1.3822 | 2.3716 | | 2.8849 | 2.0160 | 2.0133 | 3.3019 | 2.5686 | F4 | 2.3716 | 1.3822 | 2.8849 | | 3.3019 | 2.5686 | 2.0160 | 2.0133 | H5 | 1.0932 | 2.1353 | 2.0160 | 3.3019 | | 1.7857 | 2.4041 | 2.5397 | H6 | 1.0914 | 2.1443 | 2.0133 | 2.5686 | 1.7857 | | 2.5397 | 3.0458 | H7 | 2.1353 | 1.0932 | 3.3019 | 2.0160 | 2.4041 | 2.5397 | | 1.7857 | H8 | 2.1443 | 1.0914 | 2.5686 | 2.0133 | 2.5397 | 3.0458 | 1.7857 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.394 |
|
C1 |
C2 |
H7 |
109.534 |
| C1 |
C2 |
H8 |
110.353 |
|
C2 |
C1 |
F3 |
110.394 |
| C2 |
C1 |
H5 |
109.534 |
|
C2 |
C1 |
H6 |
110.353 |
| F3 |
C1 |
H5 |
108.491 |
|
F3 |
C1 |
H6 |
108.380 |
| F4 |
C2 |
H7 |
108.491 |
|
F4 |
C2 |
H8 |
108.380 |
| H5 |
C1 |
H6 |
109.654 |
|
H7 |
C2 |
H8 |
109.654 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
