Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.934950 |
| Energy at 298.15K | |
| HF Energy | -277.034811 |
| Nuclear repulsion energy | 126.450120 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3127 |
2969 |
0.00 |
|
|
|
| 2 |
Ag |
1553 |
1474 |
0.00 |
|
|
|
| 3 |
Ag |
1484 |
1409 |
0.00 |
|
|
|
| 4 |
Ag |
1132 |
1075 |
0.00 |
|
|
|
| 5 |
Ag |
1104 |
1048 |
0.00 |
|
|
|
| 6 |
Ag |
470 |
446 |
0.00 |
|
|
|
| 7 |
Au |
3193 |
3031 |
40.10 |
|
|
|
| 8 |
Au |
1270 |
1206 |
4.18 |
|
|
|
| 9 |
Au |
829 |
787 |
0.03 |
|
|
|
| 10 |
Au |
136 |
129 |
12.77 |
|
|
|
| 11 |
Bg |
3167 |
3007 |
0.00 |
|
|
|
| 12 |
Bg |
1333 |
1266 |
0.00 |
|
|
|
| 13 |
Bg |
1203 |
1142 |
0.00 |
|
|
|
| 14 |
Bu |
3134 |
2975 |
50.85 |
|
|
|
| 15 |
Bu |
1563 |
1484 |
3.26 |
|
|
|
| 16 |
Bu |
1389 |
1319 |
8.25 |
|
|
|
| 17 |
Bu |
1118 |
1061 |
189.55 |
|
|
|
| 18 |
Bu |
281 |
267 |
18.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13743.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 13047.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.422 |
0.623 |
0.000 |
| C2 |
-0.422 |
-0.623 |
0.000 |
| F3 |
-0.422 |
1.715 |
0.000 |
| F4 |
0.422 |
-1.715 |
0.000 |
| H5 |
1.046 |
0.659 |
0.885 |
| H6 |
1.046 |
0.659 |
-0.885 |
| H7 |
-1.046 |
-0.659 |
0.885 |
| H8 |
-1.046 |
-0.659 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5042 | 1.3802 | 2.3371 | 1.0839 | 1.0839 | 2.1407 | 2.1407 |
C2 | 1.5042 | | 2.3371 | 1.3802 | 2.1407 | 2.1407 | 1.0839 | 1.0839 | F3 | 1.3802 | 2.3371 | | 3.5316 | 2.0131 | 2.0131 | 2.6095 | 2.6095 | F4 | 2.3371 | 1.3802 | 3.5316 | | 2.6095 | 2.6095 | 2.0131 | 2.0131 | H5 | 1.0839 | 2.1407 | 2.0131 | 2.6095 | | 1.7707 | 2.4733 | 3.0418 | H6 | 1.0839 | 2.1407 | 2.0131 | 2.6095 | 1.7707 | | 3.0418 | 2.4733 | H7 | 2.1407 | 1.0839 | 2.6095 | 2.0131 | 2.4733 | 3.0418 | | 1.7707 | H8 | 2.1407 | 1.0839 | 2.6095 | 2.0131 | 3.0418 | 2.4733 | 1.7707 | |
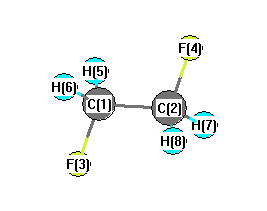 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.168 |
|
C1 |
C2 |
H7 |
110.566 |
| C1 |
C2 |
H8 |
110.566 |
|
C2 |
C1 |
F3 |
108.168 |
| C2 |
C1 |
H5 |
110.566 |
|
C2 |
C1 |
H6 |
110.566 |
| F3 |
C1 |
H5 |
108.977 |
|
F3 |
C1 |
H6 |
108.977 |
| F4 |
C2 |
H7 |
108.977 |
|
F4 |
C2 |
H8 |
108.977 |
| H5 |
C1 |
H6 |
109.544 |
|
H7 |
C2 |
H8 |
109.544 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.936187 |
| Energy at 298.15K | |
| HF Energy | -277.035154 |
| Nuclear repulsion energy | 128.637451 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3161 |
3001 |
11.77 |
|
|
|
| 2 |
A |
3119 |
2961 |
36.42 |
|
|
|
| 3 |
A |
1528 |
1451 |
0.23 |
|
|
|
| 4 |
A |
1474 |
1399 |
13.87 |
|
|
|
| 5 |
A |
1339 |
1271 |
1.78 |
|
|
|
| 6 |
A |
1165 |
1106 |
58.19 |
|
|
|
| 7 |
A |
1159 |
1100 |
25.26 |
|
|
|
| 8 |
A |
902 |
856 |
23.70 |
|
|
|
| 9 |
A |
330 |
313 |
0.34 |
|
|
|
| 10 |
A |
161 |
153 |
3.39 |
|
|
|
| 11 |
B |
3174 |
3014 |
31.31 |
|
|
|
| 12 |
B |
3110 |
2953 |
8.14 |
|
|
|
| 13 |
B |
1526 |
1449 |
7.87 |
|
|
|
| 14 |
B |
1434 |
1362 |
7.48 |
|
|
|
| 15 |
B |
1296 |
1230 |
6.19 |
|
|
|
| 16 |
B |
1126 |
1069 |
46.45 |
|
|
|
| 17 |
B |
928 |
881 |
36.43 |
|
|
|
| 18 |
B |
508 |
483 |
15.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13719.1 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 13024.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.261 |
0.699 |
0.512 |
| C2 |
-0.261 |
-0.699 |
0.512 |
| F3 |
-0.261 |
1.408 |
-0.548 |
| F4 |
0.261 |
-1.408 |
-0.548 |
| H5 |
-0.029 |
1.193 |
1.435 |
| H6 |
1.342 |
0.696 |
0.424 |
| H7 |
0.029 |
-1.193 |
1.435 |
| H8 |
-1.342 |
-0.696 |
0.424 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4925 | 1.3785 | 2.3586 | 1.0864 | 1.0846 | 2.1181 | 2.1274 |
C2 | 1.4925 | | 2.3586 | 1.3785 | 2.1181 | 2.1274 | 1.0864 | 1.0846 | F3 | 1.3785 | 2.3586 | | 2.8637 | 2.0082 | 2.0056 | 3.2839 | 2.5573 | F4 | 2.3586 | 1.3785 | 2.8637 | | 3.2839 | 2.5573 | 2.0082 | 2.0056 | H5 | 1.0864 | 2.1181 | 2.0082 | 3.2839 | | 1.7750 | 2.3876 | 2.5135 | H6 | 1.0846 | 2.1274 | 2.0056 | 2.5573 | 1.7750 | | 2.5135 | 3.0242 | H7 | 2.1181 | 1.0864 | 3.2839 | 2.0082 | 2.3876 | 2.5135 | | 1.7750 | H8 | 2.1274 | 1.0846 | 2.5573 | 2.0056 | 2.5135 | 3.0242 | 1.7750 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.418 |
|
C1 |
C2 |
H7 |
109.433 |
| C1 |
C2 |
H8 |
110.282 |
|
C2 |
C1 |
F3 |
110.418 |
| C2 |
C1 |
H5 |
109.433 |
|
C2 |
C1 |
H6 |
110.282 |
| F3 |
C1 |
H5 |
108.543 |
|
F3 |
C1 |
H6 |
108.440 |
| F4 |
C2 |
H7 |
108.543 |
|
F4 |
C2 |
H8 |
108.440 |
| H5 |
C1 |
H6 |
109.695 |
|
H7 |
C2 |
H8 |
109.695 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
