Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -278.307440 |
| Energy at 298.15K | |
| HF Energy | -278.307440 |
| Nuclear repulsion energy | 126.199638 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3060 |
2928 |
0.00 |
|
|
|
| 2 |
Ag |
1512 |
1447 |
0.00 |
|
|
|
| 3 |
Ag |
1447 |
1385 |
0.00 |
|
|
|
| 4 |
Ag |
1097 |
1050 |
0.00 |
|
|
|
| 5 |
Ag |
1087 |
1040 |
0.00 |
|
|
|
| 6 |
Ag |
460 |
440 |
0.00 |
|
|
|
| 7 |
Au |
3131 |
2997 |
51.33 |
|
|
|
| 8 |
Au |
1234 |
1181 |
4.00 |
|
|
|
| 9 |
Au |
810 |
776 |
0.00 |
|
|
|
| 10 |
Au |
112 |
107 |
12.19 |
|
|
|
| 11 |
Bg |
3104 |
2971 |
0.00 |
|
|
|
| 12 |
Bg |
1299 |
1244 |
0.00 |
|
|
|
| 13 |
Bg |
1175 |
1124 |
0.00 |
|
|
|
| 14 |
Bu |
3067 |
2935 |
60.19 |
|
|
|
| 15 |
Bu |
1523 |
1458 |
3.00 |
|
|
|
| 16 |
Bu |
1357 |
1299 |
7.40 |
|
|
|
| 17 |
Bu |
1103 |
1056 |
206.00 |
|
|
|
| 18 |
Bu |
272 |
260 |
17.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13424.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12848.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
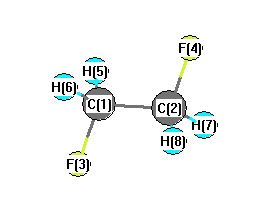
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.422 |
0.626 |
0.000 |
| C2 |
-0.422 |
-0.626 |
0.000 |
| F3 |
-0.422 |
1.717 |
0.000 |
| F4 |
0.422 |
-1.717 |
0.000 |
| H5 |
1.051 |
0.667 |
0.889 |
| H6 |
1.051 |
0.667 |
-0.889 |
| H7 |
-1.051 |
-0.667 |
0.889 |
| H8 |
-1.051 |
-0.667 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5098 | 1.3793 | 2.3435 | 1.0898 | 1.0898 | 2.1522 | 2.1522 |
C2 | 1.5098 | | 2.3435 | 1.3793 | 2.1522 | 2.1522 | 1.0898 | 1.0898 | F3 | 1.3793 | 2.3435 | | 3.5368 | 2.0152 | 2.0152 | 2.6218 | 2.6218 | F4 | 2.3435 | 1.3793 | 3.5368 | | 2.6218 | 2.6218 | 2.0152 | 2.0152 | H5 | 1.0898 | 2.1522 | 2.0152 | 2.6218 | | 1.7780 | 2.4898 | 3.0594 | H6 | 1.0898 | 2.1522 | 2.0152 | 2.6218 | 1.7780 | | 3.0594 | 2.4898 | H7 | 2.1522 | 1.0898 | 2.6218 | 2.0152 | 2.4898 | 3.0594 | | 1.7780 | H8 | 2.1522 | 1.0898 | 2.6218 | 2.0152 | 3.0594 | 2.4898 | 1.7780 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.333 |
|
C1 |
C2 |
H7 |
110.731 |
| C1 |
C2 |
H8 |
110.731 |
|
C2 |
C1 |
F3 |
108.333 |
| C2 |
C1 |
H5 |
110.731 |
|
C2 |
C1 |
H6 |
110.731 |
| F3 |
C1 |
H5 |
108.841 |
|
F3 |
C1 |
H6 |
108.841 |
| F4 |
C2 |
H7 |
108.841 |
|
F4 |
C2 |
H8 |
108.841 |
| H5 |
C1 |
H6 |
109.315 |
|
H7 |
C2 |
H8 |
109.315 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.066 |
|
|
|
| 2 |
C |
0.066 |
|
|
|
| 3 |
F |
-0.231 |
|
|
|
| 4 |
F |
-0.231 |
|
|
|
| 5 |
H |
0.082 |
|
|
|
| 6 |
H |
0.082 |
|
|
|
| 7 |
H |
0.082 |
|
|
|
| 8 |
H |
0.082 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.931 |
2.805 |
0.000 |
| y |
2.805 |
-28.698 |
0.000 |
| z |
0.000 |
0.000 |
-21.439 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.137 |
2.805 |
0.000 |
| y |
2.805 |
-7.013 |
0.000 |
| z |
0.000 |
0.000 |
3.876 |
|
| Polar |
|---|
| 3z2-r2 | 7.752 |
|---|
| x2-y2 | 6.767 |
|---|
| xy | 2.805 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.977 |
0.020 |
0.000 |
| y |
0.020 |
4.068 |
0.000 |
| z |
0.000 |
0.000 |
3.676 |
<r2> (average value of r
2) Å
2
| <r2> |
87.495 |
| (<r2>)1/2 |
9.354 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -278.308992 |
| Energy at 298.15K | |
| HF Energy | -278.308992 |
| Nuclear repulsion energy | 128.219296 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3100 |
2967 |
16.67 |
|
|
|
| 2 |
A |
3054 |
2923 |
37.15 |
|
|
|
| 3 |
A |
1484 |
1421 |
0.17 |
|
|
|
| 4 |
A |
1435 |
1373 |
13.01 |
|
|
|
| 5 |
A |
1305 |
1249 |
2.71 |
|
|
|
| 6 |
A |
1137 |
1088 |
84.93 |
|
|
|
| 7 |
A |
1127 |
1078 |
8.49 |
|
|
|
| 8 |
A |
879 |
841 |
20.95 |
|
|
|
| 9 |
A |
320 |
306 |
0.35 |
|
|
|
| 10 |
A |
147 |
141 |
3.55 |
|
|
|
| 11 |
B |
3114 |
2980 |
38.02 |
|
|
|
| 12 |
B |
3044 |
2913 |
10.16 |
|
|
|
| 13 |
B |
1483 |
1419 |
8.59 |
|
|
|
| 14 |
B |
1399 |
1339 |
7.36 |
|
|
|
| 15 |
B |
1264 |
1209 |
6.92 |
|
|
|
| 16 |
B |
1106 |
1059 |
56.38 |
|
|
|
| 17 |
B |
910 |
871 |
38.45 |
|
|
|
| 18 |
B |
491 |
470 |
15.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13398.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12823.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.266 |
0.700 |
0.504 |
| C2 |
-0.266 |
-0.700 |
0.504 |
| F3 |
-0.266 |
1.425 |
-0.541 |
| F4 |
0.266 |
-1.425 |
-0.541 |
| H5 |
-0.008 |
1.193 |
1.439 |
| H6 |
1.351 |
0.695 |
0.400 |
| H7 |
0.008 |
-1.193 |
1.439 |
| H8 |
-1.351 |
-0.695 |
0.400 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4968 | 1.3783 | 2.3675 | 1.0920 | 1.0904 | 2.1269 | 2.1376 |
C2 | 1.4968 | | 2.3675 | 1.3783 | 2.1269 | 2.1376 | 1.0920 | 1.0904 | F3 | 1.3783 | 2.3675 | | 2.8983 | 2.0103 | 2.0076 | 3.2936 | 2.5607 | F4 | 2.3675 | 1.3783 | 2.8983 | | 3.2936 | 2.5607 | 2.0103 | 2.0076 | H5 | 1.0920 | 2.1269 | 2.0103 | 3.2936 | | 1.7813 | 2.3862 | 2.5396 | H6 | 1.0904 | 2.1376 | 2.0076 | 2.5607 | 1.7813 | | 2.5396 | 3.0385 | H7 | 2.1269 | 1.0920 | 3.2936 | 2.0103 | 2.3862 | 2.5396 | | 1.7813 | H8 | 2.1376 | 1.0904 | 2.5607 | 2.0076 | 2.5396 | 3.0385 | 1.7813 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.797 |
|
C1 |
C2 |
H7 |
109.494 |
| C1 |
C2 |
H8 |
110.441 |
|
C2 |
C1 |
F3 |
110.797 |
| C2 |
C1 |
H5 |
109.494 |
|
C2 |
C1 |
H6 |
110.441 |
| F3 |
C1 |
H5 |
108.383 |
|
F3 |
C1 |
H6 |
108.265 |
| F4 |
C2 |
H7 |
108.383 |
|
F4 |
C2 |
H8 |
108.265 |
| H5 |
C1 |
H6 |
109.418 |
|
H7 |
C2 |
H8 |
109.418 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.080 |
|
|
|
| 2 |
C |
0.080 |
|
|
|
| 3 |
F |
-0.227 |
|
|
|
| 4 |
F |
-0.227 |
|
|
|
| 5 |
H |
0.066 |
|
|
|
| 6 |
H |
0.081 |
|
|
|
| 7 |
H |
0.066 |
|
|
|
| 8 |
H |
0.081 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.562 |
2.562 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.592 |
1.559 |
0.000 |
| y |
1.559 |
-25.886 |
0.000 |
| z |
0.000 |
0.000 |
-22.189 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.446 |
1.559 |
0.000 |
| y |
1.559 |
-3.995 |
0.000 |
| z |
0.000 |
0.000 |
1.550 |
|
| Polar |
|---|
| 3z2-r2 | 3.100 |
|---|
| x2-y2 | 4.294 |
|---|
| xy | 1.559 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.812 |
0.070 |
0.000 |
| y |
0.070 |
4.098 |
0.000 |
| z |
0.000 |
0.000 |
3.916 |
<r2> (average value of r
2) Å
2
| <r2> |
79.270 |
| (<r2>)1/2 |
8.903 |
