return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5NH2 (aniline)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-287.103096
Energy at 298.15K-287.111216
HF Energy-285.833381
Nuclear repulsion energy272.928679
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3597 3415 18.72      
2 A' 3255 3090 7.83      
3 A' 3227 3064 5.74      
4 A' 3203 3041 9.11      
5 A' 1683 1598 122.03      
6 A' 1660 1576 10.70      
7 A' 1545 1467 58.20      
8 A' 1322 1255 47.02      
9 A' 1212 1150 5.32      
10 A' 1059 1005 3.02      
11 A' 1027 975 0.27      
12 A' 979 929 0.03      
13 A' 916 870 10.52      
14 A' 839 797 3.64      
15 A' 786 747 76.57      
16 A' 713 677 22.92      
17 A' 652 619 266.69      
18 A' 535 507 0.66      
19 A' 516 490 51.45      
20 A' 226 215 5.00      
21 A" 3705 3518 16.64      
22 A" 3234 3071 23.72      
23 A" 3210 3047 0.94      
24 A" 1651 1567 0.92      
25 A" 1509 1433 0.92      
26 A" 1481 1406 8.26      
27 A" 1363 1294 0.02      
28 A" 1194 1133 0.75      
29 A" 1152 1093 2.72      
30 A" 1079 1025 4.58      
31 A" 971 922 0.00      
32 A" 851 808 0.03      
33 A" 628 596 0.24      
34 A" 420 399 0.33      
35 A" 378 359 0.13      
36 A" 269 255 21.13      

Unscaled Zero Point Vibrational Energy (zpe) 26022.6 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 24705.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.19011 0.08744 0.06001

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.001 0.932 0.000
C2 0.004 0.221 1.197
C3 0.004 -1.164 1.196
C4 0.004 -1.869 0.000
C5 0.004 -1.164 -1.196
C6 0.004 0.221 -1.197
N7 0.065 2.323 0.000
H8 0.012 0.759 2.131
H9 0.004 -1.693 2.134
H10 0.003 -2.944 0.000
H11 0.004 -1.693 -2.134
H12 0.012 0.759 -2.131
H13 -0.314 2.747 -0.829
H14 -0.314 2.747 0.829

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14
C11.39232.41302.80012.41301.39231.39272.13803.38273.87553.38272.13802.02052.0205
C21.39231.38542.40832.76522.39462.41961.07762.13103.38393.84233.37153.25382.5724
C32.41301.38541.38782.39162.76523.68702.13871.07712.14413.37183.84284.41583.9413
C42.80012.40831.38781.38782.40834.19183.38312.14161.07542.14163.38314.70024.7002
C52.41302.76522.39161.38781.38543.68703.84283.37182.14411.07712.13873.94134.4158
C61.39232.39462.76522.40831.38542.41963.37153.84233.38392.13101.07762.57243.2538
N71.39272.41963.68704.19183.68702.41962.64384.54805.26724.54802.64381.00541.0054
H82.13801.07762.13873.38313.84283.37152.64382.45184.27244.91994.26213.58052.3986
H93.38272.13101.07712.14163.37183.84234.54802.45182.47404.26874.91995.34744.6387
H103.87553.38392.14411.07542.14413.38395.26724.27242.47402.47404.27245.75975.7597
H113.38273.84233.37182.14161.07712.13104.54804.91994.26872.47402.45184.63875.3474
H122.13803.37153.84283.38312.13871.07762.64384.26214.91994.27242.45182.39863.5805
H132.02053.25384.41584.70023.94132.57241.00543.58055.34745.75974.63872.39861.6580
H142.02052.57243.94134.70024.41583.25381.00542.39864.63875.75975.34743.58051.6580

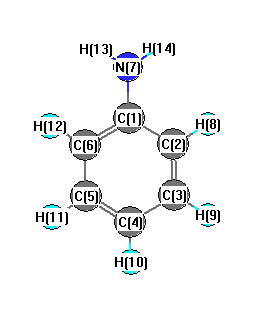
picture of aniline state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.623 C1 C2 H8 119.367
C1 C6 C5 120.623 C1 C6 H12 119.367
C1 N7 H13 113.833 C1 N7 H14 113.833
C2 C1 C6 118.628 C2 C1 N7 120.643
C2 C3 C4 120.559 C2 C3 H9 119.321
C3 C2 H8 120.009 C3 C4 C5 119.009
C3 C4 H10 120.496 C4 C3 H9 120.121
C4 C5 C6 120.559 C4 C5 H11 120.121
C5 C4 H10 120.496 C5 C6 H12 120.009
C6 C1 N7 120.643 C6 C5 H11 119.321
H13 N7 H14 111.077
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability