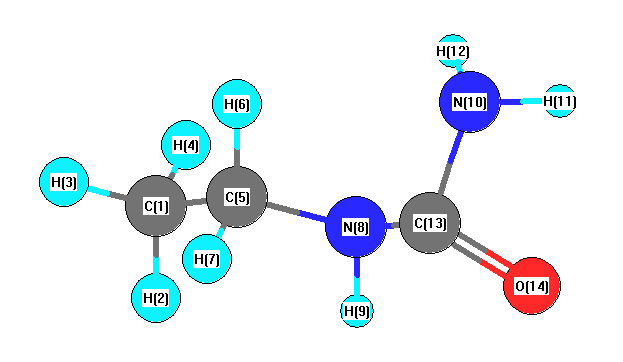
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3740 |
3579 |
39.16 |
|
|
|
| 2 |
A |
3673 |
3515 |
34.05 |
|
|
|
| 3 |
A |
3618 |
3463 |
21.38 |
|
|
|
| 4 |
A |
3149 |
3014 |
20.17 |
|
|
|
| 5 |
A |
3134 |
2999 |
26.53 |
|
|
|
| 6 |
A |
3062 |
2931 |
18.51 |
|
|
|
| 7 |
A |
3043 |
2912 |
26.35 |
|
|
|
| 8 |
A |
3010 |
2881 |
44.12 |
|
|
|
| 9 |
A |
1816 |
1738 |
538.86 |
|
|
|
| 10 |
A |
1620 |
1550 |
175.23 |
|
|
|
| 11 |
A |
1536 |
1470 |
4.05 |
|
|
|
| 12 |
A |
1505 |
1441 |
2.75 |
|
|
|
| 13 |
A |
1493 |
1429 |
7.50 |
|
|
|
| 14 |
A |
1465 |
1402 |
126.26 |
|
|
|
| 15 |
A |
1453 |
1390 |
150.63 |
|
|
|
| 16 |
A |
1414 |
1353 |
16.40 |
|
|
|
| 17 |
A |
1367 |
1308 |
18.00 |
|
|
|
| 18 |
A |
1312 |
1255 |
0.47 |
|
|
|
| 19 |
A |
1217 |
1165 |
19.17 |
|
|
|
| 20 |
A |
1172 |
1122 |
0.94 |
|
|
|
| 21 |
A |
1126 |
1078 |
17.94 |
|
|
|
| 22 |
A |
1074 |
1028 |
15.17 |
|
|
|
| 23 |
A |
1008 |
965 |
2.44 |
|
|
|
| 24 |
A |
906 |
867 |
2.68 |
|
|
|
| 25 |
A |
829 |
793 |
0.61 |
|
|
|
| 26 |
A |
783 |
749 |
36.20 |
|
|
|
| 27 |
A |
582 |
557 |
36.59 |
|
|
|
| 28 |
A |
552 |
529 |
14.25 |
|
|
|
| 29 |
A |
517 |
495 |
105.21 |
|
|
|
| 30 |
A |
459 |
440 |
21.28 |
|
|
|
| 31 |
A |
369 |
353 |
125.53 |
|
|
|
| 32 |
A |
358 |
343 |
3.78 |
|
|
|
| 33 |
A |
265 |
253 |
2.92 |
|
|
|
| 34 |
A |
198 |
190 |
4.14 |
|
|
|
| 35 |
A |
96 |
92 |
4.78 |
|
|
|
| 36 |
A |
68 |
65 |
1.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26493.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 25356.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.378 |
|
|
|
| 2 |
H |
0.123 |
|
|
|
| 3 |
H |
0.122 |
|
|
|
| 4 |
H |
0.126 |
|
|
|
| 5 |
C |
-0.121 |
|
|
|
| 6 |
H |
0.096 |
|
|
|
| 7 |
H |
0.117 |
|
|
|
| 8 |
N |
-0.154 |
|
|
|
| 9 |
H |
0.163 |
|
|
|
| 10 |
N |
-0.283 |
|
|
|
| 11 |
H |
0.176 |
|
|
|
| 12 |
H |
0.165 |
|
|
|
| 13 |
C |
0.223 |
|
|
|
| 14 |
O |
-0.374 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.307 |
2.714 |
-0.597 |
4.320 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.657 |
7.011 |
-1.266 |
| y |
7.011 |
-34.829 |
-1.466 |
| z |
-1.266 |
-1.466 |
-37.841 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.321 |
7.011 |
-1.266 |
| y |
7.011 |
3.420 |
-1.466 |
| z |
-1.266 |
-1.466 |
-1.098 |
|
| Polar |
|---|
| 3z2-r2 | -2.197 |
|---|
| x2-y2 | -3.827 |
|---|
| xy | 7.011 |
|---|
| xz | -1.266 |
|---|
| yz | -1.466 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.077 |
-0.107 |
0.093 |
| y |
-0.107 |
8.241 |
-0.043 |
| z |
0.093 |
-0.043 |
5.953 |
<r2> (average value of r
2) Å
2
| <r2> |
199.862 |
| (<r2>)1/2 |
14.137 |