return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: MP4=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP4=FULL/cc-pVTZ
 hartrees
Energy at 0K-531.568328
Energy at 298.15K 
HF Energy-530.696682
Nuclear repulsion energy155.301368
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3735 3597        
2 A 3601 3468        
3 A 3182 3064        
4 A 3106 2991        
5 A 3054 2941        
6 A 1652 1591        
7 A 1505 1449        
8 A 1500 1444        
9 A 1415 1363        
10 A 1395 1343        
11 A 1336 1286        
12 A 1040 1001        
13 A 1023 985        
14 A 998 961        
15 A 740 712        
16 A 620 597        
17 A 519 500        
18 A 421 406        
19 A 375 361        
20 A 255 246        
21 A 67 65        

Unscaled Zero Point Vibrational Energy (zpe) 15770.2 cm-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 15185.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4=FULL/cc-pVTZ
ABC
0.32760 0.16536 0.11218

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.276 0.057 -0.002
S2 -1.366 -0.110 -0.000
C3 1.227 -1.108 0.000
N4 0.888 1.257 0.003
H5 0.675 -2.038 -0.028
H6 1.889 -1.055 -0.864
H7 1.843 -1.082 0.899
H8 1.888 1.340 -0.014
H9 0.327 2.090 -0.002

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.65091.50371.34702.13292.13992.13632.05932.0335
S21.65092.77842.63602.80773.49753.47183.56182.7756
C31.50372.77842.38961.08201.09001.08992.53513.3223
N41.34702.63602.38963.30252.66452.68111.00341.0041
H52.13292.80771.08203.30251.77201.77163.58874.1428
H62.13993.49751.09002.66451.77201.76372.54063.6155
H72.13633.47181.08992.68111.77161.76372.58823.6297
H82.05933.56182.53511.00343.58872.54062.58821.7322
H92.03352.77563.32231.00414.14283.61553.62971.7322

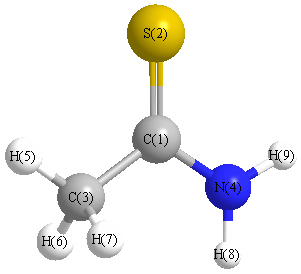
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.087 C1 C3 H6 110.167
C1 C3 H7 109.891 C1 N4 H8 121.675
C1 N4 H9 119.024 S2 C1 C3 123.401
S2 C1 N4 122.796 C3 C1 N4 113.802
H5 C3 H6 109.339 H5 C3 H7 109.313
H6 C3 H7 108.008 H8 N4 H9 119.277
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability