return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1-pentyne)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-194.912191
Energy at 298.15K-194.919482
HF Energy-194.003387
Nuclear repulsion energy154.767637
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3507 3330 62.16      
2 A' 3169 3009 23.16      
3 A' 3107 2950 23.02      
4 A' 3088 2932 18.60      
5 A' 3079 2924 10.22      
6 A' 2157 2048 3.16      
7 A' 1533 1455 6.68      
8 A' 1517 1440 0.84      
9 A' 1494 1419 1.93      
10 A' 1426 1354 2.83      
11 A' 1396 1325 1.18      
12 A' 1312 1245 4.54      
13 A' 1135 1077 1.70      
14 A' 1088 1033 0.26      
15 A' 993 942 1.52      
16 A' 898 852 2.84      
17 A' 650 617 42.56      
18 A' 507 482 5.50      
19 A' 343 326 1.14      
20 A' 155 147 0.36      
21 A" 3162 3002 39.31      
22 A" 3138 2979 3.22      
23 A" 3110 2952 1.82      
24 A" 1528 1451 7.80      
25 A" 1344 1276 0.01      
26 A" 1276 1212 0.04      
27 A" 1136 1078 0.00      
28 A" 882 837 0.80      
29 A" 755 717 1.62      
30 A" 651 618 43.04      
31 A" 348 331 6.45      
32 A" 259 246 0.04      
33 A" 104 99 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 25122.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 23851.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.79294 0.07529 0.07146

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.627 0.202 0.000
C2 1.436 0.416 0.000
C3 0.000 0.635 0.000
C4 -0.792 -0.667 0.000
C5 -2.286 -0.410 0.000
H6 3.667 0.018 0.000
H7 -0.274 1.225 0.873
H8 -0.274 1.225 -0.873
H9 -0.509 -1.250 0.872
H10 -0.509 -1.250 -0.872
H11 -2.847 -1.338 0.000
H12 -2.580 0.159 0.878
H13 -2.580 0.159 -0.878

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.21072.66273.52884.95151.05613.19793.19793.56503.56505.68755.28115.2811
C21.21071.45232.47753.81222.26682.08322.08322.70532.70534.62834.11864.1186
C32.66271.45231.52442.51353.71881.08871.08872.13842.13843.46422.76662.7666
C43.52882.47751.52441.51564.51202.14732.14731.08631.08632.16162.15612.1561
C54.95153.81222.51351.51565.96862.73532.73532.15012.15011.08481.08651.0865
H61.05612.26683.71884.51205.96864.21364.21364.45094.45096.65426.31036.3103
H73.19792.08321.08872.14732.73534.21361.74592.48613.03723.73522.54033.0851
H83.19792.08321.08872.14732.73534.21361.74593.03722.48613.73523.08512.5403
H93.56502.70532.13841.08632.15014.45092.48613.03721.74372.49702.50483.0553
H103.56502.70532.13841.08632.15014.45093.03722.48611.74372.49703.05532.5048
H115.68754.62833.46422.16161.08486.65423.73523.73522.49702.49701.75591.7559
H125.28114.11862.76662.15611.08656.31032.54033.08512.50483.05531.75591.7554
H135.28114.11862.76662.15611.08656.31033.08512.54033.05532.50481.75591.7554

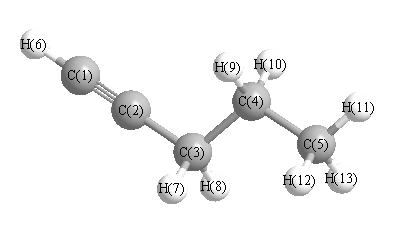
picture of 1-pentyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 178.517 C2 C1 H6 179.928
C2 C3 C4 112.651 C2 C3 H7 109.309
C2 C3 H8 109.309 C3 C4 C5 111.542
C3 C4 H9 108.832 C3 C4 H10 108.832
C4 C3 H7 109.387 C4 C3 H8 109.387
C4 C5 H11 111.384 C4 C5 H12 110.834
C4 C5 H13 110.834 C5 C4 H9 110.370
C5 C4 H10 110.370 H7 C3 H8 106.615
H9 C4 H10 106.757 H11 C5 H12 107.935
H11 C5 H13 107.935 H12 C5 H13 107.774
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability