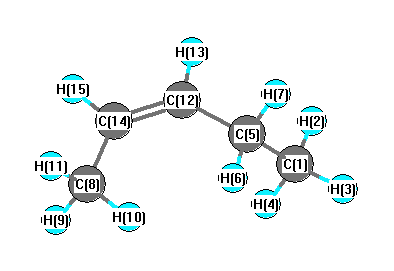
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3031 |
51.29 |
|
|
|
| 2 |
A |
3147 |
3012 |
9.35 |
|
|
|
| 3 |
A |
3138 |
3003 |
22.66 |
|
|
|
| 4 |
A |
3132 |
2997 |
29.64 |
|
|
|
| 5 |
A |
3127 |
2993 |
25.81 |
|
|
|
| 6 |
A |
3102 |
2969 |
3.24 |
|
|
|
| 7 |
A |
3094 |
2962 |
17.92 |
|
|
|
| 8 |
A |
3054 |
2923 |
32.33 |
|
|
|
| 9 |
A |
3047 |
2916 |
28.25 |
|
|
|
| 10 |
A |
3038 |
2908 |
25.12 |
|
|
|
| 11 |
A |
1754 |
1679 |
3.29 |
|
|
|
| 12 |
A |
1505 |
1441 |
5.59 |
|
|
|
| 13 |
A |
1496 |
1432 |
6.94 |
|
|
|
| 14 |
A |
1490 |
1426 |
5.14 |
|
|
|
| 15 |
A |
1482 |
1418 |
6.04 |
|
|
|
| 16 |
A |
1478 |
1415 |
0.92 |
|
|
|
| 17 |
A |
1441 |
1380 |
8.84 |
|
|
|
| 18 |
A |
1405 |
1344 |
1.73 |
|
|
|
| 19 |
A |
1399 |
1339 |
2.42 |
|
|
|
| 20 |
A |
1332 |
1275 |
4.55 |
|
|
|
| 21 |
A |
1297 |
1242 |
0.13 |
|
|
|
| 22 |
A |
1277 |
1222 |
0.12 |
|
|
|
| 23 |
A |
1173 |
1123 |
0.30 |
|
|
|
| 24 |
A |
1094 |
1047 |
3.92 |
|
|
|
| 25 |
A |
1063 |
1018 |
1.56 |
|
|
|
| 26 |
A |
1049 |
1004 |
3.48 |
|
|
|
| 27 |
A |
1042 |
997 |
0.06 |
|
|
|
| 28 |
A |
1017 |
974 |
0.51 |
|
|
|
| 29 |
A |
947 |
907 |
12.79 |
|
|
|
| 30 |
A |
874 |
837 |
1.54 |
|
|
|
| 31 |
A |
791 |
757 |
4.27 |
|
|
|
| 32 |
A |
722 |
691 |
32.03 |
|
|
|
| 33 |
A |
567 |
543 |
6.47 |
|
|
|
| 34 |
A |
471 |
451 |
1.32 |
|
|
|
| 35 |
A |
290 |
278 |
0.02 |
|
|
|
| 36 |
A |
253 |
242 |
0.02 |
|
|
|
| 37 |
A |
208 |
199 |
0.01 |
|
|
|
| 38 |
A |
137 |
131 |
0.35 |
|
|
|
| 39 |
A |
54 |
51 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30076.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 28786.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.349 |
|
|
|
| 2 |
H |
0.112 |
|
|
|
| 3 |
H |
0.115 |
|
|
|
| 4 |
H |
0.118 |
|
|
|
| 5 |
C |
-0.201 |
|
|
|
| 6 |
H |
0.111 |
|
|
|
| 7 |
H |
0.119 |
|
|
|
| 8 |
C |
-0.341 |
|
|
|
| 9 |
H |
0.119 |
|
|
|
| 10 |
H |
0.113 |
|
|
|
| 11 |
H |
0.120 |
|
|
|
| 12 |
C |
-0.147 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
C |
-0.143 |
|
|
|
| 15 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.016 |
-0.260 |
0.064 |
0.268 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.985 |
0.303 |
0.875 |
| y |
0.303 |
-32.616 |
-0.277 |
| z |
0.875 |
-0.277 |
-34.373 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.510 |
0.303 |
0.875 |
| y |
0.303 |
0.563 |
-0.277 |
| z |
0.875 |
-0.277 |
-2.073 |
|
| Polar |
|---|
| 3z2-r2 | -4.146 |
|---|
| x2-y2 | 0.631 |
|---|
| xy | 0.303 |
|---|
| xz | 0.875 |
|---|
| yz | -0.277 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.893 |
0.304 |
0.483 |
| y |
0.304 |
8.289 |
-0.113 |
| z |
0.483 |
-0.113 |
7.219 |
<r2> (average value of r
2) Å
2
| <r2> |
163.952 |
| (<r2>)1/2 |
12.804 |