Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -553.433693 |
| Energy at 298.15K | |
| HF Energy | -552.510978 |
| Nuclear repulsion energy | 199.938061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3296 |
3130 |
3.65 |
|
|
|
| 2 |
A |
3230 |
3066 |
2.06 |
|
|
|
| 3 |
A |
3210 |
3047 |
3.45 |
|
|
|
| 4 |
A |
1668 |
1584 |
2.93 |
|
|
|
| 5 |
A |
1444 |
1371 |
9.46 |
|
|
|
| 6 |
A |
1324 |
1257 |
0.32 |
|
|
|
| 7 |
A |
1074 |
1019 |
0.14 |
|
|
|
| 8 |
A |
997 |
946 |
0.00 |
|
|
|
| 9 |
A |
913 |
867 |
0.00 |
|
|
|
| 10 |
A |
742 |
704 |
0.54 |
|
|
|
| 11 |
A |
633 |
601 |
0.00 |
|
|
|
| 12 |
A |
411 |
390 |
0.51 |
|
|
|
| 13 |
A |
179 |
170 |
0.19 |
|
|
|
| 14 |
A |
15i |
14i |
0.00 |
|
|
|
| 15 |
B |
3297 |
3130 |
0.22 |
|
|
|
| 16 |
B |
3229 |
3066 |
3.12 |
|
|
|
| 17 |
B |
3210 |
3048 |
0.49 |
|
|
|
| 18 |
B |
1646 |
1563 |
121.01 |
|
|
|
| 19 |
B |
1435 |
1363 |
10.08 |
|
|
|
| 20 |
B |
1287 |
1222 |
12.00 |
|
|
|
| 21 |
B |
1041 |
988 |
24.43 |
|
|
|
| 22 |
B |
1004 |
953 |
51.75 |
|
|
|
| 23 |
B |
914 |
868 |
69.08 |
|
|
|
| 24 |
B |
763 |
725 |
38.13 |
|
|
|
| 25 |
B |
571 |
542 |
22.55 |
|
|
|
| 26 |
B |
387 |
367 |
4.25 |
|
|
|
| 27 |
B |
21i |
20i |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18934.5 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17976.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.636 |
| C2 |
0.011 |
1.338 |
-0.493 |
| C3 |
-0.011 |
-1.338 |
-0.493 |
| C4 |
0.000 |
2.613 |
-0.112 |
| C5 |
0.000 |
-2.613 |
-0.112 |
| H6 |
0.030 |
1.060 |
-1.534 |
| H7 |
-0.030 |
-1.060 |
-1.534 |
| H8 |
-0.037 |
3.395 |
-0.850 |
| H9 |
-0.006 |
2.905 |
0.924 |
| H10 |
0.037 |
-3.395 |
-0.850 |
| H11 |
0.006 |
-2.905 |
0.924 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7507 | 1.7507 | 2.7181 | 2.7181 | 2.4151 | 2.4151 | 3.7058 | 2.9197 | 3.7058 | 2.9197 |
C2 | 1.7507 | | 2.6766 | 1.3307 | 3.9698 | 1.0777 | 2.6151 | 2.0877 | 2.1130 | 4.7464 | 4.4741 | C3 | 1.7507 | 2.6766 | | 3.9698 | 1.3307 | 2.6151 | 1.0777 | 4.7464 | 4.4741 | 2.0877 | 2.1130 | C4 | 2.7181 | 1.3307 | 3.9698 | | 5.2265 | 2.1057 | 3.9393 | 1.0756 | 1.0767 | 6.0531 | 5.6151 | C5 | 2.7181 | 3.9698 | 1.3307 | 5.2265 | | 3.9393 | 2.1057 | 6.0531 | 5.6151 | 1.0756 | 1.0767 | H6 | 2.4151 | 1.0777 | 2.6151 | 2.1057 | 3.9393 | | 2.1216 | 2.4331 | 3.0738 | 4.5070 | 4.6659 | H7 | 2.4151 | 2.6151 | 1.0777 | 3.9393 | 2.1057 | 2.1216 | | 4.5070 | 4.6659 | 2.4331 | 3.0738 | H8 | 3.7058 | 2.0877 | 4.7464 | 1.0756 | 6.0531 | 2.4331 | 4.5070 | | 1.8410 | 6.7895 | 6.5453 | H9 | 2.9197 | 2.1130 | 4.4741 | 1.0767 | 5.6151 | 3.0738 | 4.6659 | 1.8410 | | 6.5453 | 5.8109 | H10 | 3.7058 | 4.7464 | 2.0877 | 6.0531 | 1.0756 | 4.5070 | 2.4331 | 6.7895 | 6.5453 | | 1.8410 | H11 | 2.9197 | 4.4741 | 2.1130 | 5.6151 | 1.0767 | 4.6659 | 3.0738 | 6.5453 | 5.8109 | 1.8410 | |
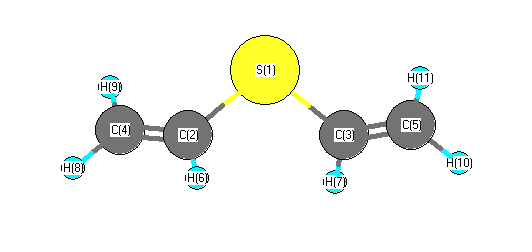 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.223 |
|
S1 |
C2 |
H6 |
115.199 |
| S1 |
C3 |
C5 |
123.223 |
|
S1 |
C3 |
H7 |
115.199 |
| C2 |
S1 |
C3 |
99.718 |
|
C2 |
C4 |
H8 |
119.990 |
| C2 |
C4 |
H9 |
122.378 |
|
C3 |
C5 |
H10 |
119.990 |
| C3 |
C5 |
H11 |
122.378 |
|
C4 |
C2 |
H6 |
121.578 |
| C5 |
C3 |
H7 |
121.578 |
|
H8 |
C4 |
H9 |
117.600 |
| H10 |
C5 |
H11 |
117.600 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -553.435735 |
| Energy at 298.15K | -553.441314 |
| HF Energy | -552.511936 |
| Nuclear repulsion energy | 203.730108 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3294 |
3127 |
3.03 |
|
|
|
| 2 |
A |
3292 |
3125 |
1.95 |
|
|
|
| 3 |
A |
3232 |
3068 |
2.93 |
|
|
|
| 4 |
A |
3218 |
3056 |
4.27 |
|
|
|
| 5 |
A |
3209 |
3047 |
0.74 |
|
|
|
| 6 |
A |
3200 |
3038 |
1.13 |
|
|
|
| 7 |
A |
1657 |
1573 |
32.53 |
|
|
|
| 8 |
A |
1646 |
1563 |
27.12 |
|
|
|
| 9 |
A |
1440 |
1367 |
4.89 |
|
|
|
| 10 |
A |
1436 |
1363 |
6.53 |
|
|
|
| 11 |
A |
1310 |
1244 |
2.43 |
|
|
|
| 12 |
A |
1297 |
1231 |
7.53 |
|
|
|
| 13 |
A |
1071 |
1017 |
5.56 |
|
|
|
| 14 |
A |
1043 |
990 |
15.16 |
|
|
|
| 15 |
A |
1019 |
967 |
23.02 |
|
|
|
| 16 |
A |
1004 |
954 |
34.51 |
|
|
|
| 17 |
A |
956 |
907 |
28.60 |
|
|
|
| 18 |
A |
915 |
868 |
36.56 |
|
|
|
| 19 |
A |
766 |
727 |
10.93 |
|
|
|
| 20 |
A |
719 |
683 |
4.95 |
|
|
|
| 21 |
A |
646 |
613 |
14.81 |
|
|
|
| 22 |
A |
623 |
591 |
12.60 |
|
|
|
| 23 |
A |
463 |
440 |
0.41 |
|
|
|
| 24 |
A |
371 |
352 |
0.51 |
|
|
|
| 25 |
A |
239 |
227 |
1.02 |
|
|
|
| 26 |
A |
160 |
151 |
3.85 |
|
|
|
| 27 |
A |
87 |
83 |
1.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19154.9 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18185.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.013 |
-0.984 |
0.009 |
| C2 |
1.054 |
0.375 |
0.366 |
| C3 |
-1.546 |
-0.195 |
-0.034 |
| C4 |
2.236 |
0.550 |
-0.221 |
| C5 |
-1.784 |
1.113 |
-0.124 |
| H6 |
0.712 |
1.025 |
1.155 |
| H7 |
-2.360 |
-0.898 |
0.024 |
| H8 |
2.893 |
1.338 |
0.106 |
| H9 |
2.565 |
-0.092 |
-1.018 |
| H10 |
-0.990 |
1.833 |
-0.215 |
| H11 |
-2.798 |
1.472 |
-0.118 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7487 | 1.7478 | 2.7107 | 2.7647 | 2.4165 | 2.3746 | 3.7005 | 2.8918 | 2.9987 | 3.7348 |
C2 | 1.7487 | | 2.6918 | 1.3313 | 2.9727 | 1.0781 | 3.6597 | 2.0920 | 2.1021 | 2.5770 | 4.0341 | C3 | 1.7478 | 2.6918 | | 3.8596 | 1.3328 | 2.8292 | 1.0770 | 4.6985 | 4.2287 | 2.1110 | 2.0865 | C4 | 2.7107 | 1.3313 | 3.8596 | | 4.0604 | 2.1074 | 4.8252 | 1.0764 | 1.0760 | 3.4723 | 5.1190 | C5 | 2.7647 | 2.9727 | 1.3328 | 4.0604 | | 2.8061 | 2.0976 | 4.6878 | 4.6008 | 1.0754 | 1.0758 | H6 | 2.4165 | 1.0781 | 2.8292 | 2.1074 | 2.8061 | | 3.7972 | 2.4401 | 3.0671 | 2.3298 | 3.7608 | H7 | 2.3746 | 3.6597 | 1.0770 | 4.8252 | 2.0976 | 3.7972 | | 5.7097 | 5.0982 | 3.0650 | 2.4145 | H8 | 3.7005 | 2.0920 | 4.6985 | 1.0764 | 4.6878 | 2.4401 | 5.7097 | | 1.8489 | 3.9277 | 5.6969 | H9 | 2.8918 | 2.1021 | 4.2287 | 1.0760 | 4.6008 | 3.0671 | 5.0982 | 1.8489 | | 4.1226 | 5.6587 | H10 | 2.9987 | 2.5770 | 2.1110 | 3.4723 | 1.0754 | 2.3298 | 3.0650 | 3.9277 | 4.1226 | | 1.8460 | H11 | 3.7348 | 4.0341 | 2.0865 | 5.1190 | 1.0758 | 3.7608 | 2.4145 | 5.6969 | 5.6587 | 1.8460 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.733 |
|
S1 |
C2 |
H6 |
115.429 |
| S1 |
C3 |
C5 |
127.130 |
|
S1 |
C3 |
H7 |
112.223 |
| C2 |
S1 |
C3 |
100.681 |
|
C2 |
C4 |
H8 |
120.283 |
| C2 |
C4 |
H9 |
121.307 |
|
C3 |
C5 |
H10 |
122.103 |
| C3 |
C5 |
H11 |
119.676 |
|
C4 |
C2 |
H6 |
121.656 |
| C5 |
C3 |
H7 |
120.648 |
|
H8 |
C4 |
H9 |
118.406 |
| H10 |
C5 |
H11 |
118.217 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
